return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H12O (1-Butanol, 2-methyl-)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-272.634704
Energy at 298.15K-272.648123
HF Energy-272.359338
Nuclear repulsion energy258.105217
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3780 3588 17.16      
2 A 3182 3020 16.88      
3 A 3158 2997 47.84      
4 A 3149 2989 39.67      
5 A 3139 2979 39.33      
6 A 3098 2941 23.01      
7 A 3081 2924 58.50      
8 A 3074 2918 64.46      
9 A 3069 2913 17.66      
10 A 3058 2903 19.45      
11 A 3052 2897 10.28      
12 A 3011 2858 47.65      
13 A 1572 1492 2.13      
14 A 1561 1481 5.21      
15 A 1557 1478 8.61      
16 A 1552 1473 4.00      
17 A 1545 1467 5.55      
18 A 1536 1458 1.29      
19 A 1493 1417 4.23      
20 A 1462 1388 2.64      
21 A 1451 1377 4.23      
22 A 1429 1356 1.30      
23 A 1398 1327 3.72      
24 A 1364 1294 2.14      
25 A 1334 1266 10.63      
26 A 1295 1230 2.22      
27 A 1278 1213 36.52      
28 A 1213 1151 0.48      
29 A 1202 1141 5.94      
30 A 1149 1090 5.59      
31 A 1092 1036 77.39      
32 A 1065 1011 1.34      
33 A 1047 994 20.76      
34 A 1001 950 5.10      
35 A 950 902 1.68      
36 A 934 886 1.26      
37 A 842 799 3.74      
38 A 779 740 5.15      
39 A 501 476 6.84      
40 A 450 427 1.62      
41 A 397 377 2.21      
42 A 286 271 4.71      
43 A 275 261 75.53      
44 A 252 240 38.73      
45 A 238 226 3.98      
46 A 224 213 2.15      
47 A 120 114 4.02      
48 A 75 71 4.53      

Unscaled Zero Point Vibrational Energy (zpe) 36883.1 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 35009.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.16556 0.07543 0.05641

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.799 1.653 0.019
H2 -1.821 1.604 -0.359
H3 -0.836 1.811 1.101
H4 -0.295 2.509 -0.437
O5 -2.130 -0.822 -0.207
H6 -2.598 -1.570 0.199
C7 -0.800 -0.824 0.313
H8 -0.815 -0.706 1.404
H9 -0.313 -1.771 0.071
C10 -0.039 0.355 -0.297
H11 -0.023 0.224 -1.383
C12 2.342 -0.723 -0.134
H13 3.359 -0.534 0.220
H14 2.001 -1.646 0.326
H15 2.383 -0.868 -1.216
C16 1.419 0.465 0.211
H17 1.853 1.363 -0.236
H18 1.405 0.608 1.296

Atom - Atom Distances (Å)
  C1 H2 H3 H4 O5 H6 C7 H8 H9 C10 H11 C12 H13 H14 H15 C16 H17 H18
C11.09051.09331.09322.81933.69562.49422.73553.45871.53692.14713.94174.70194.33804.24342.52372.67962.7528
H21.09051.77231.77542.45093.31582.71833.07543.72162.17672.48684.77465.63295.06344.95143.48133.68313.7595
H31.09331.77231.77343.21173.91722.75012.53503.76342.16953.05714.24854.88584.53874.78612.77323.03622.5510
H41.09321.77541.77343.81004.72743.45323.74114.31002.17382.48794.18234.79974.80824.37982.74542.44263.0830
O52.81932.45093.21173.81000.97161.42772.08312.06842.40062.62934.47385.51244.24544.62393.79794.54234.0985
H63.69563.31583.91724.72740.97161.94972.31892.29713.23993.51425.02336.04584.60065.22474.50275.34754.6869
C72.49422.71832.75013.45321.42771.94971.09801.09221.52932.13903.17574.16972.91893.53092.56793.48102.8069
H82.73553.07542.53503.74112.08312.31891.09801.77892.14913.04273.51214.34153.15794.13692.78983.75282.5817
H93.45873.72163.76344.31002.06842.29711.09221.77892.17462.48512.86203.87722.33093.12012.83143.82133.1799
C101.53692.17672.16952.17382.40063.23991.52932.14912.17461.09422.61933.54992.92472.86491.54812.14442.1645
H112.14712.48683.05712.48792.62933.51422.13903.04272.48511.09422.83753.81823.24262.64752.16302.47583.0597
C123.94174.77464.24854.18234.47385.02333.17573.51212.86202.61932.83751.09261.08711.09211.54342.14462.1670
H134.70195.63294.88584.79975.51246.04584.16974.34153.87723.54993.81821.09261.75891.76802.18142.46412.5058
H144.33805.06344.53874.80824.24544.60062.91893.15792.33092.92473.24261.08711.75891.76892.19263.06462.5255
H154.24344.95144.78614.37984.62395.22473.53094.13693.12012.86492.64751.09211.76801.76892.17762.49373.0733
C162.52373.48132.77322.74543.79794.50272.56792.78982.83141.54812.16301.54342.18142.19262.17761.09291.0942
H172.67963.68313.03622.44264.54235.34753.48103.75283.82132.14442.47582.14462.46413.06462.49371.09291.7653
H182.75283.75952.55103.08304.09854.68692.80692.58173.17992.16453.05972.16702.50582.52553.07331.09421.7653

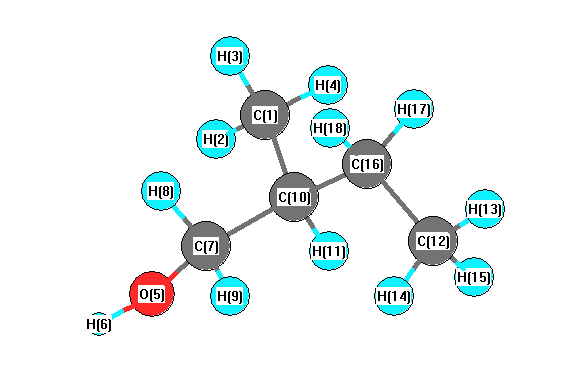
picture of 1-Butanol, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C10 C7 108.864 C1 C10 H11 108.200
C1 C10 C16 109.778 H2 C1 H3 108.506
H2 C1 H4 108.790 H2 C1 C10 110.738
H3 C1 H4 108.403 H3 C1 C10 109.999
H4 C1 C10 110.345 O5 C7 H8 110.451
O5 C7 H9 109.614 O5 C7 C10 108.499
H6 O5 C7 107.168 C7 C10 H11 108.089
C7 C10 C16 113.114 H8 C7 H9 108.623
H8 C7 C10 108.654 H9 C7 C10 110.995
C10 C16 C12 115.828 C10 C16 H17 107.320
C10 C16 H18 108.793 H11 C10 C16 108.671
C12 C16 H17 107.650 C12 C16 H18 109.306
H13 C12 H14 107.594 H13 C12 H15 108.049
H13 C12 C16 110.529 H14 C12 H15 108.531
H14 C12 C16 111.755 H15 C12 C16 110.262
H17 C16 H18 107.641
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.465      
2 H 0.182      
3 H 0.138      
4 H 0.145      
5 O -0.677      
6 H 0.410      
7 C -0.010      
8 H 0.122      
9 H 0.132      
10 C -0.118      
11 H 0.148      
12 C -0.467      
13 H 0.153      
14 H 0.149      
15 H 0.152      
16 C -0.279      
17 H 0.147      
18 H 0.138      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 9.237 0.036 -0.022
y 0.036 8.444 -0.121
z -0.022 -0.121 7.521


<r2> (average value of r2) Å2
<r2> 215.768
(<r2>)1/2 14.689