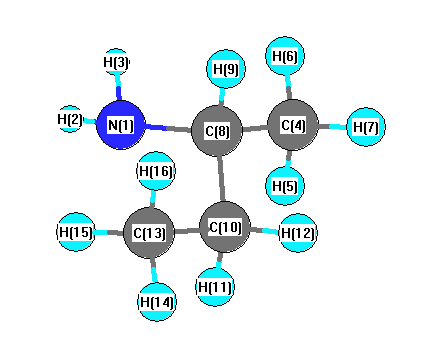
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3578 |
3423 |
0.47 |
|
|
|
| 2 |
A |
3490 |
3339 |
2.45 |
|
|
|
| 3 |
A |
3142 |
3006 |
35.24 |
|
|
|
| 4 |
A |
3130 |
2994 |
40.54 |
|
|
|
| 5 |
A |
3123 |
2988 |
50.11 |
|
|
|
| 6 |
A |
3123 |
2987 |
25.45 |
|
|
|
| 7 |
A |
3091 |
2957 |
9.55 |
|
|
|
| 8 |
A |
3053 |
2920 |
30.70 |
|
|
|
| 9 |
A |
3048 |
2916 |
37.32 |
|
|
|
| 10 |
A |
3047 |
2915 |
16.15 |
|
|
|
| 11 |
A |
2925 |
2798 |
83.17 |
|
|
|
| 12 |
A |
1698 |
1624 |
40.49 |
|
|
|
| 13 |
A |
1536 |
1469 |
8.32 |
|
|
|
| 14 |
A |
1526 |
1460 |
3.02 |
|
|
|
| 15 |
A |
1523 |
1457 |
5.04 |
|
|
|
| 16 |
A |
1516 |
1450 |
3.48 |
|
|
|
| 17 |
A |
1500 |
1435 |
0.67 |
|
|
|
| 18 |
A |
1442 |
1379 |
6.18 |
|
|
|
| 19 |
A |
1430 |
1368 |
8.27 |
|
|
|
| 20 |
A |
1426 |
1364 |
8.17 |
|
|
|
| 21 |
A |
1402 |
1341 |
14.02 |
|
|
|
| 22 |
A |
1348 |
1289 |
4.11 |
|
|
|
| 23 |
A |
1314 |
1257 |
1.69 |
|
|
|
| 24 |
A |
1273 |
1218 |
0.36 |
|
|
|
| 25 |
A |
1209 |
1156 |
1.56 |
|
|
|
| 26 |
A |
1175 |
1124 |
12.48 |
|
|
|
| 27 |
A |
1079 |
1033 |
1.50 |
|
|
|
| 28 |
A |
1043 |
998 |
2.88 |
|
|
|
| 29 |
A |
1028 |
983 |
6.86 |
|
|
|
| 30 |
A |
991 |
948 |
3.86 |
|
|
|
| 31 |
A |
957 |
916 |
13.93 |
|
|
|
| 32 |
A |
887 |
849 |
116.85 |
|
|
|
| 33 |
A |
822 |
786 |
3.79 |
|
|
|
| 34 |
A |
775 |
741 |
1.23 |
|
|
|
| 35 |
A |
479 |
459 |
8.18 |
|
|
|
| 36 |
A |
454 |
435 |
5.62 |
|
|
|
| 37 |
A |
373 |
357 |
0.83 |
|
|
|
| 38 |
A |
299 |
286 |
41.27 |
|
|
|
| 39 |
A |
256 |
245 |
0.17 |
|
|
|
| 40 |
A |
231 |
221 |
6.22 |
|
|
|
| 41 |
A |
224 |
214 |
11.41 |
|
|
|
| 42 |
A |
102 |
98 |
1.91 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 33031.8 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 31601.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.748 |
|
|
-0.945 |
| 2 |
H |
0.313 |
|
|
0.336 |
| 3 |
H |
0.309 |
|
|
0.357 |
| 4 |
C |
-0.508 |
|
|
-0.486 |
| 5 |
H |
0.179 |
|
|
0.137 |
| 6 |
H |
0.156 |
|
|
0.129 |
| 7 |
H |
0.164 |
|
|
0.101 |
| 8 |
C |
-0.004 |
|
|
0.407 |
| 9 |
H |
0.127 |
|
|
-0.040 |
| 10 |
C |
-0.284 |
|
|
0.037 |
| 11 |
H |
0.168 |
|
|
0.028 |
| 12 |
H |
0.152 |
|
|
0.003 |
| 13 |
C |
-0.526 |
|
|
-0.273 |
| 14 |
H |
0.171 |
|
|
0.074 |
| 15 |
H |
0.169 |
|
|
0.080 |
| 16 |
H |
0.161 |
|
|
0.056 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.018 |
0.206 |
1.283 |
1.299 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.006 |
0.228 |
1.264 |
1.285 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.636 |
0.388 |
1.046 |
| y |
0.388 |
-31.737 |
2.228 |
| z |
1.046 |
2.228 |
-35.168 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.816 |
0.388 |
1.046 |
| y |
0.388 |
2.165 |
2.228 |
| z |
1.046 |
2.228 |
-2.982 |
|
| Polar |
|---|
| 3z2-r2 | -5.963 |
|---|
| x2-y2 | -0.899 |
|---|
| xy | 0.388 |
|---|
| xz | 1.046 |
|---|
| yz | 2.228 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
150.593 |
| (<r2>)1/2 |
12.272 |