Jump to
S1C2
S1C3
Vibrational Frequencies calculated at CID/6-31G*
Geometric Data calculated at CID/6-31G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Vibrational Frequencies calculated at CID/6-31G*
Geometric Data calculated at CID/6-31G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -209.651600 |
| Energy at 298.15K | -209.660899 |
| HF Energy | -209.098571 |
| Nuclear repulsion energy | 135.055334 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3865 |
3570 |
68.78 |
|
|
|
| 2 |
A |
3716 |
3433 |
2.78 |
|
|
|
| 3 |
A |
3625 |
3349 |
0.88 |
|
|
|
| 4 |
A |
3219 |
2973 |
49.21 |
|
|
|
| 5 |
A |
3209 |
2964 |
27.67 |
|
|
|
| 6 |
A |
3120 |
2882 |
71.75 |
|
|
|
| 7 |
A |
3098 |
2862 |
62.41 |
|
|
|
| 8 |
A |
1768 |
1633 |
38.96 |
|
|
|
| 9 |
A |
1614 |
1491 |
0.06 |
|
|
|
| 10 |
A |
1595 |
1473 |
3.67 |
|
|
|
| 11 |
A |
1536 |
1418 |
73.92 |
|
|
|
| 12 |
A |
1493 |
1379 |
25.29 |
|
|
|
| 13 |
A |
1456 |
1345 |
9.48 |
|
|
|
| 14 |
A |
1403 |
1296 |
2.60 |
|
|
|
| 15 |
A |
1328 |
1227 |
29.72 |
|
|
|
| 16 |
A |
1255 |
1159 |
7.80 |
|
|
|
| 17 |
A |
1191 |
1100 |
66.96 |
|
|
|
| 18 |
A |
1137 |
1050 |
27.24 |
|
|
|
| 19 |
A |
1069 |
988 |
21.67 |
|
|
|
| 20 |
A |
1005 |
928 |
80.98 |
|
|
|
| 21 |
A |
934 |
863 |
13.64 |
|
|
|
| 22 |
A |
891 |
823 |
81.50 |
|
|
|
| 23 |
A |
578 |
534 |
120.62 |
|
|
|
| 24 |
A |
548 |
506 |
62.73 |
|
|
|
| 25 |
A |
346 |
320 |
0.50 |
|
|
|
| 26 |
A |
283 |
262 |
15.07 |
|
|
|
| 27 |
A |
204 |
188 |
5.65 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22743.0 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 21007.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
-1.358 |
-0.560 |
0.121 |
| C2 |
-0.631 |
0.646 |
-0.277 |
| C3 |
0.782 |
0.552 |
0.274 |
| O4 |
1.408 |
-0.626 |
-0.171 |
| H5 |
-1.652 |
-0.491 |
1.087 |
| H6 |
-2.195 |
-0.678 |
-0.431 |
| H7 |
-1.104 |
1.579 |
0.048 |
| H8 |
-0.577 |
0.660 |
-1.365 |
| H9 |
1.383 |
1.393 |
-0.067 |
| H10 |
0.750 |
0.591 |
1.370 |
| H11 |
0.733 |
-1.313 |
-0.102 |
Atom - Atom Distances (Å)
| |
N1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| N1 | | 1.4631 | 2.4160 | 2.7823 | 1.0117 | 1.0100 | 2.1550 | 2.0750 | 3.3710 | 2.7074 | 2.2336 |
C2 | 1.4631 | | 1.5188 | 2.4051 | 2.0477 | 2.0549 | 1.0957 | 1.0892 | 2.1584 | 2.1502 | 2.3928 | C3 | 2.4160 | 1.5188 | | 1.4060 | 2.7696 | 3.2971 | 2.1594 | 2.1311 | 1.0889 | 1.0974 | 1.9026 | O4 | 2.7823 | 2.4051 | 1.4060 | | 3.3115 | 3.6132 | 3.3497 | 2.6491 | 2.0211 | 2.0706 | 0.9657 | H5 | 1.0117 | 2.0477 | 2.7696 | 3.3115 | | 1.6230 | 2.3794 | 2.9135 | 3.7542 | 2.6502 | 2.7889 | H6 | 1.0100 | 2.0549 | 3.2971 | 3.6132 | 1.6230 | | 2.5517 | 2.2977 | 4.1504 | 3.6786 | 3.0145 | H7 | 2.1550 | 1.0957 | 2.1594 | 3.3497 | 2.3794 | 2.5517 | | 1.7656 | 2.4974 | 2.4830 | 3.4292 | H8 | 2.0750 | 1.0892 | 2.1311 | 2.6491 | 2.9135 | 2.2977 | 1.7656 | | 2.4622 | 3.0406 | 2.6837 | H9 | 3.3710 | 2.1584 | 1.0889 | 2.0211 | 3.7542 | 4.1504 | 2.4974 | 2.4622 | | 1.7635 | 2.7827 | H10 | 2.7074 | 2.1502 | 1.0974 | 2.0706 | 2.6502 | 3.6786 | 2.4830 | 3.0406 | 1.7635 | | 2.4063 | H11 | 2.2336 | 2.3928 | 1.9026 | 0.9657 | 2.7889 | 3.0145 | 3.4292 | 2.6837 | 2.7827 | 2.4063 | |
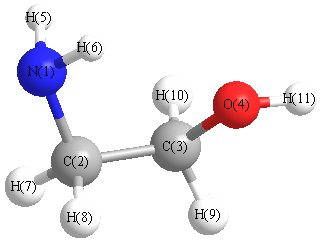 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C2 |
C3 |
108.223 |
|
N1 |
C2 |
H7 |
113.977 |
| N1 |
C2 |
H8 |
107.882 |
|
C2 |
N1 |
H5 |
110.341 |
| C2 |
N1 |
H6 |
111.056 |
|
C2 |
C3 |
O4 |
110.574 |
| C2 |
C3 |
H9 |
110.648 |
|
C2 |
C3 |
H10 |
109.488 |
| C3 |
C2 |
H7 |
110.317 |
|
C3 |
C2 |
H8 |
108.474 |
| C3 |
O4 |
H11 |
105.168 |
|
O4 |
C3 |
H9 |
107.535 |
| O4 |
C3 |
H10 |
111.008 |
|
H5 |
N1 |
H6 |
106.792 |
| H7 |
C2 |
H8 |
107.818 |
|
H9 |
C3 |
H10 |
107.531 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
