Jump to
S1C2
S1C3
Vibrational Frequencies calculated at B3PW91/6-31G*
Geometric Data calculated at B3PW91/6-31G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Vibrational Frequencies calculated at B3PW91/6-31G*
Geometric Data calculated at B3PW91/6-31G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at B3PW91/6-31G*
| | hartrees |
|---|
| Energy at 0K | -210.303269 |
| Energy at 298.15K | -210.312480 |
| HF Energy | -210.303269 |
| Nuclear repulsion energy | 134.973352 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3634 |
3477 |
80.16 |
|
|
|
| 2 |
A |
3603 |
3447 |
1.69 |
|
|
|
| 3 |
A |
3509 |
3357 |
0.38 |
|
|
|
| 4 |
A |
3109 |
2975 |
44.60 |
|
|
|
| 5 |
A |
3097 |
2963 |
36.53 |
|
|
|
| 6 |
A |
3006 |
2876 |
79.62 |
|
|
|
| 7 |
A |
2958 |
2830 |
85.88 |
|
|
|
| 8 |
A |
1695 |
1622 |
33.40 |
|
|
|
| 9 |
A |
1541 |
1475 |
0.14 |
|
|
|
| 10 |
A |
1520 |
1454 |
7.16 |
|
|
|
| 11 |
A |
1475 |
1411 |
104.10 |
|
|
|
| 12 |
A |
1423 |
1361 |
10.98 |
|
|
|
| 13 |
A |
1392 |
1331 |
2.36 |
|
|
|
| 14 |
A |
1341 |
1283 |
2.64 |
|
|
|
| 15 |
A |
1270 |
1215 |
25.77 |
|
|
|
| 16 |
A |
1207 |
1155 |
5.88 |
|
|
|
| 17 |
A |
1134 |
1085 |
67.09 |
|
|
|
| 18 |
A |
1089 |
1042 |
27.46 |
|
|
|
| 19 |
A |
1020 |
976 |
10.09 |
|
|
|
| 20 |
A |
949 |
908 |
72.02 |
|
|
|
| 21 |
A |
895 |
856 |
8.39 |
|
|
|
| 22 |
A |
842 |
806 |
88.21 |
|
|
|
| 23 |
A |
618 |
591 |
140.39 |
|
|
|
| 24 |
A |
546 |
522 |
9.79 |
|
|
|
| 25 |
A |
330 |
316 |
0.64 |
|
|
|
| 26 |
A |
284 |
272 |
15.09 |
|
|
|
| 27 |
A |
193 |
185 |
7.33 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21840.1 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 20894.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
-1.339 |
-0.562 |
0.118 |
| C2 |
-0.632 |
0.665 |
-0.261 |
| C3 |
0.800 |
0.556 |
0.259 |
| O4 |
1.380 |
-0.652 |
-0.168 |
| H5 |
-1.632 |
-0.516 |
1.092 |
| H6 |
-2.181 |
-0.686 |
-0.437 |
| H7 |
-1.105 |
1.594 |
0.097 |
| H8 |
-0.604 |
0.705 |
-1.356 |
| H9 |
1.411 |
1.383 |
-0.121 |
| H10 |
0.794 |
0.634 |
1.363 |
| H11 |
0.642 |
-1.289 |
-0.107 |
Atom - Atom Distances (Å)
| |
N1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| N1 | | 1.4659 | 2.4181 | 2.7361 | 1.0175 | 1.0155 | 2.1687 | 2.0779 | 3.3771 | 2.7445 | 2.1225 |
C2 | 1.4659 | | 1.5274 | 2.4064 | 2.0553 | 2.0623 | 1.1027 | 1.0959 | 2.1701 | 2.1614 | 2.3372 | C3 | 2.4181 | 1.5274 | | 1.4058 | 2.7854 | 3.3032 | 2.1760 | 2.1448 | 1.0961 | 1.1069 | 1.8870 | O4 | 2.7361 | 2.4064 | 1.4058 | | 3.2676 | 3.5715 | 3.3603 | 2.6813 | 2.0353 | 2.0828 | 0.9769 | H5 | 1.0175 | 2.0553 | 2.7854 | 3.2676 | | 1.6329 | 2.3915 | 2.9219 | 3.7865 | 2.6987 | 2.6839 | H6 | 1.0155 | 2.0623 | 3.3032 | 3.5715 | 1.6329 | | 2.5764 | 2.2942 | 4.1569 | 3.7190 | 2.9055 | H7 | 2.1687 | 1.1027 | 2.1760 | 3.3603 | 2.3915 | 2.5764 | | 1.7758 | 2.5348 | 2.4763 | 3.3771 | H8 | 2.0779 | 1.0959 | 2.1448 | 2.6813 | 2.9219 | 2.2942 | 1.7758 | | 2.4586 | 3.0580 | 2.6621 | H9 | 3.3771 | 2.1701 | 1.0961 | 2.0353 | 3.7865 | 4.1569 | 2.5348 | 2.4586 | | 1.7730 | 2.7800 | H10 | 2.7445 | 2.1614 | 1.1069 | 2.0828 | 2.6987 | 3.7190 | 2.4763 | 3.0580 | 1.7730 | | 2.4246 | H11 | 2.1225 | 2.3372 | 1.8870 | 0.9769 | 2.6839 | 2.9055 | 3.3771 | 2.6621 | 2.7800 | 2.4246 | |
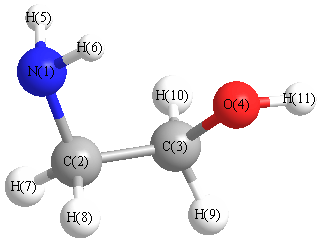 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C2 |
C3 |
107.752 |
|
N1 |
C2 |
H7 |
114.456 |
| N1 |
C2 |
H8 |
107.528 |
|
C2 |
N1 |
H5 |
110.397 |
| C2 |
N1 |
H6 |
111.120 |
|
C2 |
C3 |
O4 |
110.181 |
| C2 |
C3 |
H9 |
110.547 |
|
C2 |
C3 |
H10 |
109.228 |
| C3 |
C2 |
H7 |
110.614 |
|
C3 |
C2 |
H8 |
108.576 |
| C3 |
O4 |
H11 |
103.258 |
|
O4 |
C3 |
H9 |
108.241 |
| O4 |
C3 |
H10 |
111.418 |
|
H5 |
N1 |
H6 |
106.867 |
| H7 |
C2 |
H8 |
107.742 |
|
H9 |
C3 |
H10 |
107.182 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.776 |
|
|
|
| 2 |
C |
-0.227 |
|
|
|
| 3 |
C |
-0.050 |
|
|
|
| 4 |
O |
-0.639 |
|
|
|
| 5 |
H |
0.324 |
|
|
|
| 6 |
H |
0.331 |
|
|
|
| 7 |
H |
0.147 |
|
|
|
| 8 |
H |
0.180 |
|
|
|
| 9 |
H |
0.162 |
|
|
|
| 10 |
H |
0.129 |
|
|
|
| 11 |
H |
0.421 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.110 |
1.097 |
0.574 |
3.347 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.589 |
2.177 |
0.268 |
| y |
2.177 |
-25.216 |
-0.340 |
| z |
0.268 |
-0.340 |
-24.166 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.898 |
2.177 |
0.268 |
| y |
2.177 |
-0.339 |
-0.340 |
| z |
0.268 |
-0.340 |
1.237 |
|
| Polar |
|---|
| 3z2-r2 | 2.473 |
|---|
| x2-y2 | -0.373 |
|---|
| xy | 2.177 |
|---|
| xz | 0.268 |
|---|
| yz | -0.340 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.164 |
0.272 |
-0.009 |
| y |
0.272 |
4.932 |
0.054 |
| z |
-0.009 |
0.054 |
4.562 |
<r2> (average value of r
2) Å
2
| <r2> |
85.795 |
| (<r2>)1/2 |
9.263 |
