Jump to
S1C2
S1C3
Vibrational Frequencies calculated at QCISD/6-31G*
Geometric Data calculated at QCISD/6-31G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Vibrational Frequencies calculated at QCISD/6-31G*
Geometric Data calculated at QCISD/6-31G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at QCISD/6-31G*
| | hartrees |
|---|
| Energy at 0K | -209.744795 |
| Energy at 298.15K | -209.754026 |
| HF Energy | -209.096749 |
| Nuclear repulsion energy | 134.287684 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3702 |
3525 |
54.19 |
|
|
|
| 2 |
A |
3583 |
3412 |
0.48 |
|
|
|
| 3 |
A |
3492 |
3325 |
0.25 |
|
|
|
| 4 |
A |
3122 |
2973 |
45.92 |
|
|
|
| 5 |
A |
3113 |
2965 |
27.59 |
|
|
|
| 6 |
A |
3024 |
2880 |
68.13 |
|
|
|
| 7 |
A |
2997 |
2854 |
65.15 |
|
|
|
| 8 |
A |
1723 |
1640 |
34.72 |
|
|
|
| 9 |
A |
1569 |
1494 |
0.03 |
|
|
|
| 10 |
A |
1550 |
1476 |
4.10 |
|
|
|
| 11 |
A |
1493 |
1421 |
84.50 |
|
|
|
| 12 |
A |
1449 |
1380 |
19.62 |
|
|
|
| 13 |
A |
1416 |
1348 |
3.66 |
|
|
|
| 14 |
A |
1363 |
1298 |
2.13 |
|
|
|
| 15 |
A |
1291 |
1229 |
24.54 |
|
|
|
| 16 |
A |
1218 |
1160 |
9.91 |
|
|
|
| 17 |
A |
1147 |
1092 |
45.00 |
|
|
|
| 18 |
A |
1095 |
1043 |
30.47 |
|
|
|
| 19 |
A |
1043 |
993 |
22.87 |
|
|
|
| 20 |
A |
984 |
937 |
80.05 |
|
|
|
| 21 |
A |
908 |
865 |
13.37 |
|
|
|
| 22 |
A |
869 |
828 |
67.83 |
|
|
|
| 23 |
A |
574 |
547 |
146.22 |
|
|
|
| 24 |
A |
541 |
515 |
27.25 |
|
|
|
| 25 |
A |
340 |
323 |
0.41 |
|
|
|
| 26 |
A |
277 |
264 |
15.79 |
|
|
|
| 27 |
A |
202 |
193 |
5.70 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22041.0 cm
-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 20989.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
-1.358 |
-0.567 |
0.122 |
| C2 |
-0.633 |
0.652 |
-0.279 |
| C3 |
0.784 |
0.557 |
0.276 |
| O4 |
1.412 |
-0.633 |
-0.172 |
| H5 |
-1.660 |
-0.484 |
1.094 |
| H6 |
-2.206 |
-0.678 |
-0.433 |
| H7 |
-1.114 |
1.590 |
0.046 |
| H8 |
-0.577 |
0.663 |
-1.375 |
| H9 |
1.394 |
1.401 |
-0.070 |
| H10 |
0.751 |
0.599 |
1.380 |
| H11 |
0.716 |
-1.313 |
-0.097 |
Atom - Atom Distances (Å)
| |
N1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| N1 | | 1.4737 | 2.4240 | 2.7858 | 1.0210 | 1.0192 | 2.1725 | 2.0886 | 3.3887 | 2.7181 | 2.2145 |
C2 | 1.4737 | | 1.5248 | 2.4170 | 2.0564 | 2.0646 | 1.1036 | 1.0969 | 2.1715 | 2.1612 | 2.3904 | C3 | 2.4240 | 1.5248 | | 1.4181 | 2.7795 | 3.3113 | 2.1729 | 2.1420 | 1.0968 | 1.1059 | 1.9089 | O4 | 2.7858 | 2.4170 | 1.4181 | | 3.3255 | 3.6269 | 3.3716 | 2.6609 | 2.0361 | 2.0893 | 0.9764 | H5 | 1.0210 | 2.0564 | 2.7795 | 3.3255 | | 1.6326 | 2.3869 | 2.9292 | 3.7726 | 2.6579 | 2.7837 | H6 | 1.0192 | 2.0646 | 3.3113 | 3.6269 | 1.6326 | | 2.5618 | 2.3098 | 4.1725 | 3.6952 | 3.0086 | H7 | 2.1725 | 1.1036 | 2.1729 | 3.3716 | 2.3869 | 2.5618 | | 1.7794 | 2.5180 | 2.4976 | 3.4350 | H8 | 2.0886 | 1.0969 | 2.1420 | 2.6609 | 2.9292 | 2.3098 | 1.7794 | | 2.4769 | 3.0589 | 2.6853 | H9 | 3.3887 | 2.1715 | 1.0968 | 2.0361 | 3.7726 | 4.1725 | 2.5180 | 2.4769 | | 1.7774 | 2.7979 | H10 | 2.7181 | 2.1612 | 1.1059 | 2.0893 | 2.6579 | 3.6952 | 2.4976 | 3.0589 | 1.7774 | | 2.4169 | H11 | 2.2145 | 2.3904 | 1.9089 | 0.9764 | 2.7837 | 3.0086 | 3.4350 | 2.6853 | 2.7979 | 2.4169 | |
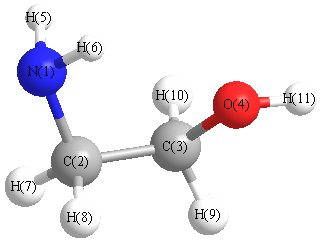 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C2 |
C3 |
107.870 |
|
N1 |
C2 |
H7 |
114.135 |
| N1 |
C2 |
H8 |
107.786 |
|
C2 |
N1 |
H5 |
109.700 |
| C2 |
N1 |
H6 |
110.508 |
|
C2 |
C3 |
O4 |
110.375 |
| C2 |
C3 |
H9 |
110.799 |
|
C2 |
C3 |
H10 |
109.449 |
| C3 |
C2 |
H7 |
110.499 |
|
C3 |
C2 |
H8 |
108.473 |
| C3 |
O4 |
H11 |
104.199 |
|
O4 |
C3 |
H9 |
107.433 |
| O4 |
C3 |
H10 |
111.141 |
|
H5 |
N1 |
H6 |
106.302 |
| H7 |
C2 |
H8 |
107.919 |
|
H9 |
C3 |
H10 |
107.597 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
