Jump to
S1C2
S1C3
Vibrational Frequencies calculated at B2PLYP/6-31G*
Geometric Data calculated at B2PLYP/6-31G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Vibrational Frequencies calculated at B2PLYP/6-31G*
Geometric Data calculated at B2PLYP/6-31G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at B2PLYP/6-31G*
| | hartrees |
|---|
| Energy at 0K | -210.142918 |
| Energy at 298.15K | -210.152137 |
| HF Energy | -209.944131 |
| Nuclear repulsion energy | 134.624435 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3667 |
3481 |
64.71 |
|
|
|
| 2 |
A |
3603 |
3420 |
1.07 |
|
|
|
| 3 |
A |
3509 |
3331 |
0.30 |
|
|
|
| 4 |
A |
3137 |
2977 |
46.04 |
|
|
|
| 5 |
A |
3127 |
2968 |
31.16 |
|
|
|
| 6 |
A |
3033 |
2879 |
76.13 |
|
|
|
| 7 |
A |
2998 |
2845 |
76.02 |
|
|
|
| 8 |
A |
1712 |
1625 |
31.88 |
|
|
|
| 9 |
A |
1565 |
1486 |
0.16 |
|
|
|
| 10 |
A |
1546 |
1467 |
4.32 |
|
|
|
| 11 |
A |
1486 |
1410 |
92.95 |
|
|
|
| 12 |
A |
1443 |
1369 |
15.46 |
|
|
|
| 13 |
A |
1409 |
1337 |
4.24 |
|
|
|
| 14 |
A |
1356 |
1287 |
2.48 |
|
|
|
| 15 |
A |
1284 |
1219 |
25.97 |
|
|
|
| 16 |
A |
1214 |
1153 |
10.67 |
|
|
|
| 17 |
A |
1132 |
1075 |
49.61 |
|
|
|
| 18 |
A |
1086 |
1031 |
31.27 |
|
|
|
| 19 |
A |
1033 |
980 |
13.80 |
|
|
|
| 20 |
A |
965 |
916 |
77.72 |
|
|
|
| 21 |
A |
899 |
853 |
12.43 |
|
|
|
| 22 |
A |
856 |
813 |
78.76 |
|
|
|
| 23 |
A |
595 |
565 |
147.06 |
|
|
|
| 24 |
A |
546 |
518 |
12.24 |
|
|
|
| 25 |
A |
335 |
318 |
0.47 |
|
|
|
| 26 |
A |
280 |
266 |
15.37 |
|
|
|
| 27 |
A |
197 |
187 |
6.53 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22005.0 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 20887.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
-1.350 |
-0.565 |
0.120 |
| C2 |
-0.633 |
0.658 |
-0.271 |
| C3 |
0.791 |
0.558 |
0.268 |
| O4 |
1.398 |
-0.642 |
-0.170 |
| H5 |
-1.645 |
-0.502 |
1.092 |
| H6 |
-2.193 |
-0.683 |
-0.433 |
| H7 |
-1.109 |
1.590 |
0.070 |
| H8 |
-0.592 |
0.683 |
-1.363 |
| H9 |
1.400 |
1.391 |
-0.093 |
| H10 |
0.769 |
0.617 |
1.369 |
| H11 |
0.683 |
-1.301 |
-0.103 |
Atom - Atom Distances (Å)
| |
N1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| N1 | | 1.4709 | 2.4222 | 2.7644 | 1.0176 | 1.0157 | 2.1691 | 2.0813 | 3.3814 | 2.7291 | 2.1732 |
C2 | 1.4709 | | 1.5262 | 2.4140 | 2.0562 | 2.0638 | 1.1004 | 1.0934 | 2.1684 | 2.1584 | 2.3667 | C3 | 2.4222 | 1.5262 | | 1.4135 | 2.7818 | 3.3073 | 2.1719 | 2.1420 | 1.0935 | 1.1035 | 1.8987 | O4 | 2.7644 | 2.4140 | 1.4135 | | 3.2975 | 3.6016 | 3.3653 | 2.6716 | 2.0342 | 2.0852 | 0.9757 | H5 | 1.0176 | 2.0562 | 2.7818 | 3.2975 | | 1.6308 | 2.3891 | 2.9225 | 3.7765 | 2.6754 | 2.7357 | H6 | 1.0157 | 2.0638 | 3.3073 | 3.6016 | 1.6308 | | 2.5679 | 2.3010 | 4.1627 | 3.7035 | 2.9605 | H7 | 2.1691 | 1.1004 | 2.1719 | 3.3653 | 2.3891 | 2.5679 | | 1.7737 | 2.5224 | 2.4827 | 3.4061 | H8 | 2.0813 | 1.0934 | 2.1420 | 2.6716 | 2.9225 | 2.3010 | 1.7737 | | 2.4659 | 3.0534 | 2.6740 | H9 | 3.3814 | 2.1684 | 1.0935 | 2.0342 | 3.7765 | 4.1627 | 2.5224 | 2.4659 | | 1.7712 | 2.7865 | H10 | 2.7291 | 2.1584 | 1.1035 | 2.0852 | 2.6754 | 3.7035 | 2.4827 | 3.0534 | 1.7712 | | 2.4193 | H11 | 2.1732 | 2.3667 | 1.8987 | 0.9757 | 2.7357 | 2.9605 | 3.4061 | 2.6740 | 2.7865 | 2.4193 | |
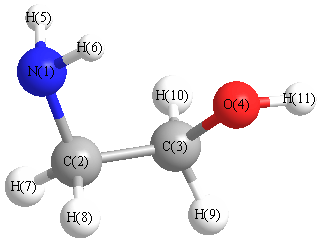 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C2 |
C3 |
107.818 |
|
N1 |
C2 |
H7 |
114.271 |
| N1 |
C2 |
H8 |
107.605 |
|
C2 |
N1 |
H5 |
110.098 |
| C2 |
N1 |
H6 |
110.863 |
|
C2 |
C3 |
O4 |
110.342 |
| C2 |
C3 |
H9 |
110.644 |
|
C2 |
C3 |
H10 |
109.274 |
| C3 |
C2 |
H7 |
110.510 |
|
C3 |
C2 |
H8 |
108.572 |
| C3 |
O4 |
H11 |
103.738 |
|
O4 |
C3 |
H9 |
107.790 |
| O4 |
C3 |
H10 |
111.289 |
|
H5 |
N1 |
H6 |
106.647 |
| H7 |
C2 |
H8 |
107.902 |
|
H9 |
C3 |
H10 |
107.458 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.769 |
|
|
|
| 2 |
C |
-0.180 |
|
|
|
| 3 |
C |
-0.007 |
|
|
|
| 4 |
O |
-0.655 |
|
|
|
| 5 |
H |
0.314 |
|
|
|
| 6 |
H |
0.322 |
|
|
|
| 7 |
H |
0.130 |
|
|
|
| 8 |
H |
0.162 |
|
|
|
| 9 |
H |
0.147 |
|
|
|
| 10 |
H |
0.115 |
|
|
|
| 11 |
H |
0.420 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.072 |
1.119 |
0.598 |
3.324 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.034 |
2.000 |
0.314 |
| y |
2.000 |
-25.409 |
-0.293 |
| z |
0.314 |
-0.293 |
-24.455 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.102 |
2.000 |
0.314 |
| y |
2.000 |
-0.165 |
-0.293 |
| z |
0.314 |
-0.293 |
1.267 |
|
| Polar |
|---|
| 3z2-r2 | 2.533 |
|---|
| x2-y2 | -0.625 |
|---|
| xy | 2.000 |
|---|
| xz | 0.314 |
|---|
| yz | -0.293 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.081 |
0.284 |
-0.024 |
| y |
0.284 |
4.861 |
0.051 |
| z |
-0.024 |
0.051 |
4.511 |
<r2> (average value of r
2) Å
2
| <r2> |
86.533 |
| (<r2>)1/2 |
9.302 |
