return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H8O2 (Ethyl acetate)

using model chemistry: CID/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CID/6-31G*
 hartrees
Energy at 0K-306.619564
Energy at 298.15K 
HF Energy-305.876012
Nuclear repulsion energy245.063738
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3311 3058 6.76      
2 A' 3262 3013 25.28      
3 A' 3208 2963 14.63      
4 A' 3194 2951 2.04      
5 A' 3182 2939 15.33      
6 A' 1955 1806 277.44      
7 A' 1625 1501 3.93      
8 A' 1599 1477 2.85      
9 A' 1572 1452 13.16      
10 A' 1539 1422 7.13      
11 A' 1514 1399 108.32      
12 A' 1486 1373 7.38      
13 A' 1394 1287 450.31      
14 A' 1218 1125 29.44      
15 A' 1169 1080 69.81      
16 A' 1083 1001 4.77      
17 A' 1014 937 4.97      
18 A' 928 858 10.48      
19 A' 677 625 11.36      
20 A' 450 416 0.96      
21 A' 390 360 11.02      
22 A' 205 189 5.77      
23 A" 3276 3026 43.16      
24 A" 3270 3020 4.95      
25 A" 3252 3004 2.70      
26 A" 1587 1465 5.54      
27 A" 1574 1454 8.15      
28 A" 1378 1272 0.78      
29 A" 1262 1166 5.57      
30 A" 1140 1053 8.77      
31 A" 860 794 0.17      
32 A" 642 593 9.87      
33 A" 282 261 1.12      
34 A" 159 146 5.21      
35 A" 90 83 0.11      
36 A" 74 69 0.69      

Unscaled Zero Point Vibrational Energy (zpe) 27409.0 cm-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 25317.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CID/6-31G*
ABC
0.28419 0.07064 0.05841

See section I.F.4 to change rotational constant units
Geometric Data calculated at CID/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -2.297 0.012 0.000
C2 -0.896 -0.524 0.000
O3 0.000 0.468 0.000
O4 -0.596 -1.687 0.000
C5 1.376 0.074 0.000
C6 2.200 1.338 0.000
H7 -2.999 -0.813 0.000
H8 -2.452 0.635 0.877
H9 -2.452 0.635 -0.877
H10 1.568 -0.537 0.878
H11 1.568 -0.537 -0.878
H12 3.260 1.090 0.000
H13 1.986 1.936 -0.882
H14 1.986 1.936 0.882

Atom - Atom Distances (Å)
  C1 C2 O3 O4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14
C11.50052.34182.40403.67304.68861.08321.08701.08704.00154.00155.66024.77754.7775
C21.50051.33681.20042.34863.61282.12282.12962.12962.61562.61564.45783.89063.8906
O32.34181.33682.23581.43092.36603.26092.60932.60932.05922.05923.31852.62252.6225
O42.40401.20042.23582.64324.11912.55673.09903.09902.60302.60304.75124.53554.5355
C53.67302.34861.43092.64321.50924.46333.96643.96641.08651.08652.14052.14912.1491
C64.68863.61282.36604.11911.50925.62644.78614.78612.16442.16441.08811.08701.0870
H71.08322.12283.26092.55674.46335.62641.77921.77924.65884.65886.54135.76075.7607
H81.08702.12962.60933.09903.96644.78611.77921.75484.18744.54035.79644.94814.6248
H91.08702.12962.60933.09903.96644.78611.77921.75484.54034.18745.79644.62484.9481
H104.00152.61562.05922.60301.08652.16444.65884.18744.54031.75512.50543.06382.5083
H114.00152.61562.05922.60301.08652.16444.65884.54034.18741.75512.50542.50833.0638
H125.66024.45783.31854.75122.14051.08816.54135.79645.79642.50542.50541.76541.7654
H134.77753.89062.62254.53552.14911.08705.76074.94814.62483.06382.50831.76541.7637
H144.77753.89062.62254.53552.14911.08705.76074.62484.94812.50833.06381.76541.7637

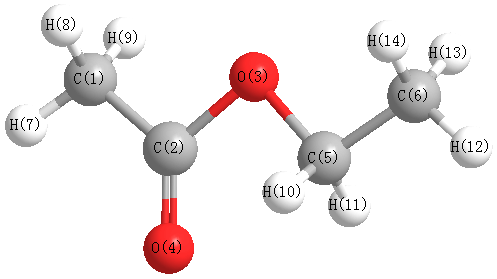
picture of Ethyl acetate state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O3 111.116 C1 C2 O4 125.401
C2 C1 H7 109.435 C2 C1 H8 109.744
C2 C1 H9 109.744 C2 O3 C5 116.069
O3 C2 O4 123.483 O3 C5 C6 107.136
O3 C5 H10 108.996 O3 C5 H11 108.996
C5 C6 H12 109.938 C5 C6 H13 110.688
C5 C6 H14 110.688 C6 C5 H10 111.959
C6 C5 H11 111.959 H7 C1 H8 110.130
H7 C1 H9 110.130 H8 C1 H9 107.636
H10 C5 H11 107.735 H12 C6 H13 108.510
H12 C6 H14 108.510 H13 C6 H14 108.441
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability