return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H8O2 (Ethyl acetate)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-306.793400
Energy at 298.15K 
HF Energy-305.872967
Nuclear repulsion energy243.070365
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3203 3050 7.46      
2 A' 3155 3005 24.05      
3 A' 3095 2947 15.25      
4 A' 3091 2943 2.01      
5 A' 3077 2930 15.31      
6 A' 1851 1763 215.56      
7 A' 1571 1496 3.83      
8 A' 1548 1474 2.99      
9 A' 1525 1452 11.97      
10 A' 1480 1410 4.38      
11 A' 1456 1387 75.54      
12 A' 1430 1362 5.65      
13 A' 1328 1265 402.42      
14 A' 1173 1117 15.21      
15 A' 1119 1065 83.05      
16 A' 1045 995 6.67      
17 A' 973 927 7.30      
18 A' 887 844 10.39      
19 A' 650 619 9.05      
20 A' 435 414 0.77      
21 A' 378 360 10.16      
22 A' 200 190 5.16      
23 A" 3167 3016 38.02      
24 A" 3164 3013 5.38      
25 A" 3138 2988 7.81      
26 A" 1537 1463 5.35      
27 A" 1524 1452 8.06      
28 A" 1323 1260 0.42      
29 A" 1217 1159 4.78      
30 A" 1095 1043 6.46      
31 A" 833 793 0.27      
32 A" 609 580 7.52      
33 A" 274 261 1.07      
34 A" 154 147 4.84      
35 A" 81 77 0.06      
36 A" 68 65 0.67      

Unscaled Zero Point Vibrational Energy (zpe) 26426.1 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 25165.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
0.27868 0.06980 0.05763

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -2.313 0.031 0.000
C2 -0.908 -0.519 0.000
O3 0.000 0.487 0.000
O4 -0.606 -1.696 0.000
C5 1.383 0.062 0.000
C6 2.228 1.320 0.000
H7 -3.024 -0.798 0.000
H8 -2.466 0.658 0.885
H9 -2.466 0.658 -0.885
H10 1.565 -0.558 0.885
H11 1.565 -0.558 -0.885
H12 3.293 1.055 0.000
H13 2.020 1.925 -0.889
H14 2.020 1.925 0.889

Atom - Atom Distances (Å)
  C1 C2 O3 O4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14
C11.50832.35732.42753.69584.72061.09201.09561.09564.02044.02045.69854.81154.8115
C21.50831.35471.21552.36343.63552.13452.14352.14352.62642.62644.48603.91593.9159
O32.35731.35472.26561.44672.37903.28572.62512.62512.07922.07923.34162.63422.6342
O42.42751.21552.26562.65514.13922.57913.12793.12792.60602.60604.77234.56094.5609
C53.69582.36341.44672.65511.51544.49023.99373.99371.09601.09602.15272.16022.1602
C64.72063.63552.37904.13921.51545.66344.82224.82222.17982.17981.09691.09561.0956
H71.09202.13453.28572.57914.49025.66341.79331.79334.67944.67946.58325.80065.8006
H81.09562.14352.62513.12793.99374.82221.79331.76944.20984.56675.83954.98694.6608
H91.09562.14352.62513.12793.99374.82221.79331.76944.56674.20985.83954.66084.9869
H104.02042.62642.07922.60601.09602.17984.67944.20984.56671.77002.52463.08582.5248
H114.02042.62642.07922.60601.09602.17984.67944.56674.20981.77002.52462.52483.0858
H125.69854.48603.34164.77232.15271.09696.58325.83955.83952.52462.52461.77991.7799
H134.81153.91592.63424.56092.16021.09565.80064.98694.66083.08582.52481.77991.7782
H144.81153.91592.63424.56092.16021.09565.80064.66084.98692.52483.08581.77991.7782

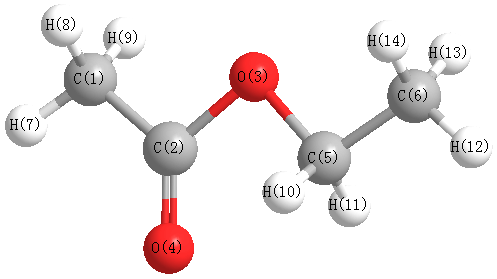
picture of Ethyl acetate state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O3 110.730 C1 C2 O4 125.716
C2 C1 H7 109.299 C2 C1 H8 109.793
C2 C1 H9 109.793 C2 O3 C5 115.016
O3 C2 O4 123.554 O3 C5 C6 106.844
O3 C5 H10 108.939 O3 C5 H11 108.939
C5 C6 H12 109.949 C5 C6 H13 110.627
C5 C6 H14 110.627 C6 C5 H10 112.172
C6 C5 H11 112.172 H7 C1 H8 110.117
H7 C1 H9 110.117 H8 C1 H9 107.703
H10 C5 H11 107.703 H12 C6 H13 108.546
H12 C6 H14 108.546 H13 C6 H14 108.486
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability