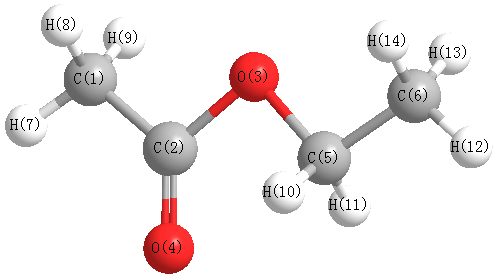
Vibrational Frequencies calculated at PBE1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3216 |
3057 |
5.22 |
64.31 |
0.68 |
0.81 |
| 2 |
A' |
3168 |
3010 |
19.79 |
73.69 |
0.71 |
0.83 |
| 3 |
A' |
3095 |
2941 |
5.97 |
55.33 |
0.00 |
0.00 |
| 4 |
A' |
3094 |
2940 |
11.66 |
140.07 |
0.06 |
0.11 |
| 5 |
A' |
3084 |
2931 |
14.49 |
109.66 |
0.01 |
0.01 |
| 6 |
A' |
1871 |
1778 |
239.07 |
3.21 |
0.34 |
0.51 |
| 7 |
A' |
1545 |
1468 |
6.23 |
1.01 |
0.69 |
0.81 |
| 8 |
A' |
1525 |
1449 |
2.25 |
23.16 |
0.75 |
0.86 |
| 9 |
A' |
1497 |
1423 |
13.78 |
14.66 |
0.73 |
0.84 |
| 10 |
A' |
1455 |
1383 |
7.51 |
5.50 |
0.74 |
0.85 |
| 11 |
A' |
1428 |
1357 |
92.29 |
3.46 |
0.74 |
0.85 |
| 12 |
A' |
1405 |
1335 |
11.60 |
3.28 |
0.62 |
0.77 |
| 13 |
A' |
1315 |
1250 |
400.95 |
0.39 |
0.22 |
0.36 |
| 14 |
A' |
1159 |
1102 |
21.85 |
6.28 |
0.16 |
0.28 |
| 15 |
A' |
1114 |
1058 |
75.45 |
2.80 |
0.75 |
0.86 |
| 16 |
A' |
1027 |
976 |
5.84 |
1.10 |
0.22 |
0.36 |
| 17 |
A' |
975 |
927 |
3.89 |
2.11 |
0.20 |
0.33 |
| 18 |
A' |
883 |
839 |
6.72 |
8.44 |
0.35 |
0.52 |
| 19 |
A' |
650 |
618 |
7.06 |
7.30 |
0.33 |
0.50 |
| 20 |
A' |
430 |
409 |
0.86 |
0.34 |
0.69 |
0.82 |
| 21 |
A' |
373 |
355 |
10.29 |
2.49 |
0.32 |
0.48 |
| 22 |
A' |
195 |
186 |
5.21 |
0.28 |
0.66 |
0.80 |
| 23 |
A" |
3174 |
3016 |
30.95 |
25.62 |
0.75 |
0.86 |
| 24 |
A" |
3169 |
3012 |
3.50 |
57.22 |
0.75 |
0.86 |
| 25 |
A" |
3138 |
2982 |
9.32 |
81.29 |
0.75 |
0.86 |
| 26 |
A" |
1514 |
1439 |
6.82 |
19.40 |
0.75 |
0.86 |
| 27 |
A" |
1501 |
1427 |
9.56 |
16.27 |
0.75 |
0.86 |
| 28 |
A" |
1311 |
1246 |
0.59 |
11.57 |
0.75 |
0.86 |
| 29 |
A" |
1195 |
1135 |
4.91 |
1.37 |
0.75 |
0.86 |
| 30 |
A" |
1076 |
1022 |
8.72 |
0.69 |
0.75 |
0.86 |
| 31 |
A" |
816 |
776 |
1.05 |
0.20 |
0.75 |
0.86 |
| 32 |
A" |
610 |
580 |
5.00 |
1.37 |
0.75 |
0.86 |
| 33 |
A" |
270 |
257 |
1.02 |
0.01 |
0.75 |
0.86 |
| 34 |
A" |
157 |
150 |
4.22 |
0.01 |
0.75 |
0.86 |
| 35 |
A" |
65 |
62 |
0.05 |
0.35 |
0.75 |
0.86 |
| 36 |
A" |
50 |
47 |
0.90 |
0.03 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 26274.6 cm
-1
Scaled (by 0.9503) Zero Point Vibrational Energy (zpe) 24968.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.609 |
|
|
|
| 2 |
C |
0.610 |
|
|
|
| 3 |
O |
-0.454 |
|
|
|
| 4 |
O |
-0.472 |
|
|
|
| 5 |
C |
-0.093 |
|
|
|
| 6 |
C |
-0.542 |
|
|
|
| 7 |
H |
0.211 |
|
|
|
| 8 |
H |
0.209 |
|
|
|
| 9 |
H |
0.209 |
|
|
|
| 10 |
H |
0.187 |
|
|
|
| 11 |
H |
0.187 |
|
|
|
| 12 |
H |
0.181 |
|
|
|
| 13 |
H |
0.188 |
|
|
|
| 14 |
H |
0.188 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.280 |
1.944 |
0.000 |
1.964 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-30.103 |
-1.015 |
0.000 |
| y |
-1.015 |
-41.930 |
0.000 |
| z |
0.000 |
0.000 |
-35.062 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
8.393 |
-1.015 |
0.000 |
| y |
-1.015 |
-9.348 |
0.000 |
| z |
0.000 |
0.000 |
0.955 |
|
| Polar |
|---|
| 3z2-r2 | 1.910 |
|---|
| x2-y2 | 11.827 |
|---|
| xy | -1.015 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.401 |
0.456 |
0.000 |
| y |
0.456 |
7.265 |
0.000 |
| z |
0.000 |
0.000 |
5.602 |
<r2> (average value of r
2) Å
2
| <r2> |
199.863 |
| (<r2>)1/2 |
14.137 |