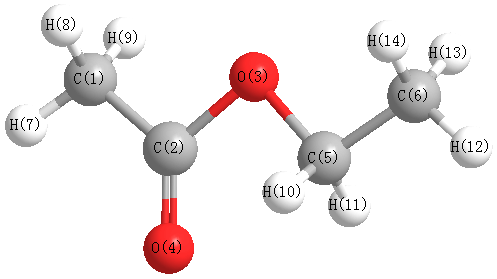
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3125 |
3073 |
6.25 |
67.84 |
0.67 |
0.80 |
| 2 |
A' |
3077 |
3026 |
20.41 |
80.41 |
0.70 |
0.83 |
| 3 |
A' |
3007 |
2957 |
3.11 |
110.81 |
0.01 |
0.01 |
| 4 |
A' |
2998 |
2949 |
17.69 |
70.44 |
0.02 |
0.04 |
| 5 |
A' |
2996 |
2947 |
15.23 |
142.25 |
0.04 |
0.08 |
| 6 |
A' |
1778 |
1749 |
200.48 |
3.19 |
0.26 |
0.41 |
| 7 |
A' |
1496 |
1471 |
5.60 |
1.61 |
0.67 |
0.80 |
| 8 |
A' |
1478 |
1453 |
2.57 |
24.18 |
0.75 |
0.86 |
| 9 |
A' |
1453 |
1429 |
13.27 |
15.75 |
0.71 |
0.83 |
| 10 |
A' |
1398 |
1375 |
6.03 |
9.32 |
0.66 |
0.80 |
| 11 |
A' |
1371 |
1348 |
44.41 |
3.50 |
0.66 |
0.80 |
| 12 |
A' |
1353 |
1331 |
9.70 |
5.12 |
0.66 |
0.79 |
| 13 |
A' |
1237 |
1216 |
366.62 |
0.57 |
0.04 |
0.08 |
| 14 |
A' |
1115 |
1097 |
11.99 |
5.57 |
0.18 |
0.31 |
| 15 |
A' |
1051 |
1033 |
101.67 |
2.52 |
0.68 |
0.81 |
| 16 |
A' |
987 |
971 |
10.65 |
1.01 |
0.21 |
0.34 |
| 17 |
A' |
932 |
916 |
6.27 |
2.00 |
0.16 |
0.28 |
| 18 |
A' |
840 |
826 |
6.56 |
9.42 |
0.34 |
0.50 |
| 19 |
A' |
623 |
613 |
4.44 |
8.01 |
0.33 |
0.50 |
| 20 |
A' |
416 |
409 |
0.72 |
0.46 |
0.65 |
0.79 |
| 21 |
A' |
359 |
353 |
9.15 |
3.13 |
0.32 |
0.49 |
| 22 |
A' |
188 |
185 |
4.72 |
0.31 |
0.65 |
0.79 |
| 23 |
A" |
3082 |
3031 |
29.91 |
32.49 |
0.75 |
0.86 |
| 24 |
A" |
3075 |
3025 |
4.72 |
60.62 |
0.75 |
0.86 |
| 25 |
A" |
3036 |
2986 |
12.76 |
77.07 |
0.75 |
0.86 |
| 26 |
A" |
1469 |
1445 |
6.32 |
20.64 |
0.75 |
0.86 |
| 27 |
A" |
1457 |
1433 |
9.11 |
17.30 |
0.75 |
0.86 |
| 28 |
A" |
1260 |
1240 |
0.31 |
12.05 |
0.75 |
0.86 |
| 29 |
A" |
1151 |
1132 |
4.14 |
1.97 |
0.75 |
0.86 |
| 30 |
A" |
1033 |
1016 |
6.95 |
0.66 |
0.75 |
0.86 |
| 31 |
A" |
791 |
778 |
1.41 |
0.20 |
0.75 |
0.86 |
| 32 |
A" |
582 |
573 |
3.08 |
1.37 |
0.75 |
0.86 |
| 33 |
A" |
264 |
259 |
0.93 |
0.01 |
0.75 |
0.86 |
| 34 |
A" |
154 |
152 |
3.76 |
0.02 |
0.75 |
0.86 |
| 35 |
A" |
54 |
53 |
0.19 |
0.36 |
0.75 |
0.86 |
| 36 |
A" |
20 |
20 |
0.83 |
0.07 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 25351.4 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 24933.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.570 |
|
|
|
| 2 |
C |
0.554 |
|
|
|
| 3 |
O |
-0.401 |
|
|
|
| 4 |
O |
-0.435 |
|
|
|
| 5 |
C |
-0.083 |
|
|
|
| 6 |
C |
-0.505 |
|
|
|
| 7 |
H |
0.194 |
|
|
|
| 8 |
H |
0.193 |
|
|
|
| 9 |
H |
0.193 |
|
|
|
| 10 |
H |
0.171 |
|
|
|
| 11 |
H |
0.171 |
|
|
|
| 12 |
H |
0.167 |
|
|
|
| 13 |
H |
0.175 |
|
|
|
| 14 |
H |
0.175 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.325 |
1.879 |
0.000 |
1.907 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-30.333 |
-0.980 |
0.000 |
| y |
-0.980 |
-41.688 |
0.000 |
| z |
0.000 |
0.000 |
-35.216 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
8.120 |
-0.980 |
0.000 |
| y |
-0.980 |
-8.914 |
0.000 |
| z |
0.000 |
0.000 |
0.794 |
|
| Polar |
|---|
| 3z2-r2 | 1.588 |
|---|
| x2-y2 | 11.356 |
|---|
| xy | -0.980 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.054 |
0.549 |
0.000 |
| y |
0.549 |
7.541 |
0.000 |
| z |
0.000 |
0.000 |
5.732 |
<r2> (average value of r
2) Å
2
| <r2> |
203.194 |
| (<r2>)1/2 |
14.255 |