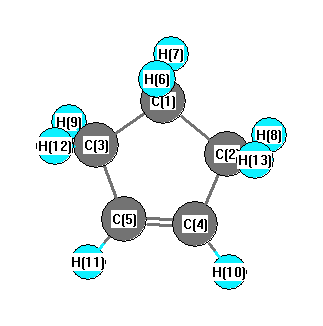
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3226 |
3086 |
28.00 |
|
|
|
| 2 |
A' |
3129 |
2993 |
40.42 |
|
|
|
| 3 |
A' |
3088 |
2954 |
55.24 |
|
|
|
| 4 |
A' |
3069 |
2936 |
8.27 |
|
|
|
| 5 |
A' |
3027 |
2896 |
29.14 |
|
|
|
| 6 |
A' |
1710 |
1636 |
1.53 |
|
|
|
| 7 |
A' |
1532 |
1466 |
1.18 |
|
|
|
| 8 |
A' |
1505 |
1440 |
1.16 |
|
|
|
| 9 |
A' |
1348 |
1289 |
0.45 |
|
|
|
| 10 |
A' |
1242 |
1188 |
1.29 |
|
|
|
| 11 |
A' |
1138 |
1089 |
0.13 |
|
|
|
| 12 |
A' |
1077 |
1031 |
5.16 |
|
|
|
| 13 |
A' |
992 |
949 |
0.84 |
|
|
|
| 14 |
A' |
922 |
882 |
0.46 |
|
|
|
| 15 |
A' |
830 |
794 |
0.44 |
|
|
|
| 16 |
A' |
718 |
687 |
29.91 |
|
|
|
| 17 |
A' |
612 |
585 |
12.11 |
|
|
|
| 18 |
A' |
144 |
137 |
0.22 |
|
|
|
| 19 |
A" |
3201 |
3062 |
8.92 |
|
|
|
| 20 |
A" |
3079 |
2946 |
29.77 |
|
|
|
| 21 |
A" |
3028 |
2897 |
60.13 |
|
|
|
| 22 |
A" |
1512 |
1447 |
0.80 |
|
|
|
| 23 |
A" |
1397 |
1337 |
1.26 |
|
|
|
| 24 |
A" |
1337 |
1279 |
0.49 |
|
|
|
| 25 |
A" |
1317 |
1260 |
1.92 |
|
|
|
| 26 |
A" |
1242 |
1188 |
0.01 |
|
|
|
| 27 |
A" |
1161 |
1111 |
1.47 |
|
|
|
| 28 |
A" |
1057 |
1012 |
1.73 |
|
|
|
| 29 |
A" |
971 |
929 |
0.12 |
|
|
|
| 30 |
A" |
934 |
893 |
8.20 |
|
|
|
| 31 |
A" |
905 |
866 |
1.80 |
|
|
|
| 32 |
A" |
782 |
748 |
0.22 |
|
|
|
| 33 |
A" |
390 |
374 |
0.08 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25810.0 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 24692.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.335 |
|
|
-0.146 |
| 2 |
C |
-0.354 |
|
|
0.118 |
| 3 |
C |
-0.354 |
|
|
0.118 |
| 4 |
C |
-0.134 |
|
|
-0.221 |
| 5 |
C |
-0.134 |
|
|
-0.221 |
| 6 |
H |
0.171 |
|
|
0.054 |
| 7 |
H |
0.162 |
|
|
0.027 |
| 8 |
H |
0.169 |
|
|
0.013 |
| 9 |
H |
0.169 |
|
|
0.013 |
| 10 |
H |
0.153 |
|
|
0.124 |
| 11 |
H |
0.153 |
|
|
0.124 |
| 12 |
H |
0.166 |
|
|
-0.001 |
| 13 |
H |
0.166 |
|
|
-0.001 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.047 |
-0.157 |
0.000 |
0.164 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.071 |
-0.179 |
0.000 |
0.193 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.988 |
-0.038 |
0.000 |
| y |
-0.038 |
-30.552 |
0.000 |
| z |
0.000 |
0.000 |
-30.008 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.708 |
-0.038 |
0.000 |
| y |
-0.038 |
0.446 |
0.000 |
| z |
0.000 |
0.000 |
1.262 |
|
| Polar |
|---|
| 3z2-r2 | 2.523 |
|---|
| x2-y2 | -1.436 |
|---|
| xy | -0.038 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
104.655 |
| (<r2>)1/2 |
10.230 |