return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8 (Cyclopentene)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-195.077389
Energy at 298.15K-195.086367
HF Energy-194.864905
Nuclear repulsion energy171.188930
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3245 3080 26.51      
2 A' 3149 2989 43.77      
3 A' 3111 2953 51.97      
4 A' 3092 2935 10.19      
5 A' 3054 2898 29.41      
6 A' 1700 1613 1.23      
7 A' 1558 1479 0.60      
8 A' 1533 1455 1.05      
9 A' 1364 1295 0.12      
10 A' 1260 1196 1.29      
11 A' 1151 1092 0.09      
12 A' 1097 1041 5.13      
13 A' 994 943 0.82      
14 A' 925 878 0.40      
15 A' 843 801 0.38      
16 A' 723 686 27.89      
17 A' 614 583 10.04      
18 A' 156 148 0.18      
19 A" 3221 3057 8.17      
20 A" 3105 2947 28.60      
21 A" 3054 2899 55.70      
22 A" 1540 1462 0.29      
23 A" 1412 1340 0.97      
24 A" 1357 1288 0.84      
25 A" 1337 1269 1.66      
26 A" 1251 1187 0.03      
27 A" 1174 1115 1.54      
28 A" 1061 1007 1.34      
29 A" 968 919 0.08      
30 A" 940 892 5.44      
31 A" 914 867 2.33      
32 A" 786 746 0.17      
33 A" 393 373 0.07      

Unscaled Zero Point Vibrational Energy (zpe) 26038.0 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 24715.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.24393 0.24105 0.13170

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.267 -1.202 0.000
C2 -0.069 -0.328 1.236
C3 -0.069 -0.328 -1.236
C4 -0.069 1.073 0.669
C5 -0.069 1.073 -0.669
H6 1.336 -1.431 0.000
H7 -0.270 -2.154 0.000
H8 -1.054 -0.574 1.655
H9 -1.054 -0.574 -1.655
H10 -0.107 1.959 1.295
H11 -0.107 1.959 -1.295
H12 0.656 -0.459 -2.047
H13 0.656 -0.459 2.047

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13
C11.55041.55042.39472.39471.09391.09292.20862.20863.43643.43642.21222.2122
C21.55042.47261.51212.36512.17232.21401.09823.06462.28873.41233.36511.0954
C31.55042.47262.36511.51212.17232.21403.06461.09823.41232.28871.09543.3651
C42.39471.51212.36511.33752.94823.30182.15803.01411.08592.15513.20152.1848
C52.39472.36511.51211.33752.94823.30183.01412.15802.15511.08592.18483.2015
H61.09392.17232.17232.94822.94821.76133.03113.03113.90553.90552.36622.3662
H71.09292.21402.21403.30183.30181.76132.41892.41894.31554.31552.81422.8142
H82.20861.09823.06462.15803.01413.03112.41893.31052.72854.00274.07981.7578
H92.20863.06461.09823.01412.15803.03112.41893.31054.00272.72851.75784.0798
H103.43642.28873.41231.08592.15513.90554.31552.72854.00272.59074.19562.6449
H113.43643.41232.28872.15511.08593.90554.31554.00272.72852.59072.64494.1956
H122.21223.36511.09543.20152.18482.36622.81424.07981.75784.19562.64494.0946
H132.21221.09543.36512.18483.20152.36622.81421.75784.07982.64494.19564.0946

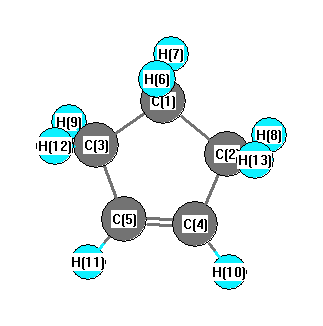
picture of Cyclopentene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 102.873 C1 C2 H8 111.859
C1 C2 H13 112.327 C1 C3 C5 102.873
C1 C3 H9 111.859 C1 C3 H12 112.327
C2 C1 C3 105.764 C2 C1 H6 109.248
C2 C1 H7 112.616 C2 C4 C5 112.046
C2 C4 H10 122.665 C3 C1 H6 109.248
C3 C1 H7 112.616 C3 C5 C4 112.046
C3 C5 H11 122.665 C4 C2 H8 110.527
C4 C2 H13 112.863 C4 C5 H11 125.242
C5 C3 H9 110.527 C5 C3 H12 112.863
C5 C4 H10 125.242 H6 C1 H7 107.299
H8 C2 H13 106.508 H9 C3 H12 106.508
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.302      
2 C -0.316      
3 C -0.316      
4 C -0.138      
5 C -0.138      
6 H 0.155      
7 H 0.148      
8 H 0.153      
9 H 0.153      
10 H 0.149      
11 H 0.149      
12 H 0.151      
13 H 0.151      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 5.325 -0.067 0.000
y -0.067 7.310 0.000
z 0.000 0.000 8.656


<r2> (average value of r2) Å2
<r2> 104.944
(<r2>)1/2 10.244