return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H12O2 (Hexanoic acid)

using model chemistry: CID/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A1
Energy calculated at CID/6-31G*
 hartrees
Energy at 0K-384.878302
Energy at 298.15K-384.892390
HF Energy-383.950129
Nuclear repulsion energy371.784544
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3922 3623 93.76      
2 A 3252 3004 40.35      
3 A 3248 3001 97.26      
4 A 3244 2996 7.67      
5 A 3221 2975 0.79      
6 A 3208 2964 19.36      
7 A 3194 2951 33.70      
8 A 3186 2943 2.87      
9 A 3184 2941 4.54      
10 A 3173 2931 26.69      
11 A 3168 2927 36.15      
12 A 3153 2913 14.03      
13 A 1980 1829 306.61      
14 A 1620 1497 6.14      
15 A 1610 1487 3.49      
16 A 1607 1484 6.66      
17 A 1600 1478 0.72      
18 A 1599 1477 0.77      
19 A 1575 1455 18.63      
20 A 1551 1432 60.19      
21 A 1529 1412 2.61      
22 A 1517 1401 20.04      
23 A 1462 1351 37.97      
24 A 1422 1313 0.89      
25 A 1416 1308 0.07      
26 A 1404 1297 0.60      
27 A 1379 1273 0.05      
28 A 1350 1247 27.56      
29 A 1314 1213 0.09      
30 A 1262 1166 255.03      
31 A 1218 1125 2.21      
32 A 1207 1115 23.72      
33 A 1143 1055 2.56      
34 A 1133 1046 4.33      
35 A 1091 1008 8.18      
36 A 1040 961 0.92      
37 A 981 906 3.72      
38 A 932 861 1.02      
39 A 914 844 7.84      
40 A 813 751 5.32      
41 A 770 712 10.00      
42 A 699 646 115.10      
43 A 676 624 38.94      
44 A 553 510 25.54      
45 A 537 496 23.55      
46 A 428 395 1.36      
47 A 322 298 1.91      
48 A 264 243 0.00      
49 A 239 221 2.31      
50 A 147 135 0.02      
51 A 121 112 0.14      
52 A 103 95 0.94      
53 A 62 57 0.10      
54 A 45 42 0.31      

Unscaled Zero Point Vibrational Energy (zpe) 40891.9 cm-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 37771.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CID/6-31G*
ABC
0.25044 0.02186 0.02048

See section I.F.4 to change rotational constant units
Geometric Data calculated at CID/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 4.048 -0.220 0.000
C2 2.719 0.522 -0.000
C3 1.518 -0.416 0.000
C4 0.187 0.325 -0.000
C5 -0.999 -0.625 0.000
C6 -2.322 0.083 -0.000
O7 -3.346 -0.784 0.000
O8 -2.489 1.270 -0.000
H9 4.887 0.472 -0.000
H10 4.142 -0.855 0.879
H11 4.142 -0.855 -0.879
H12 2.664 1.174 -0.872
H13 2.664 1.174 0.872
H14 1.572 -1.069 0.873
H15 1.572 -1.070 -0.872
H16 0.125 0.976 -0.870
H17 0.125 0.977 0.869
H18 -0.980 -1.283 0.868
H19 -0.980 -1.284 -0.868
H20 -4.157 -0.263 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 O7 O8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18 H19 H20
C11.52232.53793.89925.06296.37767.41596.70441.08761.08851.08852.14892.14892.75942.75944.19284.19285.21205.21208.2051
C21.52231.52342.53903.89015.05996.20375.26072.16902.16642.16641.09031.09032.14672.14672.77322.77324.20604.20606.9200
C32.53791.52341.52292.52513.87254.87804.34683.48432.80172.80172.14542.14541.09161.09162.15302.15302.78292.78295.6768
C43.89922.53901.52291.51942.52103.70322.83794.70224.21934.21932.76012.76012.15022.15021.08841.08842.16842.16844.3836
C55.06293.89012.52511.51941.50152.35312.41105.98715.21995.21994.17324.17322.75082.75082.14102.14101.08961.08963.1790
C66.37765.05993.87252.52101.50151.34171.19837.21986.59066.59065.17855.17854.15424.15422.74622.74622.10352.10351.8670
O77.41596.20374.87803.70322.35311.34172.22558.32867.53977.53976.38136.38135.00345.00343.98773.98772.56992.56990.9632
O86.70445.26074.34682.83792.41101.19832.22557.41877.01787.01795.22725.22714.76714.76722.76992.76983.09053.09062.2657
H91.08762.16903.48434.70225.98717.21988.32867.41871.75791.75792.48852.48853.75873.75874.86714.86726.18526.18529.0737
H101.08852.16642.80174.21935.21996.59067.53977.01781.75791.75763.06022.51002.57853.11714.74844.41485.13945.42818.3659
H111.08852.16642.80174.21935.21996.59067.53977.01791.75791.75762.51003.06023.11712.57854.41484.74845.42815.13948.3659
H122.14891.09032.14542.76014.17325.17856.38135.22722.48853.06022.51001.74373.04482.49552.54733.08564.72734.39557.0253
H132.14891.09032.14542.76014.17325.17856.38135.22712.48852.51003.06021.74372.49553.04483.08562.54734.39554.72747.0253
H142.75942.14671.09162.15022.75084.15425.00344.76713.75872.57853.11713.04482.49551.74523.05222.50632.56083.09645.8509
H152.75942.14671.09162.15022.75084.15425.00344.76723.75873.11712.57852.49553.04481.74522.50633.05223.09642.56085.8509
H164.19282.77322.15301.08842.14102.74623.98772.76994.86714.74844.41482.54733.08563.05222.50631.73883.05732.51564.5415
H174.19282.77322.15301.08842.14102.74623.98772.76984.86724.41484.74843.08562.54732.50633.05221.73882.51563.05734.5414
H185.21204.20602.78292.16841.08962.10352.56993.09056.18525.13945.42814.72734.39552.56083.09643.05732.51561.73623.4480
H195.21204.20602.78292.16841.08962.10352.56993.09066.18525.42815.13944.39554.72743.09642.56082.51563.05731.73623.4481
H208.20516.92005.67684.38363.17901.86700.96322.26579.07378.36598.36597.02537.02535.85095.85094.54154.54143.44803.4481

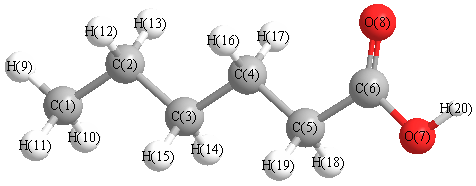
picture of Hexanoic acid state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 112.866 C1 C2 H12 109.561
C1 C2 H13 109.561 C2 C1 H9 111.329
C2 C1 H10 111.063 C2 C1 H11 111.063
C2 C3 C4 112.914 C2 C3 H14 109.233
C2 C3 H15 109.233 C3 C2 H12 109.217
C3 C2 H13 109.217 C3 C4 C5 112.198
C3 C4 H16 109.959 C3 C4 H17 109.959
C4 C3 H14 109.549 C4 C3 H15 109.549
C4 C5 C6 113.133 C4 C5 H18 111.365
C4 C5 H19 111.365 C5 C4 H16 109.253
C5 C4 H17 109.253 C5 C6 O7 111.586
C5 C6 O8 126.144 C6 C5 H18 107.478
C6 C5 H19 107.478 C6 O7 H20 107.045
O7 C6 O8 122.270 H9 C1 H10 107.770
H9 C1 H11 107.770 H10 C1 H11 107.680
H12 C2 H13 106.199 H14 C3 H15 106.135
H16 C4 H17 106.026 H18 C5 H19 105.632
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability