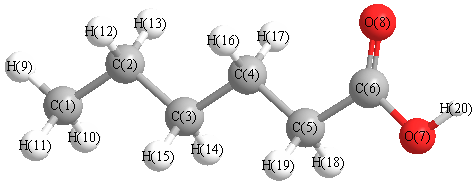
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3719 |
3558 |
48.29 |
|
|
|
| 2 |
A |
3131 |
2995 |
37.91 |
|
|
|
| 3 |
A |
3124 |
2989 |
70.07 |
|
|
|
| 4 |
A |
3115 |
2980 |
30.35 |
|
|
|
| 5 |
A |
3090 |
2956 |
1.49 |
|
|
|
| 6 |
A |
3081 |
2947 |
19.62 |
|
|
|
| 7 |
A |
3068 |
2935 |
38.24 |
|
|
|
| 8 |
A |
3056 |
2924 |
3.51 |
|
|
|
| 9 |
A |
3055 |
2923 |
4.65 |
|
|
|
| 10 |
A |
3051 |
2919 |
37.39 |
|
|
|
| 11 |
A |
3042 |
2910 |
30.80 |
|
|
|
| 12 |
A |
3025 |
2894 |
13.62 |
|
|
|
| 13 |
A |
1863 |
1783 |
251.76 |
|
|
|
| 14 |
A |
1535 |
1469 |
6.45 |
|
|
|
| 15 |
A |
1525 |
1459 |
7.45 |
|
|
|
| 16 |
A |
1525 |
1459 |
2.91 |
|
|
|
| 17 |
A |
1513 |
1448 |
0.54 |
|
|
|
| 18 |
A |
1510 |
1445 |
0.81 |
|
|
|
| 19 |
A |
1484 |
1420 |
15.44 |
|
|
|
| 20 |
A |
1446 |
1384 |
55.18 |
|
|
|
| 21 |
A |
1435 |
1373 |
7.44 |
|
|
|
| 22 |
A |
1424 |
1362 |
9.11 |
|
|
|
| 23 |
A |
1379 |
1319 |
31.13 |
|
|
|
| 24 |
A |
1348 |
1290 |
0.72 |
|
|
|
| 25 |
A |
1341 |
1283 |
0.10 |
|
|
|
| 26 |
A |
1325 |
1268 |
0.37 |
|
|
|
| 27 |
A |
1305 |
1248 |
0.08 |
|
|
|
| 28 |
A |
1272 |
1217 |
19.30 |
|
|
|
| 29 |
A |
1236 |
1183 |
0.02 |
|
|
|
| 30 |
A |
1181 |
1130 |
215.14 |
|
|
|
| 31 |
A |
1144 |
1095 |
1.44 |
|
|
|
| 32 |
A |
1138 |
1089 |
43.90 |
|
|
|
| 33 |
A |
1087 |
1039 |
3.59 |
|
|
|
| 34 |
A |
1079 |
1032 |
5.03 |
|
|
|
| 35 |
A |
1037 |
992 |
11.40 |
|
|
|
| 36 |
A |
983 |
941 |
0.42 |
|
|
|
| 37 |
A |
929 |
888 |
6.55 |
|
|
|
| 38 |
A |
882 |
844 |
2.91 |
|
|
|
| 39 |
A |
862 |
825 |
7.54 |
|
|
|
| 40 |
A |
770 |
736 |
4.52 |
|
|
|
| 41 |
A |
735 |
704 |
13.69 |
|
|
|
| 42 |
A |
681 |
652 |
92.98 |
|
|
|
| 43 |
A |
636 |
609 |
29.94 |
|
|
|
| 44 |
A |
526 |
503 |
21.01 |
|
|
|
| 45 |
A |
508 |
486 |
22.93 |
|
|
|
| 46 |
A |
403 |
386 |
1.11 |
|
|
|
| 47 |
A |
306 |
293 |
1.72 |
|
|
|
| 48 |
A |
247 |
237 |
0.01 |
|
|
|
| 49 |
A |
224 |
215 |
2.10 |
|
|
|
| 50 |
A |
140 |
134 |
0.01 |
|
|
|
| 51 |
A |
118 |
113 |
0.17 |
|
|
|
| 52 |
A |
97 |
93 |
0.94 |
|
|
|
| 53 |
A |
60 |
58 |
0.05 |
|
|
|
| 54 |
A |
38 |
36 |
0.26 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 38915.2 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 37230.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.509 |
|
|
|
| 2 |
C |
-0.306 |
|
|
|
| 3 |
C |
-0.305 |
|
|
|
| 4 |
C |
-0.306 |
|
|
|
| 5 |
C |
-0.417 |
|
|
|
| 6 |
C |
0.588 |
|
|
|
| 7 |
O |
-0.582 |
|
|
|
| 8 |
O |
-0.467 |
|
|
|
| 9 |
H |
0.168 |
|
|
|
| 10 |
H |
0.165 |
|
|
|
| 11 |
H |
0.165 |
|
|
|
| 12 |
H |
0.161 |
|
|
|
| 13 |
H |
0.161 |
|
|
|
| 14 |
H |
0.154 |
|
|
|
| 15 |
H |
0.154 |
|
|
|
| 16 |
H |
0.177 |
|
|
|
| 17 |
H |
0.177 |
|
|
|
| 18 |
H |
0.201 |
|
|
|
| 19 |
H |
0.201 |
|
|
|
| 20 |
H |
0.421 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.213 |
-1.375 |
0.000 |
1.391 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-44.876 |
2.681 |
-0.000 |
| y |
2.681 |
-54.183 |
0.001 |
| z |
-0.000 |
0.001 |
-48.254 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.343 |
2.681 |
-0.000 |
| y |
2.681 |
-7.618 |
0.001 |
| z |
-0.000 |
0.001 |
1.275 |
|
| Polar |
|---|
| 3z2-r2 | 2.550 |
|---|
| x2-y2 | 9.307 |
|---|
| xy | 2.681 |
|---|
| xz | -0.000 |
|---|
| yz | 0.001 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.716 |
0.044 |
0.000 |
| y |
0.044 |
9.856 |
-0.000 |
| z |
0.000 |
-0.000 |
8.158 |
<r2> (average value of r
2) Å
2
| <r2> |
510.095 |
| (<r2>)1/2 |
22.585 |