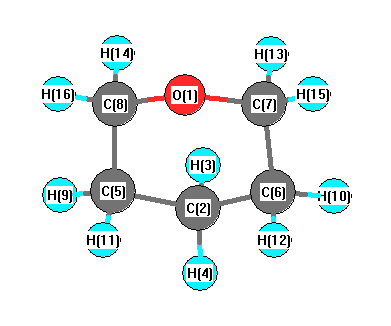
Vibrational Frequencies calculated at PBE1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3133 |
2977 |
42.90 |
|
|
|
| 2 |
A' |
3123 |
2968 |
50.06 |
|
|
|
| 3 |
A' |
3111 |
2956 |
35.26 |
|
|
|
| 4 |
A' |
3069 |
2916 |
14.84 |
|
|
|
| 5 |
A' |
3053 |
2901 |
25.73 |
|
|
|
| 6 |
A' |
2991 |
2842 |
101.39 |
|
|
|
| 7 |
A' |
1532 |
1456 |
3.71 |
|
|
|
| 8 |
A' |
1517 |
1442 |
6.83 |
|
|
|
| 9 |
A' |
1506 |
1432 |
7.93 |
|
|
|
| 10 |
A' |
1444 |
1372 |
9.30 |
|
|
|
| 11 |
A' |
1402 |
1332 |
0.89 |
|
|
|
| 12 |
A' |
1348 |
1281 |
4.05 |
|
|
|
| 13 |
A' |
1303 |
1238 |
5.91 |
|
|
|
| 14 |
A' |
1192 |
1132 |
3.64 |
|
|
|
| 15 |
A' |
1075 |
1022 |
9.71 |
|
|
|
| 16 |
A' |
1037 |
986 |
9.79 |
|
|
|
| 17 |
A' |
918 |
873 |
19.41 |
|
|
|
| 18 |
A' |
871 |
828 |
5.63 |
|
|
|
| 19 |
A' |
850 |
808 |
1.62 |
|
|
|
| 20 |
A' |
564 |
536 |
3.78 |
|
|
|
| 21 |
A' |
438 |
416 |
1.51 |
|
|
|
| 22 |
A' |
405 |
384 |
4.40 |
|
|
|
| 23 |
A' |
255 |
242 |
3.55 |
|
|
|
| 24 |
A" |
3130 |
2974 |
43.89 |
|
|
|
| 25 |
A" |
3118 |
2963 |
37.75 |
|
|
|
| 26 |
A" |
3069 |
2916 |
28.16 |
|
|
|
| 27 |
A" |
2984 |
2836 |
20.48 |
|
|
|
| 28 |
A" |
1516 |
1441 |
0.62 |
|
|
|
| 29 |
A" |
1496 |
1422 |
2.47 |
|
|
|
| 30 |
A" |
1413 |
1342 |
10.02 |
|
|
|
| 31 |
A" |
1402 |
1332 |
0.97 |
|
|
|
| 32 |
A" |
1367 |
1299 |
0.04 |
|
|
|
| 33 |
A" |
1319 |
1254 |
10.14 |
|
|
|
| 34 |
A" |
1256 |
1194 |
54.75 |
|
|
|
| 35 |
A" |
1208 |
1148 |
0.01 |
|
|
|
| 36 |
A" |
1164 |
1106 |
64.42 |
|
|
|
| 37 |
A" |
1083 |
1029 |
9.34 |
|
|
|
| 38 |
A" |
994 |
945 |
3.15 |
|
|
|
| 39 |
A" |
908 |
863 |
0.41 |
|
|
|
| 40 |
A" |
829 |
788 |
0.22 |
|
|
|
| 41 |
A" |
465 |
442 |
0.01 |
|
|
|
| 42 |
A" |
240 |
228 |
0.52 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32547.5 cm
-1
Scaled (by 0.9503) Zero Point Vibrational Energy (zpe) 30929.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.465 |
|
|
|
| 2 |
C |
-0.335 |
|
|
|
| 3 |
H |
0.161 |
|
|
|
| 4 |
H |
0.172 |
|
|
|
| 5 |
C |
-0.341 |
|
|
|
| 6 |
C |
-0.341 |
|
|
|
| 7 |
C |
-0.082 |
|
|
|
| 8 |
C |
-0.082 |
|
|
|
| 9 |
H |
0.166 |
|
|
|
| 10 |
H |
0.166 |
|
|
|
| 11 |
H |
0.174 |
|
|
|
| 12 |
H |
0.174 |
|
|
|
| 13 |
H |
0.141 |
|
|
|
| 14 |
H |
0.141 |
|
|
|
| 15 |
H |
0.175 |
|
|
|
| 16 |
H |
0.175 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.948 |
1.046 |
0.000 |
1.412 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.603 |
-1.579 |
0.000 |
| y |
-1.579 |
-39.854 |
0.000 |
| z |
0.000 |
0.000 |
-35.071 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.140 |
-1.579 |
0.000 |
| y |
-1.579 |
-3.017 |
0.000 |
| z |
0.000 |
0.000 |
4.157 |
|
| Polar |
|---|
| 3z2-r2 | 8.314 |
|---|
| x2-y2 | 1.252 |
|---|
| xy | -1.579 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.366 |
0.068 |
0.000 |
| y |
0.068 |
7.853 |
0.000 |
| z |
0.000 |
0.000 |
8.774 |
<r2> (average value of r
2) Å
2
| <r2> |
151.412 |
| (<r2>)1/2 |
12.305 |