return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10O (2H-Pyran, tetrahydro-)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A'
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-271.446076
Energy at 298.15K-271.459004
HF Energy-271.173483
Nuclear repulsion energy259.866345
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3142 2982 42.73      
2 A' 3126 2967 64.36      
3 A' 3113 2954 35.75      
4 A' 3075 2919 11.27      
5 A' 3060 2904 26.72      
6 A' 3008 2856 98.45      
7 A' 1556 1477 3.20      
8 A' 1541 1463 3.79      
9 A' 1532 1455 5.71      
10 A' 1455 1381 10.41      
11 A' 1417 1345 0.25      
12 A' 1352 1283 3.18      
13 A' 1313 1246 5.89      
14 A' 1207 1145 3.40      
15 A' 1071 1017 6.41      
16 A' 1038 986 10.57      
17 A' 902 856 19.19      
18 A' 882 837 5.23      
19 A' 841 798 2.76      
20 A' 572 543 3.85      
21 A' 440 418 1.65      
22 A' 408 387 4.75      
23 A' 255 242 3.84      
24 A" 3139 2980 47.34      
25 A" 3119 2961 32.05      
26 A" 3074 2918 35.44      
27 A" 3002 2849 19.10      
28 A" 1541 1463 0.09      
29 A" 1523 1445 1.58      
30 A" 1425 1353 12.74      
31 A" 1415 1343 0.83      
32 A" 1385 1314 0.18      
33 A" 1328 1261 8.14      
34 A" 1252 1189 35.75      
35 A" 1217 1155 0.15      
36 A" 1143 1085 74.83      
37 A" 1084 1029 14.15      
38 A" 1005 954 3.54      
39 A" 903 857 1.10      
40 A" 838 795 0.74      
41 A" 467 443 0.01      
42 A" 243 230 0.50      

Unscaled Zero Point Vibrational Energy (zpe) 32704.1 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 31042.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.15571 0.15005 0.08666

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
O1 -0.680 -1.257 0.000
C2 0.653 1.314 0.000
H3 1.708 1.015 0.000
H4 0.620 2.408 0.000
C5 -0.023 0.736 1.252
C6 -0.023 0.736 -1.252
C7 -0.023 -0.796 -1.174
C8 -0.023 -0.796 1.174
H9 0.477 1.063 2.169
H10 0.477 1.063 -2.169
H11 -1.064 1.073 1.293
H12 -1.064 1.073 -1.293
H13 1.013 -1.165 -1.188
H14 1.013 -1.165 1.188
H15 -0.555 -1.235 -2.022
H16 -0.555 -1.235 2.022

Atom - Atom Distances (Å)
  O1 C2 H3 H4 C5 C6 C7 C8 H9 H10 H11 H12 H13 H14 H15 H16
O12.89663.29623.88912.44342.44341.42251.42253.38063.38062.69182.69182.07002.07002.02562.0256
C22.89661.09581.09441.53621.53622.50802.50802.19042.19042.16272.16272.77222.77223.47083.4708
H33.29621.09581.76702.15432.15432.76692.76692.49392.49393.05843.05842.57802.57803.77783.7778
H43.88911.09441.76702.18602.18603.47263.47262.55592.55592.50792.50793.78553.78554.32914.3291
C52.44341.53622.15432.18602.50392.86931.53351.09483.47271.09442.76973.26142.16523.85782.1814
C62.44341.53622.15432.18602.50391.53352.86933.47271.09482.76971.09442.16523.26142.18143.8578
C71.42252.50802.76693.47262.86931.53352.34903.85822.16713.26502.14191.09972.60571.09263.2697
C81.42252.50802.76693.47261.53352.86932.34902.16713.85822.14193.26502.60571.09973.26971.0926
H93.38062.19042.49392.55591.09483.47273.85822.16714.33771.77263.78904.06432.49274.88972.5240
H103.38062.19042.49392.55593.47271.09482.16713.85824.33773.78901.77262.49274.06432.52404.8897
H112.69182.16273.05842.50791.09442.76973.26502.14191.77263.78902.58533.93313.05414.07032.4727
H122.69182.16273.05842.50792.76971.09442.14193.26503.78901.77262.58533.05413.93312.47274.0703
H132.07002.77222.57803.78553.26142.16521.09972.60574.06432.49273.93313.05412.37571.77743.5728
H142.07002.77222.57803.78552.16523.26142.60571.09972.49274.06433.05413.93312.37573.57281.7774
H152.02563.47083.77784.32913.85782.18141.09263.26974.88972.52404.07032.47271.77743.57284.0433
H162.02563.47083.77784.32912.18143.85783.26971.09262.52404.88972.47274.07033.57281.77744.0433

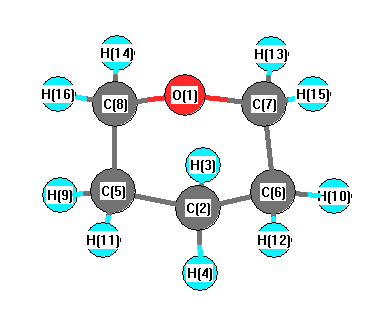
picture of 2H-Pyran, tetrahydro- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
O1 C7 C6 111.444 O1 C7 H13 109.647
O1 C7 H15 106.557 O1 C8 C5 111.444
O1 C8 H14 109.647 O1 C8 H16 106.557
C2 C5 C8 109.576 C2 C5 H9 111.622
C2 C5 H11 109.449 C2 C6 C7 109.576
C2 C6 H10 111.622 C2 C6 H12 109.449
H3 C2 H4 107.570 H3 C2 C5 108.716
H3 C2 C6 108.716 H4 C2 C5 111.294
H4 C2 C6 111.294 C5 C2 C6 109.172
C5 C8 H14 109.527 C5 C8 H16 111.231
C6 C7 H13 109.527 C6 C7 H15 111.231
C7 O1 C8 111.306 C7 C6 H10 109.962
C7 C6 H12 108.019 C8 C5 H9 109.962
C8 C5 H11 108.019 H9 C5 H11 108.132
H10 C6 H12 108.132 H13 C7 H15 108.347
H14 C8 H16 108.347
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 O -0.537      
2 C -0.295      
3 H 0.141      
4 H 0.152      
5 C -0.298      
6 C -0.298      
7 C -0.015      
8 C -0.015      
9 H 0.145      
10 H 0.145      
11 H 0.154      
12 H 0.154      
13 H 0.126      
14 H 0.126      
15 H 0.158      
16 H 0.158      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 7.303 0.081 0.000
y 0.081 7.762 0.000
z 0.000 0.000 8.723


<r2> (average value of r2) Å2
<r2> 152.509
(<r2>)1/2 12.349