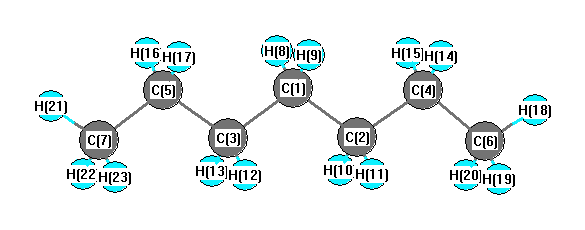
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3056 |
3005 |
44.43 |
|
|
|
| 2 |
A1 |
2978 |
2928 |
33.19 |
|
|
|
| 3 |
A1 |
2963 |
2915 |
131.10 |
|
|
|
| 4 |
A1 |
2951 |
2903 |
14.13 |
|
|
|
| 5 |
A1 |
2943 |
2895 |
0.25 |
|
|
|
| 6 |
A1 |
1495 |
1470 |
6.80 |
|
|
|
| 7 |
A1 |
1484 |
1459 |
0.19 |
|
|
|
| 8 |
A1 |
1468 |
1444 |
0.00 |
|
|
|
| 9 |
A1 |
1463 |
1439 |
0.00 |
|
|
|
| 10 |
A1 |
1389 |
1366 |
2.56 |
|
|
|
| 11 |
A1 |
1369 |
1347 |
0.23 |
|
|
|
| 12 |
A1 |
1285 |
1264 |
0.01 |
|
|
|
| 13 |
A1 |
1136 |
1118 |
0.95 |
|
|
|
| 14 |
A1 |
1060 |
1043 |
0.68 |
|
|
|
| 15 |
A1 |
991 |
975 |
0.00 |
|
|
|
| 16 |
A1 |
903 |
888 |
1.70 |
|
|
|
| 17 |
A1 |
405 |
398 |
0.08 |
|
|
|
| 18 |
A1 |
297 |
292 |
0.01 |
|
|
|
| 19 |
A1 |
96 |
94 |
0.01 |
|
|
|
| 20 |
A2 |
3049 |
2999 |
0.00 |
|
|
|
| 21 |
A2 |
3002 |
2952 |
0.00 |
|
|
|
| 22 |
A2 |
2976 |
2926 |
0.00 |
|
|
|
| 23 |
A2 |
1482 |
1457 |
0.00 |
|
|
|
| 24 |
A2 |
1303 |
1282 |
0.00 |
|
|
|
| 25 |
A2 |
1297 |
1276 |
0.00 |
|
|
|
| 26 |
A2 |
1215 |
1195 |
0.00 |
|
|
|
| 27 |
A2 |
1005 |
988 |
0.00 |
|
|
|
| 28 |
A2 |
827 |
813 |
0.00 |
|
|
|
| 29 |
A2 |
724 |
712 |
0.00 |
|
|
|
| 30 |
A2 |
246 |
241 |
0.00 |
|
|
|
| 31 |
A2 |
139 |
137 |
0.00 |
|
|
|
| 32 |
A2 |
76 |
74 |
0.00 |
|
|
|
| 33 |
B1 |
3049 |
2999 |
108.17 |
|
|
|
| 34 |
B1 |
3011 |
2961 |
115.87 |
|
|
|
| 35 |
B1 |
2989 |
2939 |
0.00 |
|
|
|
| 36 |
B1 |
2968 |
2919 |
0.08 |
|
|
|
| 37 |
B1 |
1482 |
1458 |
13.81 |
|
|
|
| 38 |
B1 |
1310 |
1288 |
0.78 |
|
|
|
| 39 |
B1 |
1266 |
1245 |
0.07 |
|
|
|
| 40 |
B1 |
1179 |
1160 |
0.13 |
|
|
|
| 41 |
B1 |
918 |
902 |
1.10 |
|
|
|
| 42 |
B1 |
757 |
744 |
1.81 |
|
|
|
| 43 |
B1 |
719 |
708 |
6.91 |
|
|
|
| 44 |
B1 |
247 |
243 |
0.00 |
|
|
|
| 45 |
B1 |
145 |
143 |
0.00 |
|
|
|
| 46 |
B1 |
62 |
61 |
0.00 |
|
|
|
| 47 |
B2 |
3055 |
3005 |
31.66 |
|
|
|
| 48 |
B2 |
2977 |
2928 |
47.46 |
|
|
|
| 49 |
B2 |
2961 |
2912 |
0.05 |
|
|
|
| 50 |
B2 |
2944 |
2896 |
0.01 |
|
|
|
| 51 |
B2 |
1491 |
1467 |
1.22 |
|
|
|
| 52 |
B2 |
1475 |
1451 |
0.96 |
|
|
|
| 53 |
B2 |
1463 |
1439 |
0.29 |
|
|
|
| 54 |
B2 |
1390 |
1367 |
0.96 |
|
|
|
| 55 |
B2 |
1374 |
1351 |
0.93 |
|
|
|
| 56 |
B2 |
1337 |
1315 |
2.80 |
|
|
|
| 57 |
B2 |
1226 |
1206 |
4.54 |
|
|
|
| 58 |
B2 |
1071 |
1053 |
3.62 |
|
|
|
| 59 |
B2 |
1056 |
1039 |
1.58 |
|
|
|
| 60 |
B2 |
1035 |
1018 |
0.02 |
|
|
|
| 61 |
B2 |
872 |
858 |
2.75 |
|
|
|
| 62 |
B2 |
469 |
462 |
0.08 |
|
|
|
| 63 |
B2 |
237 |
233 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 46802.7 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 46030.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.281 |
|
|
|
| 2 |
C |
-0.274 |
|
|
|
| 3 |
C |
-0.274 |
|
|
|
| 4 |
C |
-0.279 |
|
|
|
| 5 |
C |
-0.279 |
|
|
|
| 6 |
C |
-0.482 |
|
|
|
| 7 |
C |
-0.482 |
|
|
|
| 8 |
H |
0.141 |
|
|
|
| 9 |
H |
0.141 |
|
|
|
| 10 |
H |
0.140 |
|
|
|
| 11 |
H |
0.140 |
|
|
|
| 12 |
H |
0.140 |
|
|
|
| 13 |
H |
0.140 |
|
|
|
| 14 |
H |
0.145 |
|
|
|
| 15 |
H |
0.145 |
|
|
|
| 16 |
H |
0.145 |
|
|
|
| 17 |
H |
0.145 |
|
|
|
| 18 |
H |
0.155 |
|
|
|
| 19 |
H |
0.155 |
|
|
|
| 20 |
H |
0.155 |
|
|
|
| 21 |
H |
0.155 |
|
|
|
| 22 |
H |
0.155 |
|
|
|
| 23 |
H |
0.155 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.052 |
0.052 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.160 |
0.000 |
0.000 |
| y |
0.000 |
-48.849 |
0.000 |
| z |
0.000 |
0.000 |
-48.280 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.404 |
0.000 |
0.000 |
| y |
0.000 |
-1.129 |
0.000 |
| z |
0.000 |
0.000 |
-0.275 |
|
| Polar |
|---|
| 3z2-r2 | -0.551 |
|---|
| x2-y2 | 1.689 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.167 |
0.000 |
0.000 |
| y |
0.000 |
14.442 |
0.000 |
| z |
0.000 |
0.000 |
10.505 |
<r2> (average value of r
2) Å
2
| <r2> |
480.353 |
| (<r2>)1/2 |
21.917 |