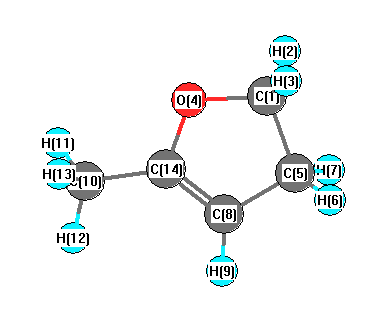
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3266 |
3125 |
4.59 |
|
|
|
| 2 |
A |
3165 |
3028 |
13.30 |
|
|
|
| 3 |
A |
3131 |
2995 |
40.44 |
|
|
|
| 4 |
A |
3120 |
2985 |
13.23 |
|
|
|
| 5 |
A |
3082 |
2949 |
43.99 |
|
|
|
| 6 |
A |
3059 |
2926 |
22.53 |
|
|
|
| 7 |
A |
3057 |
2924 |
42.19 |
|
|
|
| 8 |
A |
3028 |
2897 |
54.27 |
|
|
|
| 9 |
A |
1756 |
1680 |
55.69 |
|
|
|
| 10 |
A |
1545 |
1478 |
0.17 |
|
|
|
| 11 |
A |
1521 |
1455 |
0.24 |
|
|
|
| 12 |
A |
1513 |
1448 |
4.82 |
|
|
|
| 13 |
A |
1495 |
1431 |
8.07 |
|
|
|
| 14 |
A |
1442 |
1380 |
24.75 |
|
|
|
| 15 |
A |
1420 |
1359 |
6.23 |
|
|
|
| 16 |
A |
1339 |
1281 |
10.14 |
|
|
|
| 17 |
A |
1284 |
1228 |
57.30 |
|
|
|
| 18 |
A |
1267 |
1212 |
5.76 |
|
|
|
| 19 |
A |
1225 |
1172 |
28.68 |
|
|
|
| 20 |
A |
1218 |
1166 |
10.55 |
|
|
|
| 21 |
A |
1090 |
1043 |
0.74 |
|
|
|
| 22 |
A |
1072 |
1026 |
5.52 |
|
|
|
| 23 |
A |
1057 |
1012 |
48.79 |
|
|
|
| 24 |
A |
1038 |
993 |
5.50 |
|
|
|
| 25 |
A |
998 |
955 |
15.02 |
|
|
|
| 26 |
A |
950 |
909 |
17.64 |
|
|
|
| 27 |
A |
925 |
885 |
20.02 |
|
|
|
| 28 |
A |
862 |
824 |
2.88 |
|
|
|
| 29 |
A |
733 |
701 |
25.57 |
|
|
|
| 30 |
A |
704 |
674 |
2.05 |
|
|
|
| 31 |
A |
638 |
610 |
1.03 |
|
|
|
| 32 |
A |
558 |
534 |
1.91 |
|
|
|
| 33 |
A |
334 |
319 |
2.30 |
|
|
|
| 34 |
A |
225 |
215 |
6.82 |
|
|
|
| 35 |
A |
171 |
163 |
2.16 |
|
|
|
| 36 |
A |
101 |
96 |
1.52 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26694.9 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 25539.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.078 |
|
|
-0.019 |
| 2 |
H |
0.173 |
|
|
0.077 |
| 3 |
H |
0.169 |
|
|
0.070 |
| 4 |
O |
-0.502 |
|
|
-0.308 |
| 5 |
C |
-0.379 |
|
|
0.067 |
| 6 |
H |
0.173 |
|
|
0.024 |
| 7 |
H |
0.174 |
|
|
0.027 |
| 8 |
C |
-0.270 |
|
|
-0.419 |
| 9 |
H |
0.155 |
|
|
0.155 |
| 10 |
C |
-0.587 |
|
|
-0.717 |
| 11 |
H |
0.193 |
|
|
0.206 |
| 12 |
H |
0.185 |
|
|
0.198 |
| 13 |
H |
0.192 |
|
|
0.203 |
| 14 |
C |
0.402 |
|
|
0.436 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.391 |
0.661 |
0.118 |
0.777 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.389 |
0.625 |
0.114 |
0.745 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.715 |
1.223 |
-0.019 |
| y |
1.223 |
-37.032 |
-0.158 |
| z |
-0.019 |
-0.158 |
-36.786 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.194 |
1.223 |
-0.019 |
| y |
1.223 |
-2.781 |
-0.158 |
| z |
-0.019 |
-0.158 |
-2.413 |
|
| Polar |
|---|
| 3z2-r2 | -4.825 |
|---|
| x2-y2 | 5.317 |
|---|
| xy | 1.223 |
|---|
| xz | -0.019 |
|---|
| yz | -0.158 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
155.676 |
| (<r2>)1/2 |
12.477 |