return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8O (Furan, 2,3-dihydro-5-methyl-)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-270.235496
Energy at 298.15K-270.244668
HF Energy-269.964338
Nuclear repulsion energy237.752987
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3282 3115 4.28      
2 A 3182 3021 14.24      
3 A 3163 3002 38.61      
4 A 3146 2986 13.99      
5 A 3112 2954 40.31      
6 A 3084 2928 19.49      
7 A 3084 2927 43.25      
8 A 3058 2903 49.65      
9 A 1752 1663 46.31      
10 A 1568 1488 0.14      
11 A 1547 1468 0.12      
12 A 1540 1461 4.94      
13 A 1521 1444 6.88      
14 A 1462 1388 13.36      
15 A 1432 1359 8.98      
16 A 1355 1287 10.87      
17 A 1297 1231 40.94      
18 A 1276 1211 10.11      
19 A 1234 1171 14.52      
20 A 1225 1162 32.78      
21 A 1108 1052 0.47      
22 A 1087 1032 3.46      
23 A 1058 1005 26.42      
24 A 1041 988 21.29      
25 A 994 943 21.89      
26 A 954 905 16.32      
27 A 923 876 21.39      
28 A 873 829 6.40      
29 A 734 696 24.58      
30 A 699 664 1.49      
31 A 636 604 1.26      
32 A 553 525 2.67      
33 A 334 317 2.11      
34 A 226 215 5.89      
35 A 173 165 2.39      
36 A 124 118 1.81      

Unscaled Zero Point Vibrational Energy (zpe) 26917.8 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 25550.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.25152 0.10891 0.07971

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.322 -0.761 0.247
H2 -2.107 -1.444 -0.086
H3 -1.243 -0.798 1.338
O4 -0.014 -1.095 -0.259
C5 -1.495 0.711 -0.144
H6 -2.249 1.177 0.492
H7 -1.761 0.814 -1.205
C8 -0.100 1.132 0.058
H9 0.239 2.115 0.202
C10 2.159 0.014 0.056
H11 2.601 -0.376 -0.866
H12 2.543 1.020 0.229
H13 2.482 -0.632 0.878
C14 0.692 0.052 -0.036

Atom - Atom Distances (Å)
  C1 H2 H3 O4 C5 H6 H7 C8 H9 C10 H11 H12 H13 C14
C11.09211.09451.44121.53252.16222.18662.26123.27253.57154.09544.25543.85762.1904
H21.09211.78622.12802.24032.68722.54353.26844.27174.51034.88935.27124.75803.1739
H31.09451.78622.03672.12942.37193.05462.58263.45993.72544.45084.34383.75672.5210
O41.44122.12802.03672.33813.27422.75522.25183.25312.46062.77923.35422.78151.3658
C51.53252.24032.12942.33811.09121.09821.47102.25763.72564.29844.06674.31972.2868
H62.16222.68722.37193.27421.09121.80212.19282.67464.58035.27044.80195.07963.1932
H72.18662.54353.05462.75521.09821.80212.11012.76884.19524.53374.54074.94242.8223
C82.26123.26842.58262.25181.47102.19282.11011.04932.52153.22872.65103.23301.3433
H93.27254.27173.45993.25312.25762.67462.76881.04932.85093.59542.55163.61072.1258
C103.57154.51033.72542.46063.72564.58034.19522.52152.85091.09421.09051.09371.4705
H114.09544.88934.45082.77924.29845.27044.53373.22873.59541.09421.77511.76672.1247
H124.25545.27124.34383.35424.06674.80194.54072.65102.55161.09051.77511.77622.1050
H133.85764.75803.75672.78154.31975.07964.94243.23303.61071.09371.76671.77622.1223
C142.19043.17392.52101.36582.28683.19322.82231.34332.12581.47052.12472.10502.1223

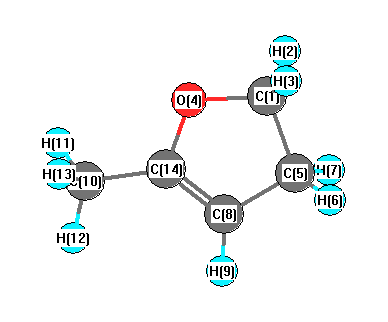
picture of Furan, 2,3-dihydro-5-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 O4 C14 102.552 C1 C5 H6 109.853
C1 C5 H7 111.367 C1 C5 C8 97.660
H2 C1 H3 109.548 H2 C1 O4 113.572
H2 C1 C5 116.198 H3 C1 O4 106.069
H3 C1 C5 107.127 O4 C1 C5 103.632
O4 C14 C8 112.440 O4 C14 C10 120.303
C5 C8 H9 126.396 C5 C8 C14 108.616
H6 C5 H7 110.793 H6 C5 C8 116.933
H7 C5 C8 109.578 C8 C14 C10 127.250
H9 C8 C14 124.917 H11 C10 H12 108.678
H11 C10 H13 107.705 H11 C10 C14 111.025
H12 C10 H13 108.818 H12 C10 C14 109.673
H13 C10 C14 110.873
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.019      
2 H 0.161      
3 H 0.155      
4 O -0.572      
5 C -0.339      
6 H 0.158      
7 H 0.160      
8 C -0.277      
9 H 0.153      
10 C -0.523      
11 H 0.179      
12 H 0.169      
13 H 0.178      
14 C 0.418      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 9.407 -0.506 0.017
y -0.506 8.095 0.019
z 0.017 0.019 5.652


<r2> (average value of r2) Å2
<r2> 156.220
(<r2>)1/2 12.499