return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H3F3 (Benzene trifluoride 123)

using model chemistry: CID/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at CID/6-31G*
 hartrees
Energy at 0K-528.240981
Energy at 298.15K-528.245134
HF Energy-527.240747
Nuclear repulsion energy435.718822
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3374 3117 0.38      
2 A1 3351 3095 3.77      
3 A1 1798 1661 0.09      
4 A1 1680 1552 220.53      
5 A1 1430 1321 54.13      
6 A1 1353 1249 31.83      
7 A1 1148 1060 9.77      
8 A1 883 816 13.79      
9 A1 744 688 24.99      
10 A1 510 471 0.07      
11 A1 319 294 2.11      
12 A2 942 870 0.00      
13 A2 612 565 0.00      
14 A2 272 251 0.00      
15 B1 1003 927 0.42      
16 B1 827 764 85.02      
17 B1 638 589 0.10      
18 B1 563 520 0.41      
19 B1 328 303 0.33      
20 B1 164 151 0.27      
21 B2 3369 3112 0.77      
22 B2 1797 1660 83.88      
23 B2 1627 1503 92.84      
24 B2 1361 1257 37.45      
25 B2 1339 1237 2.34      
26 B2 1240 1145 3.26      
27 B2 1111 1026 127.26      
28 B2 612 565 3.65      
29 B2 532 491 1.07      
30 B2 285 264 0.14      

Unscaled Zero Point Vibrational Energy (zpe) 17604.9 cm-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 16261.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CID/6-31G*
ABC
0.07829 0.05939 0.03377

See section I.F.4 to change rotational constant units
Geometric Data calculated at CID/6-31G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.716
C2 0.000 1.189 0.015
C3 0.000 -1.189 0.015
C4 0.000 1.204 -1.364
C5 0.000 -1.204 -1.364
C6 0.000 0.000 -2.048
F7 0.000 0.000 2.044
F8 0.000 2.328 0.703
F9 0.000 -2.328 0.703
H10 0.000 2.151 -1.876
H11 0.000 -2.151 -1.876
H12 0.000 0.000 -3.125

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 F7 F8 F9 H10 H11 H12
C11.38051.38052.40322.40322.76381.32762.32762.32763.36853.36853.8412
C21.38052.37861.37882.76232.38112.35151.32993.58352.12173.83873.3578
C31.38052.37862.76231.37882.38112.35153.58351.32993.83872.12173.3578
C42.40321.37882.76232.40861.38503.61382.35204.09201.07673.39452.1338
C52.40322.76231.37882.40861.38503.61384.09202.35203.39451.07672.1338
C62.76382.38112.38111.38501.38504.09133.60323.60322.15812.15811.0774
F71.32762.35152.35153.61383.61384.09132.68612.68614.47124.47125.1687
F82.32761.32993.58352.35204.09203.60322.68614.65512.58485.16824.4800
F92.32763.58351.32994.09202.35203.60322.68614.65515.16822.58484.4800
H103.36852.12173.83871.07673.39452.15814.47122.58485.16824.30252.4876
H113.36853.83872.12173.39451.07672.15814.47125.16822.58484.30252.4876
H123.84123.35783.35782.13382.13381.07745.16874.48004.48002.48762.4876

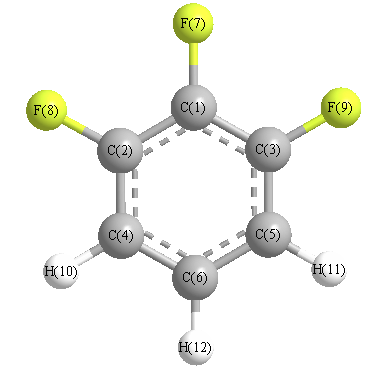
picture of Benzene trifluoride 123 state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 121.139 C1 C2 F8 118.343
C1 C3 C5 121.139 C1 C3 F9 118.343
C2 C1 C3 118.968 C2 C1 F7 120.516
C2 C4 C6 118.975 C2 C4 H10 119.041
C3 C1 F7 120.516 C3 C5 C6 118.975
C3 C5 H11 119.041 C4 C2 F8 120.518
C4 C6 C5 120.804 C4 C6 H12 119.598
C5 C3 F9 120.518 C5 C6 H12 119.598
C6 C4 H10 121.984 C6 C5 H11 121.984
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability