Jump to
S1C2
S1C3
Energy calculated at MP3/6-31G*
| | hartrees |
|---|
| Energy at 0K | -691.747303 |
| Energy at 298.15K | -691.746720 |
| HF Energy | -691.442185 |
| Nuclear repulsion energy | 60.462881 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at MP3/6-31G*
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| K1 |
0.000 |
0.000 |
1.323 |
| C2 |
0.000 |
0.000 |
-1.303 |
| N3 |
0.000 |
0.000 |
-2.475 |
Atom - Atom Distances (Å)
| |
K1 |
C2 |
N3 |
| K1 | | 2.6257 | 3.7981 |
C2 | 2.6257 | | 1.1724 | N3 | 3.7981 | 1.1724 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| K1 |
C2 |
N3 |
180.000 |
|
K1 |
N3 |
C2 |
0.000 |
| C2 |
K1 |
N3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at MP3/6-31G*
| | hartrees |
|---|
| Energy at 0K | -691.747748 |
| Energy at 298.15K | |
| HF Energy | -691.447574 |
| Nuclear repulsion energy | 63.875205 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at MP3/6-31G*
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| K1 |
0.000 |
0.000 |
1.224 |
| C2 |
0.000 |
0.000 |
-2.425 |
| N3 |
0.000 |
0.000 |
-1.244 |
Atom - Atom Distances (Å)
| |
K1 |
C2 |
N3 |
| K1 | | 3.6488 | 2.4685 |
C2 | 3.6488 | | 1.1803 | N3 | 2.4685 | 1.1803 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| K1 |
C2 |
N3 |
0.000 |
|
K1 |
N3 |
C2 |
180.000 |
| C2 |
K1 |
N3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at MP3/6-31G*
| | hartrees |
|---|
| Energy at 0K | -691.755050 |
| Energy at 298.15K | -691.754967 |
| HF Energy | -691.450599 |
| Nuclear repulsion energy | 68.211640 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at MP3/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| K1 |
0.000 |
1.049 |
0.000 |
| C2 |
0.633 |
-1.589 |
0.000 |
| N3 |
-0.543 |
-1.486 |
0.000 |
Atom - Atom Distances (Å)
| |
K1 |
C2 |
N3 |
| K1 | | 2.7127 | 2.5922 |
C2 | 2.7127 | | 1.1808 | N3 | 2.5922 | 1.1808 | |
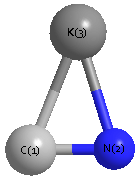 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| K1 |
C2 |
N3 |
71.495 |
|
K1 |
N3 |
C2 |
82.913 |
| C2 |
K1 |
N3 |
25.592 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
