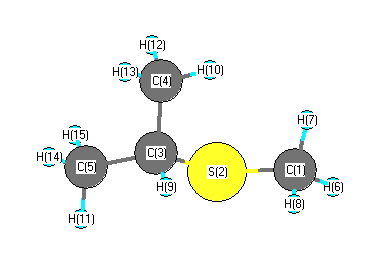
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3165 |
3028 |
7.38 |
|
|
|
| 2 |
A |
3148 |
3012 |
7.89 |
|
|
|
| 3 |
A |
3147 |
3011 |
27.32 |
|
|
|
| 4 |
A |
3144 |
3008 |
25.40 |
|
|
|
| 5 |
A |
3132 |
2996 |
32.71 |
|
|
|
| 6 |
A |
3124 |
2988 |
15.97 |
|
|
|
| 7 |
A |
3063 |
2930 |
22.11 |
|
|
|
| 8 |
A |
3058 |
2926 |
28.76 |
|
|
|
| 9 |
A |
3056 |
2923 |
21.32 |
|
|
|
| 10 |
A |
3051 |
2919 |
23.46 |
|
|
|
| 11 |
A |
1533 |
1466 |
8.16 |
|
|
|
| 12 |
A |
1524 |
1458 |
5.83 |
|
|
|
| 13 |
A |
1514 |
1448 |
9.69 |
|
|
|
| 14 |
A |
1512 |
1446 |
4.49 |
|
|
|
| 15 |
A |
1506 |
1440 |
4.67 |
|
|
|
| 16 |
A |
1493 |
1428 |
10.26 |
|
|
|
| 17 |
A |
1440 |
1378 |
5.16 |
|
|
|
| 18 |
A |
1424 |
1362 |
7.63 |
|
|
|
| 19 |
A |
1387 |
1327 |
2.73 |
|
|
|
| 20 |
A |
1359 |
1300 |
1.53 |
|
|
|
| 21 |
A |
1296 |
1240 |
32.85 |
|
|
|
| 22 |
A |
1194 |
1143 |
11.26 |
|
|
|
| 23 |
A |
1150 |
1101 |
1.14 |
|
|
|
| 24 |
A |
1091 |
1044 |
4.41 |
|
|
|
| 25 |
A |
993 |
950 |
9.62 |
|
|
|
| 26 |
A |
985 |
942 |
5.59 |
|
|
|
| 27 |
A |
970 |
928 |
0.87 |
|
|
|
| 28 |
A |
942 |
901 |
1.72 |
|
|
|
| 29 |
A |
904 |
865 |
0.91 |
|
|
|
| 30 |
A |
740 |
708 |
0.85 |
|
|
|
| 31 |
A |
646 |
618 |
2.02 |
|
|
|
| 32 |
A |
423 |
404 |
1.03 |
|
|
|
| 33 |
A |
345 |
330 |
0.84 |
|
|
|
| 34 |
A |
328 |
314 |
0.44 |
|
|
|
| 35 |
A |
257 |
245 |
0.06 |
|
|
|
| 36 |
A |
231 |
221 |
0.01 |
|
|
|
| 37 |
A |
202 |
194 |
0.53 |
|
|
|
| 38 |
A |
159 |
152 |
0.50 |
|
|
|
| 39 |
A |
68 |
65 |
1.99 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29349.6 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 28078.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.664 |
|
|
-0.320 |
| 2 |
S |
0.101 |
|
|
-0.270 |
| 3 |
C |
-0.277 |
|
|
0.225 |
| 4 |
C |
-0.498 |
|
|
-0.395 |
| 5 |
C |
-0.516 |
|
|
-0.482 |
| 6 |
H |
0.207 |
|
|
0.178 |
| 7 |
H |
0.199 |
|
|
0.119 |
| 8 |
H |
0.195 |
|
|
0.105 |
| 9 |
H |
0.186 |
|
|
0.081 |
| 10 |
H |
0.171 |
|
|
0.119 |
| 11 |
H |
0.188 |
|
|
0.124 |
| 12 |
H |
0.183 |
|
|
0.114 |
| 13 |
H |
0.169 |
|
|
0.103 |
| 14 |
H |
0.173 |
|
|
0.133 |
| 15 |
H |
0.183 |
|
|
0.166 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.273 |
1.417 |
0.983 |
1.746 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.238 |
1.436 |
0.992 |
1.762 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.506 |
1.467 |
0.530 |
| y |
1.467 |
-41.611 |
0.031 |
| z |
0.530 |
0.031 |
-40.979 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.789 |
1.467 |
0.530 |
| y |
1.467 |
-2.869 |
0.031 |
| z |
0.530 |
0.031 |
-1.921 |
|
| Polar |
|---|
| 3z2-r2 | -3.842 |
|---|
| x2-y2 | 5.105 |
|---|
| xy | 1.467 |
|---|
| xz | 0.530 |
|---|
| yz | 0.031 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
183.484 |
| (<r2>)1/2 |
13.546 |