Jump to
S1C2
Vibrational Frequencies calculated at B2PLYP/6-31G*
Geometric Data calculated at B2PLYP/6-31G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP/6-31G*
| | hartrees |
|---|
| Energy at 0K | -556.259743 |
| Energy at 298.15K | |
| HF Energy | -556.043859 |
| Nuclear repulsion energy | 240.576126 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3187 |
3025 |
8.76 |
|
|
|
| 2 |
A |
3173 |
3012 |
12.20 |
|
|
|
| 3 |
A |
3168 |
3007 |
25.93 |
|
|
|
| 4 |
A |
3165 |
3005 |
24.33 |
|
|
|
| 5 |
A |
3153 |
2993 |
36.42 |
|
|
|
| 6 |
A |
3145 |
2986 |
12.65 |
|
|
|
| 7 |
A |
3089 |
2932 |
23.22 |
|
|
|
| 8 |
A |
3081 |
2925 |
24.85 |
|
|
|
| 9 |
A |
3078 |
2922 |
20.62 |
|
|
|
| 10 |
A |
3075 |
2918 |
20.30 |
|
|
|
| 11 |
A |
1558 |
1479 |
7.10 |
|
|
|
| 12 |
A |
1549 |
1471 |
4.51 |
|
|
|
| 13 |
A |
1540 |
1462 |
8.93 |
|
|
|
| 14 |
A |
1538 |
1460 |
3.29 |
|
|
|
| 15 |
A |
1532 |
1454 |
3.78 |
|
|
|
| 16 |
A |
1518 |
1441 |
8.82 |
|
|
|
| 17 |
A |
1465 |
1391 |
3.77 |
|
|
|
| 18 |
A |
1448 |
1375 |
4.98 |
|
|
|
| 19 |
A |
1414 |
1342 |
3.90 |
|
|
|
| 20 |
A |
1381 |
1311 |
1.85 |
|
|
|
| 21 |
A |
1322 |
1255 |
33.75 |
|
|
|
| 22 |
A |
1214 |
1153 |
9.73 |
|
|
|
| 23 |
A |
1161 |
1102 |
1.17 |
|
|
|
| 24 |
A |
1109 |
1052 |
3.38 |
|
|
|
| 25 |
A |
1010 |
959 |
7.70 |
|
|
|
| 26 |
A |
1002 |
951 |
5.08 |
|
|
|
| 27 |
A |
985 |
935 |
0.43 |
|
|
|
| 28 |
A |
957 |
908 |
1.36 |
|
|
|
| 29 |
A |
911 |
865 |
0.83 |
|
|
|
| 30 |
A |
743 |
705 |
0.89 |
|
|
|
| 31 |
A |
652 |
619 |
2.03 |
|
|
|
| 32 |
A |
430 |
409 |
0.95 |
|
|
|
| 33 |
A |
351 |
333 |
0.72 |
|
|
|
| 34 |
A |
335 |
318 |
0.37 |
|
|
|
| 35 |
A |
261 |
248 |
0.06 |
|
|
|
| 36 |
A |
236 |
224 |
0.01 |
|
|
|
| 37 |
A |
212 |
201 |
0.48 |
|
|
|
| 38 |
A |
164 |
155 |
0.49 |
|
|
|
| 39 |
A |
68 |
65 |
1.86 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29691.0 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 28182.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
2.089 |
0.371 |
0.294 |
| S2 |
0.765 |
-0.715 |
-0.318 |
| C3 |
-0.720 |
0.125 |
0.367 |
| C4 |
-0.968 |
1.495 |
-0.263 |
| C5 |
-1.911 |
-0.812 |
0.160 |
| H6 |
3.035 |
-0.103 |
0.027 |
| H7 |
2.056 |
1.361 |
-0.165 |
| H8 |
2.044 |
0.469 |
1.382 |
| H9 |
-0.547 |
0.245 |
1.443 |
| H10 |
-0.124 |
2.173 |
-0.114 |
| H11 |
-1.751 |
-1.775 |
0.651 |
| H12 |
-1.140 |
1.396 |
-1.338 |
| H13 |
-1.852 |
1.960 |
0.188 |
| H14 |
-2.818 |
-0.360 |
0.573 |
| H15 |
-2.080 |
-0.995 |
-0.906 |
Atom - Atom Distances (Å)
| |
C1 |
S2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
| C1 | | 1.8185 | 2.8209 | 3.3041 | 4.1729 | 1.0914 | 1.0915 | 1.0928 | 2.8779 | 2.8832 | 4.4128 | 3.7608 | 4.2503 | 4.9686 | 4.5483 |
S2 | 1.8185 | | 1.8388 | 2.8086 | 2.7197 | 2.3767 | 2.4490 | 2.4349 | 2.3965 | 3.0287 | 2.8968 | 3.0212 | 3.7762 | 3.7088 | 2.9188 | C3 | 2.8209 | 1.8388 | | 1.5276 | 1.5291 | 3.7778 | 3.0848 | 2.9650 | 1.0961 | 2.1865 | 2.1798 | 2.1680 | 2.1634 | 2.1626 | 2.1733 | C4 | 3.3041 | 2.8086 | 1.5276 | | 2.5276 | 4.3199 | 3.0283 | 3.5818 | 2.1559 | 1.0928 | 3.4838 | 1.0935 | 1.0957 | 2.7495 | 2.8013 | C5 | 4.1729 | 2.7197 | 1.5291 | 2.5276 | | 4.9983 | 4.5342 | 4.3331 | 2.1500 | 3.4897 | 1.0925 | 2.7776 | 2.7734 | 1.0943 | 1.0943 | H6 | 1.0914 | 2.3767 | 3.7778 | 4.3199 | 4.9983 | | 1.7713 | 1.7729 | 3.8674 | 3.8964 | 5.1078 | 4.6419 | 5.3069 | 5.8840 | 5.2760 | H7 | 1.0915 | 2.4490 | 3.0848 | 3.0283 | 4.5342 | 1.7713 | | 1.7849 | 3.2562 | 2.3272 | 4.9985 | 3.4049 | 3.9688 | 5.2207 | 4.8171 | H8 | 1.0928 | 2.4349 | 2.9650 | 3.5818 | 4.3331 | 1.7729 | 1.7849 | | 2.6017 | 3.1374 | 4.4688 | 4.2894 | 4.3390 | 4.9982 | 4.9384 | H9 | 2.8779 | 2.3965 | 1.0961 | 2.1559 | 2.1500 | 3.8674 | 3.2562 | 2.6017 | | 2.5139 | 2.4807 | 3.0677 | 2.4941 | 2.5060 | 3.0663 | H10 | 2.8832 | 3.0287 | 2.1865 | 1.0928 | 3.4897 | 3.8964 | 2.3272 | 3.1374 | 2.5139 | | 4.3377 | 1.7704 | 1.7662 | 3.7607 | 3.8063 | H11 | 4.4128 | 2.8968 | 2.1798 | 3.4838 | 1.0925 | 5.1078 | 4.9985 | 4.4688 | 2.4807 | 4.3377 | | 3.7929 | 3.7652 | 1.7736 | 1.7724 | H12 | 3.7608 | 3.0212 | 2.1680 | 1.0935 | 2.7776 | 4.6419 | 3.4049 | 4.2894 | 3.0677 | 1.7704 | 3.7929 | | 1.7753 | 3.0902 | 2.6054 | H13 | 4.2503 | 3.7762 | 2.1634 | 1.0957 | 2.7734 | 5.3069 | 3.9688 | 4.3390 | 2.4941 | 1.7662 | 3.7652 | 1.7753 | | 2.5431 | 3.1591 | H14 | 4.9686 | 3.7088 | 2.1626 | 2.7495 | 1.0943 | 5.8840 | 5.2207 | 4.9982 | 2.5060 | 3.7607 | 1.7736 | 3.0902 | 2.5431 | | 1.7694 | H15 | 4.5483 | 2.9188 | 2.1733 | 2.8013 | 1.0943 | 5.2760 | 4.8171 | 4.9384 | 3.0663 | 3.8063 | 1.7724 | 2.6054 | 3.1591 | 1.7694 | |
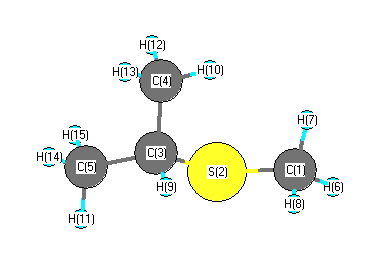 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
S2 |
C3 |
100.940 |
|
S2 |
C1 |
H6 |
106.849 |
| S2 |
C1 |
H7 |
112.197 |
|
S2 |
C1 |
H8 |
111.052 |
| S2 |
C3 |
C4 |
112.758 |
|
S2 |
C3 |
C5 |
107.354 |
| S2 |
C3 |
H9 |
106.732 |
|
C3 |
C4 |
H10 |
112.041 |
| C3 |
C4 |
H12 |
110.514 |
|
C3 |
C4 |
H13 |
110.023 |
| C3 |
C5 |
H11 |
111.415 |
|
C3 |
C5 |
H14 |
109.941 |
| C3 |
C5 |
H15 |
110.788 |
|
C4 |
C3 |
C5 |
111.561 |
| C4 |
C3 |
H9 |
109.406 |
|
C5 |
C3 |
H9 |
108.846 |
| H6 |
C1 |
H7 |
108.475 |
|
H6 |
C1 |
H8 |
108.526 |
| H7 |
C1 |
H8 |
109.608 |
|
H10 |
C4 |
H12 |
108.146 |
| H10 |
C4 |
H13 |
107.613 |
|
H11 |
C5 |
H14 |
108.403 |
| H11 |
C5 |
H15 |
108.290 |
|
H12 |
C4 |
H13 |
108.383 |
| H14 |
C5 |
H15 |
107.898 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.614 |
|
|
|
| 2 |
S |
0.097 |
|
|
|
| 3 |
C |
-0.256 |
|
|
|
| 4 |
C |
-0.453 |
|
|
|
| 5 |
C |
-0.470 |
|
|
|
| 6 |
H |
0.190 |
|
|
|
| 7 |
H |
0.183 |
|
|
|
| 8 |
H |
0.179 |
|
|
|
| 9 |
H |
0.169 |
|
|
|
| 10 |
H |
0.156 |
|
|
|
| 11 |
H |
0.173 |
|
|
|
| 12 |
H |
0.167 |
|
|
|
| 13 |
H |
0.153 |
|
|
|
| 14 |
H |
0.158 |
|
|
|
| 15 |
H |
0.167 |
|
|
|
Electric dipole moments
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.589 |
-0.025 |
-0.067 |
| y |
-0.025 |
8.360 |
0.303 |
| z |
-0.067 |
0.303 |
7.291 |
<r2> (average value of r
2) Å
2
| <r2> |
183.666 |
| (<r2>)1/2 |
13.552 |
