return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H12 (cis-1,2-dimethylcyclobutane)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-235.524687
Energy at 298.15K 
HF Energy-235.266012
Nuclear repulsion energy251.473328
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3156 2995 101.58      
2 A' 3149 2989 28.97      
3 A' 3142 2982 21.92      
4 A' 3098 2940 15.28      
5 A' 3089 2932 54.77      
6 A' 3073 2917 40.03      
7 A' 1558 1479 4.90      
8 A' 1554 1475 7.53      
9 A' 1548 1469 1.00      
10 A' 1459 1385 3.92      
11 A' 1396 1325 3.56      
12 A' 1322 1255 0.15      
13 A' 1287 1221 1.22      
14 A' 1218 1156 0.80      
15 A' 1146 1088 0.64      
16 A' 1050 997 0.52      
17 A' 999 948 1.19      
18 A' 982 932 0.38      
19 A' 923 876 0.42      
20 A' 778 739 0.38      
21 A' 754 716 1.38      
22 A' 362 344 0.20      
23 A' 284 270 0.02      
24 A' 234 222 0.02      
25 A" 3142 2983 13.66      
26 A" 3138 2979 43.87      
27 A" 3133 2974 1.57      
28 A" 3088 2931 42.80      
29 A" 3074 2917 10.74      
30 A" 3068 2912 10.90      
31 A" 1540 1462 3.26      
32 A" 1539 1461 3.68      
33 A" 1525 1448 0.82      
34 A" 1451 1378 1.85      
35 A" 1397 1326 3.13      
36 A" 1329 1261 0.72      
37 A" 1291 1226 0.03      
38 A" 1268 1204 0.53      
39 A" 1209 1148 0.22      
40 A" 1141 1083 1.89      
41 A" 1045 992 1.91      
42 A" 915 868 0.22      
43 A" 878 833 0.02      
44 A" 683 648 0.40      
45 A" 473 449 0.82      
46 A" 337 320 0.01      
47 A" 240 228 0.01      
48 A" 104i 99i 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 37179.3 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 35290.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.13769 0.12689 0.07807

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.510 -0.037 0.785
C2 0.510 -0.037 -0.785
C3 -0.211 1.341 0.776
C4 -0.211 1.341 -0.776
C5 -0.211 -1.163 1.513
C6 -0.211 -1.163 -1.513
H7 1.530 0.036 1.178
H8 1.530 0.036 -1.178
H9 -1.208 1.321 1.227
H10 -1.208 1.321 -1.227
H11 0.349 2.157 1.238
H12 0.349 2.157 -1.238
H13 -1.251 -1.256 1.186
H14 -1.251 -1.256 -1.186
H15 0.279 -2.129 1.353
H16 -0.223 -0.971 2.591
H17 0.279 -2.129 -1.353
H18 -0.223 -0.971 -2.591

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18
C11.56951.55502.20281.52342.65921.09522.21312.23392.97342.24632.98852.17932.91052.18052.16193.00023.5788
C21.56952.20281.55502.65921.52342.21311.09522.97342.23392.98852.24632.91052.17933.00023.57882.18052.1619
C31.55502.20281.55112.61063.39272.21212.92371.09432.23681.09312.24392.82743.41643.55162.93964.10014.0841
C42.20281.55501.55113.39272.61062.92372.21212.23681.09432.24391.09313.41642.82744.10014.08413.55162.9396
C51.52342.65922.61063.39273.02702.14073.42282.69213.83073.37904.34881.09422.89391.09481.09503.06414.1094
C62.65921.52343.39272.61063.02703.42282.14073.83072.69214.34883.37902.89391.09423.06414.10941.09481.0950
H71.09522.21312.21212.92372.14073.42282.35633.02423.86362.42793.42463.06653.87172.50712.46653.55834.2774
H82.21311.09522.92372.21213.42282.14072.35633.86363.02423.42462.42793.87173.06653.55834.27742.50712.4665
H92.23392.97341.09432.23682.69213.83073.02423.86362.45361.76783.03302.57793.53033.75892.84334.55724.5608
H102.97342.23392.23681.09433.83072.69213.86363.02422.45363.03301.76783.53032.57794.55724.56083.75892.8433
H112.24632.98851.09312.24393.37904.34882.42793.42461.76783.03302.47563.77064.48194.28883.45635.00924.9776
H122.98852.24632.24391.09314.34883.37903.42462.42793.03301.76782.47564.48193.77065.00924.97764.28883.4563
H132.17932.91052.82743.41641.09422.89393.06653.87172.57793.53033.77064.48192.37111.76921.76493.08963.9247
H142.91052.17933.41642.82742.89391.09423.87173.06653.53032.57794.48193.77062.37113.08963.92471.76921.7649
H152.18053.00023.55164.10011.09483.06412.50713.55833.75894.55724.28885.00921.76923.08961.76842.70574.1413
H162.16193.57882.93964.08411.09504.10942.46654.27742.84334.56083.45634.97761.76493.92471.76844.14135.1828
H173.00022.18054.10013.55163.06411.09483.55832.50714.55723.75895.00924.28883.08961.76922.70574.14131.7684
H183.57882.16194.08412.93964.10941.09504.27742.46654.56082.84334.97763.45633.92471.76494.14135.18281.7684

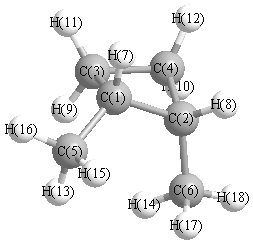
picture of cis-1,2-dimethylcyclobutane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 89.662 C1 C2 C6 118.580
C1 C2 H8 111.053 C1 C3 C4 90.338
C1 C3 H9 113.828 C1 C3 H11 114.930
C1 C5 H13 111.679 C1 C5 H15 111.735
C1 C5 H16 110.242 C2 C1 C3 89.662
C2 C1 C5 118.580 C2 C1 H7 111.053
C2 C4 C3 90.338 C2 C4 H10 113.828
C2 C4 H12 114.930 C2 C6 H14 111.679
C2 C6 H17 111.735 C2 C6 H18 110.242
C3 C1 C5 115.996 C3 C1 H7 111.998
C3 C4 H10 114.353 C3 C4 H12 115.017
C4 C2 C6 115.996 C4 C2 H8 111.998
C4 C3 H9 114.353 C4 C3 H11 115.017
C5 C1 H7 108.570 C6 C2 H8 108.570
H9 C3 H11 107.834 H10 C4 H12 107.834
H13 C5 H15 107.837 H13 C5 H16 107.444
H14 C6 H17 107.837 H14 C6 H18 107.444
H15 C5 H16 107.716 H17 C6 H18 107.716
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.109      
2 C -0.109      
3 C -0.305      
4 C -0.305      
5 C -0.465      
6 C -0.465      
7 H 0.139      
8 H 0.139      
9 H 0.146      
10 H 0.146      
11 H 0.148      
12 H 0.148      
13 H 0.147      
14 H 0.147      
15 H 0.149      
16 H 0.151      
17 H 0.149      
18 H 0.151      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 8.101 -0.023 0.000
y -0.023 9.210 0.000
z 0.000 0.000 9.329


<r2> (average value of r2) Å2
<r2> 176.010
(<r2>)1/2 13.267