return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8 (Spiropentane)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes D2D 1A1
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-195.021185
Energy at 298.15K-195.029853
HF Energy-194.807796
Nuclear repulsion energy170.336491
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3167 3006 0.00      
2 A1 1542 1463 0.00      
3 A1 1113 1057 0.00      
4 A1 1078 1023 0.00      
5 A1 615 584 0.00      
6 A2 3242 3077 0.00      
7 A2 1199 1138 0.00      
8 A2 858 814 0.00      
9 B1 3241 3076 0.00      
10 B1 1210 1148 0.00      
11 B1 1050 996 0.00      
12 B1 297 282 0.00      
13 B2 3167 3006 35.60      
14 B2 1619 1537 2.01      
15 B2 1474 1399 0.14      
16 B2 1045 992 8.80      
17 B2 925 878 18.47      
18 E 3254 3088 25.04      
19 E 3164 3003 23.38      
20 E 1509 1433 0.23      
20 E 1509 1433 0.23      
21 E 1210 1149 2.31      
21 E 1210 1149 2.31      
22 E 1100 1044 1.84      
22 E 1100 1044 1.84      
23 E 910 864 4.85      
23 E 910 864 4.85      
24 E 805 764 0.29      
24 E 805 764 0.29      
25 E 309 293 0.25      
25 E 309 293 0.25      

Unscaled Zero Point Vibrational Energy (zpe) 25680.6 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 24376.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.35356 0.13899 0.13899

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is D2d

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.000
C2 0.000 0.764 1.270
C3 0.000 -0.764 1.270
C4 -0.764 0.000 -1.270
C5 0.764 0.000 -1.270
H6 0.913 1.268 1.574
H7 -0.913 1.268 1.574
H8 -0.913 -1.268 1.574
H9 0.913 -1.268 1.574
H10 -1.268 -0.913 -1.574
H11 -1.268 0.913 -1.574
H12 1.268 0.913 -1.574
H13 1.268 -0.913 -1.574

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13
C11.48231.48231.48231.48232.21772.21772.21772.21772.21772.21772.21772.2177
C21.48231.52742.76092.76091.08611.08612.24802.24803.53673.11773.11773.5367
C31.48231.52742.76092.76092.24802.24801.08611.08613.11773.53673.53673.1177
C41.48232.76092.76091.52743.53673.11773.11773.53671.08611.08612.24802.2480
C51.48232.76092.76091.52743.11773.53673.53673.11772.24802.24801.08611.0861
H62.21771.08612.24803.53673.11771.82563.12492.53614.40683.84573.18743.8457
H72.21771.08612.24803.11773.53671.82562.53613.12493.84573.18743.84574.4068
H82.21772.24801.08613.11773.53673.12492.53611.82563.18743.84574.40683.8457
H92.21772.24801.08613.53673.11772.53613.12491.82563.84574.40683.84573.1874
H102.21773.53673.11771.08612.24804.40683.84573.18743.84571.82563.12492.5361
H112.21773.11773.53671.08612.24803.84573.18743.84574.40681.82562.53613.1249
H122.21773.11773.53672.24801.08613.18743.84574.40683.84573.12492.53611.8256
H132.21773.53673.11772.24801.08613.84574.40683.84573.18742.53613.12491.8256

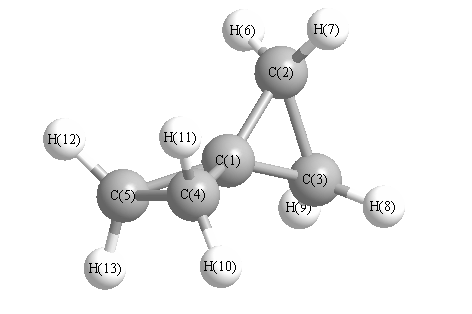
picture of Spiropentane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 58.988 C1 C2 H6 118.596
C1 C2 H7 118.596 C1 C3 C2 58.988
C1 C3 H8 118.596 C1 C3 H9 118.596
C1 C4 C5 58.988 C1 C4 H10 118.596
C1 C4 H11 118.596 C1 C5 C4 58.988
C1 C5 H12 118.596 C1 C5 H13 118.596
C2 C1 C3 62.024 C2 C1 C4 137.270
C2 C1 C5 137.270 C2 C3 H8 117.670
C2 C3 H9 117.670 C3 C1 C4 137.270
C3 C1 C5 137.270 C3 C2 H6 117.670
C3 C2 H7 117.670 C4 C1 C5 62.024
C4 C5 H12 117.670 C4 C5 H13 117.670
C5 C4 H10 117.670 C5 C4 H11 117.670
H6 C2 H7 114.375 H8 C3 H9 114.375
H10 C4 H11 114.375 H12 C5 H13 114.375
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C 0.089      
2 C -0.348      
3 C -0.348      
4 C -0.348      
5 C -0.348      
6 H 0.163      
7 H 0.163      
8 H 0.163      
9 H 0.163      
10 H 0.163      
11 H 0.163      
12 H 0.163      
13 H 0.163      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 6.834 0.000 0.000
y 0.000 6.834 0.000
z 0.000 0.000 7.817


<r2> (average value of r2) Å2
<r2> 111.534
(<r2>)1/2 10.561