return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H6 (3-Penten-1-yne, (Z)-)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-193.819760
Energy at 298.15K-193.824161
HF Energy-193.607233
Nuclear repulsion energy140.997952
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3512 3334 60.33      
2 A' 3213 3050 28.56      
3 A' 3194 3032 2.65      
4 A' 3176 3014 4.00      
5 A' 3072 2916 20.91      
6 A' 2188 2077 0.21      
7 A' 1708 1621 4.02      
8 A' 1532 1455 6.71      
9 A' 1469 1395 3.46      
10 A' 1434 1361 6.63      
11 A' 1289 1223 0.07      
12 A' 1145 1087 0.51      
13 A' 989 939 18.11      
14 A' 918 872 1.38      
15 A' 666 632 3.31      
16 A' 612 581 44.37      
17 A' 377 357 4.28      
18 A' 161 153 0.11      
19 A" 3126 2967 19.51      
20 A" 1534 1456 6.43      
21 A" 1088 1033 0.55      
22 A" 985 935 0.38      
23 A" 759 721 21.02      
24 A" 551 523 62.32      
25 A" 480 456 1.85      
26 A" 276 262 0.23      
27 A" 143 135 0.54      

Unscaled Zero Point Vibrational Energy (zpe) 19797.8 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 18792.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.39890 0.11071 0.08807

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 3.058 -0.838 0.000
C2 2.188 -0.334 0.000
C3 1.143 0.276 0.000
H4 0.061 2.022 0.000
C5 0.000 0.980 0.000
H6 -2.058 1.064 0.000
C7 -1.235 0.423 0.000
H8 -0.554 -1.579 0.000
H9 -2.068 -1.305 0.884
H10 -2.068 -1.305 -0.884
C11 -1.491 -1.022 0.000

Atom - Atom Distances (Å)
  H1 C2 C3 H4 C5 H6 C7 H8 H9 H10 C11
H11.00612.21604.14343.55835.45874.47523.68795.22285.22284.5527
C21.00611.20993.17412.55234.47033.50603.01194.45384.45383.7425
C32.21601.20992.05461.34263.29652.38252.51383.68583.68582.9356
H44.14343.17412.05461.04342.32572.05803.65344.04714.04713.4168
C53.55832.55231.34261.04342.06001.35512.61903.20613.20612.4965
H65.45874.47033.29652.32572.06001.04273.04082.52782.52782.1618
C74.47523.50602.38252.05801.35511.04272.11522.11202.11201.4680
H83.68793.01192.51383.65342.61903.04082.11521.77431.77431.0894
H95.22284.45383.68584.04713.20612.52782.11201.77431.76831.0930
H105.22284.45383.68584.04713.20612.52782.11201.77431.76831.0930
C114.55273.74252.93563.41682.49652.16181.46801.08941.09301.0930

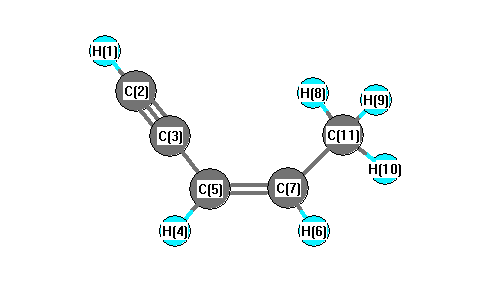
picture of 3-Penten-1-yne, (Z)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C2 C3 179.870 C2 C3 C5 178.558
C3 C5 H4 118.338 C3 C5 C7 124.055
H4 C5 C7 117.608 C5 C7 H6 117.848
C5 C7 C11 124.286 H6 C7 C11 117.866
C7 C11 H8 110.731 C7 C11 H9 110.253
C7 C11 H10 110.253 H8 C11 H9 108.778
H8 C11 H10 108.778 H9 C11 H10 107.981
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 H 0.234      
2 C -0.486      
3 C 0.227      
4 H 0.182      
5 C -0.239      
6 H 0.159      
7 C -0.091      
8 H 0.184      
9 H 0.169      
10 H 0.169      
11 C -0.508      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 11.625 -0.774 0.000
y -0.774 6.512 0.000
z 0.000 0.000 3.642


<r2> (average value of r2) Å2
<r2> 133.412
(<r2>)1/2 11.550