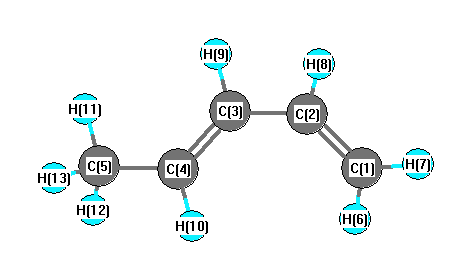
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3274 |
3113 |
15.12 |
|
|
|
| 2 |
A' |
3198 |
3041 |
37.30 |
|
|
|
| 3 |
A' |
3192 |
3036 |
13.32 |
|
|
|
| 4 |
A' |
3182 |
3026 |
9.30 |
|
|
|
| 5 |
A' |
3173 |
3017 |
5.97 |
|
|
|
| 6 |
A' |
3162 |
3007 |
3.31 |
|
|
|
| 7 |
A' |
3058 |
2908 |
27.67 |
|
|
|
| 8 |
A' |
1757 |
1671 |
9.72 |
|
|
|
| 9 |
A' |
1698 |
1615 |
2.00 |
|
|
|
| 10 |
A' |
1517 |
1443 |
7.67 |
|
|
|
| 11 |
A' |
1491 |
1418 |
12.63 |
|
|
|
| 12 |
A' |
1441 |
1370 |
3.52 |
|
|
|
| 13 |
A' |
1410 |
1341 |
1.67 |
|
|
|
| 14 |
A' |
1333 |
1267 |
1.05 |
|
|
|
| 15 |
A' |
1304 |
1240 |
1.41 |
|
|
|
| 16 |
A' |
1206 |
1147 |
1.72 |
|
|
|
| 17 |
A' |
1075 |
1022 |
0.02 |
|
|
|
| 18 |
A' |
982 |
934 |
13.44 |
|
|
|
| 19 |
A' |
911 |
867 |
1.99 |
|
|
|
| 20 |
A' |
619 |
589 |
5.80 |
|
|
|
| 21 |
A' |
388 |
369 |
4.27 |
|
|
|
| 22 |
A' |
219 |
208 |
0.31 |
|
|
|
| 23 |
A" |
3112 |
2959 |
18.66 |
|
|
|
| 24 |
A" |
1507 |
1433 |
7.44 |
|
|
|
| 25 |
A" |
1069 |
1016 |
0.76 |
|
|
|
| 26 |
A" |
1049 |
997 |
11.95 |
|
|
|
| 27 |
A" |
998 |
949 |
1.81 |
|
|
|
| 28 |
A" |
931 |
885 |
48.98 |
|
|
|
| 29 |
A" |
811 |
772 |
7.11 |
|
|
|
| 30 |
A" |
647 |
616 |
25.97 |
|
|
|
| 31 |
A" |
360 |
342 |
0.08 |
|
|
|
| 32 |
A" |
151 |
144 |
0.06 |
|
|
|
| 33 |
A" |
124 |
118 |
0.65 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25173.3 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 23939.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.405 |
|
|
|
| 2 |
C |
-0.095 |
|
|
|
| 3 |
C |
-0.149 |
|
|
|
| 4 |
C |
-0.146 |
|
|
|
| 5 |
C |
-0.561 |
|
|
|
| 6 |
H |
0.166 |
|
|
|
| 7 |
H |
0.168 |
|
|
|
| 8 |
H |
0.157 |
|
|
|
| 9 |
H |
0.154 |
|
|
|
| 10 |
H |
0.159 |
|
|
|
| 11 |
H |
0.181 |
|
|
|
| 12 |
H |
0.186 |
|
|
|
| 13 |
H |
0.186 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.488 |
-0.208 |
0.000 |
0.531 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-28.511 |
0.167 |
0.000 |
| y |
0.167 |
-29.121 |
0.000 |
| z |
0.000 |
0.000 |
-34.299 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.200 |
0.167 |
0.000 |
| y |
0.167 |
2.284 |
0.000 |
| z |
0.000 |
0.000 |
-5.483 |
|
| Polar |
|---|
| 3z2-r2 | -10.967 |
|---|
| x2-y2 | 0.611 |
|---|
| xy | 0.167 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
14.167 |
0.857 |
0.000 |
| y |
0.857 |
7.189 |
0.000 |
| z |
0.000 |
0.000 |
3.817 |
<r2> (average value of r
2) Å
2
| <r2> |
155.613 |
| (<r2>)1/2 |
12.474 |