return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H6O (cis-2-butenal)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-230.525001
Energy at 298.15K-230.530803
Nuclear repulsion energy154.751377
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3191 3039 9.06      
2 A' 3187 3035 10.16      
3 A' 3148 2998 15.05      
4 A' 3060 2914 15.69      
5 A' 2976 2834 147.44      
6 A' 1799 1713 67.77      
7 A' 1738 1655 132.94      
8 A' 1538 1464 16.60      
9 A' 1468 1398 1.24      
10 A' 1459 1390 7.09      
11 A' 1351 1287 3.17      
12 A' 1349 1284 22.05      
13 A' 1166 1110 6.13      
14 A' 1053 1002 15.72      
15 A' 920 877 25.09      
16 A' 748 713 38.68      
17 A' 400 381 3.27      
18 A' 213 202 5.24      
19 A" 3123 2974 16.20      
20 A" 1528 1455 7.08      
21 A" 1106 1053 1.14      
22 A" 1027 978 0.21      
23 A" 1005 958 34.22      
24 A" 763 727 0.45      
25 A" 242 230 5.60      
26 A" 198 189 0.01      
27 A" 140 133 6.70      

Unscaled Zero Point Vibrational Energy (zpe) 19947.5 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 18996.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
0.62942 0.08698 0.07752

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.477 0.557 0.000
C2 0.000 0.702 0.000
C3 0.803 -0.376 0.000
C4 2.301 -0.336 0.000
O5 -2.065 -0.517 0.000
H6 -2.054 1.506 0.000
H7 0.405 1.715 0.000
H8 0.327 -1.358 0.000
H9 2.677 0.694 0.000
H10 2.702 -0.853 0.882
H11 2.702 -0.853 -0.882

Atom - Atom Distances (Å)
  C1 C2 C3 C4 O5 H6 H7 H8 H9 H10 H11
C11.48402.46333.88231.22421.11002.20902.63104.15654.49764.4976
C21.48401.34352.52412.39722.20561.09102.08552.67723.23933.2393
C32.46331.34351.49902.87083.42052.12831.09162.15822.14762.1476
C43.88232.52411.49904.36954.72822.79332.22351.09601.09841.0984
O51.22422.39722.87084.36952.02233.32802.53504.89384.85894.8589
H61.11002.20563.42054.72822.02232.46713.72404.80015.38105.3810
H72.20901.09102.12832.79333.32802.46713.07402.49163.55643.5564
H82.63102.08551.09162.22352.53503.72403.07403.12012.58362.5836
H94.15652.67722.15821.09604.89384.80012.49163.12011.78071.7807
H104.49763.23932.14761.09844.85895.38103.55642.58361.78071.7650
H114.49763.23932.14761.09844.85895.38103.55642.58361.78071.7650

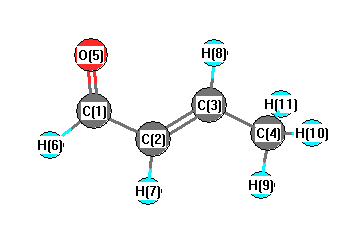
picture of cis-2-butenal state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 121.111 C1 C2 H7 117.337
C2 C1 O5 124.263 C2 C1 H6 115.728
C2 C3 C4 125.157 C2 C3 H8 117.457
C3 C2 H7 121.553 C3 C4 H9 111.599
C3 C4 H10 110.606 C3 C4 H11 110.606
C4 C3 H8 117.386 O5 C1 H6 120.009
H9 C4 H10 108.478 H9 C4 H11 108.478
H10 C4 H11 106.922
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at QCISD/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C 0.324      
2 C -0.326      
3 C -0.086      
4 C -0.527      
5 O -0.525      
6 H 0.156      
7 H 0.194      
8 H 0.235      
9 H 0.181      
10 H 0.187      
11 H 0.187      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 0.000 0.000 0.000
y 0.000 0.000 0.000
z 0.000 0.000 0.000


<r2> (average value of r2) Å2
<r2> 0.000
(<r2>)1/2 0.000