return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H6O (cis-2-butenal)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-230.973593
Energy at 298.15K-230.979438
HF Energy-230.745352
Nuclear repulsion energy155.062598
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3205 3042 20.25      
2 A' 3201 3039 0.08      
3 A' 3165 3004 13.56      
4 A' 3072 2916 13.29      
5 A' 2966 2816 168.40      
6 A' 1785 1695 67.81      
7 A' 1708 1621 177.72      
8 A' 1533 1455 24.07      
9 A' 1462 1387 2.41      
10 A' 1454 1380 3.39      
11 A' 1347 1278 1.42      
12 A' 1341 1273 26.22      
13 A' 1162 1103 5.84      
14 A' 1048 995 15.51      
15 A' 914 868 29.58      
16 A' 748 710 37.97      
17 A' 399 378 3.30      
18 A' 208 197 5.13      
19 A" 3129 2970 13.60      
20 A" 1527 1449 7.39      
21 A" 1101 1045 0.93      
22 A" 1037 985 0.20      
23 A" 1019 968 30.07      
24 A" 773 734 0.07      
25 A" 250 237 4.61      
26 A" 208 197 0.34      
27 A" 154 146 6.05      

Unscaled Zero Point Vibrational Energy (zpe) 19958.0 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 18944.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.63316 0.08720 0.07775

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.470 0.561 0.000
C2 0.000 0.700 0.000
C3 0.803 -0.377 0.000
C4 2.296 -0.345 0.000
O5 -2.066 -0.506 0.000
H6 -2.035 1.515 0.000
H7 0.408 1.708 0.000
H8 0.321 -1.353 0.000
H9 2.681 0.678 0.000
H10 2.693 -0.866 0.878
H11 2.693 -0.866 -0.878

Atom - Atom Distances (Å)
  C1 C2 C3 C4 O5 H6 H7 H8 H9 H10 H11
C11.47652.45833.87321.22251.10852.20062.62084.15234.48754.4875
C21.47651.34322.52262.39282.19161.08752.07802.68093.23673.2367
C32.45831.34321.49382.87193.41002.12231.08862.15402.14112.1411
C43.87322.52261.49384.36544.71292.78932.21781.09241.09571.0957
O51.22252.39282.87194.36542.02123.32052.53284.89264.85324.8532
H61.10852.19163.41004.71292.02122.45013.71094.78915.36565.3656
H72.20061.08752.12232.78933.32052.45013.06252.49583.55263.5526
H82.62082.07801.08862.21782.53283.71093.06253.11352.57622.5762
H94.15232.68092.15401.09244.89264.78912.49583.11351.77601.7760
H104.48753.23672.14111.09574.85325.36563.55262.57621.77601.7564
H114.48753.23672.14111.09574.85325.36563.55262.57621.77601.7564

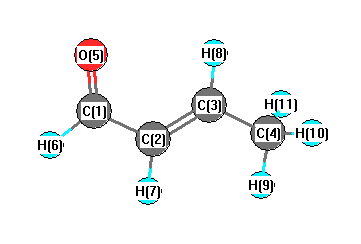
picture of cis-2-butenal state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 121.276 C1 C2 H7 117.436
C2 C1 O5 124.624 C2 C1 H6 115.210
C2 C3 C4 125.458 C2 C3 H8 117.032
C3 C2 H7 121.288 C3 C4 H9 111.858
C3 C4 H10 110.613 C3 C4 H11 110.613
C4 C3 H8 117.511 O5 C1 H6 120.166
H9 C4 H10 108.519 H9 C4 H11 108.519
H10 C4 H11 106.540
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C 0.263      
2 C -0.237      
3 C -0.066      
4 C -0.513      
5 O -0.459      
6 H 0.126      
7 H 0.158      
8 H 0.199      
9 H 0.172      
10 H 0.180      
11 H 0.180      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 9.326 -1.066 0.000
y -1.066 7.038 0.000
z 0.000 0.000 3.537


<r2> (average value of r2) Å2
<r2> 146.822
(<r2>)1/2 12.117