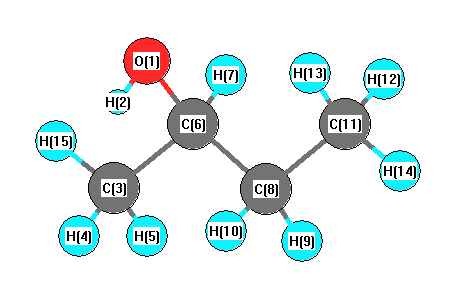
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3755 |
3593 |
8.02 |
|
|
|
| 2 |
A |
3151 |
3015 |
31.71 |
|
|
|
| 3 |
A |
3148 |
3011 |
16.92 |
|
|
|
| 4 |
A |
3130 |
2994 |
45.95 |
|
|
|
| 5 |
A |
3121 |
2986 |
37.48 |
|
|
|
| 6 |
A |
3078 |
2945 |
30.06 |
|
|
|
| 7 |
A |
3064 |
2931 |
29.33 |
|
|
|
| 8 |
A |
3059 |
2927 |
23.53 |
|
|
|
| 9 |
A |
3045 |
2913 |
19.18 |
|
|
|
| 10 |
A |
3021 |
2890 |
22.22 |
|
|
|
| 11 |
A |
1534 |
1468 |
6.64 |
|
|
|
| 12 |
A |
1522 |
1456 |
12.24 |
|
|
|
| 13 |
A |
1520 |
1454 |
1.75 |
|
|
|
| 14 |
A |
1519 |
1453 |
4.49 |
|
|
|
| 15 |
A |
1504 |
1439 |
0.55 |
|
|
|
| 16 |
A |
1441 |
1379 |
36.48 |
|
|
|
| 17 |
A |
1429 |
1367 |
16.63 |
|
|
|
| 18 |
A |
1424 |
1362 |
8.71 |
|
|
|
| 19 |
A |
1407 |
1346 |
3.39 |
|
|
|
| 20 |
A |
1356 |
1297 |
2.36 |
|
|
|
| 21 |
A |
1334 |
1277 |
5.32 |
|
|
|
| 22 |
A |
1273 |
1217 |
5.50 |
|
|
|
| 23 |
A |
1198 |
1146 |
21.48 |
|
|
|
| 24 |
A |
1146 |
1096 |
22.43 |
|
|
|
| 25 |
A |
1104 |
1057 |
73.41 |
|
|
|
| 26 |
A |
1062 |
1016 |
1.88 |
|
|
|
| 27 |
A |
1013 |
969 |
18.17 |
|
|
|
| 28 |
A |
987 |
944 |
10.51 |
|
|
|
| 29 |
A |
937 |
897 |
8.38 |
|
|
|
| 30 |
A |
824 |
789 |
5.49 |
|
|
|
| 31 |
A |
780 |
746 |
0.57 |
|
|
|
| 32 |
A |
497 |
476 |
14.41 |
|
|
|
| 33 |
A |
463 |
442 |
7.26 |
|
|
|
| 34 |
A |
374 |
358 |
14.61 |
|
|
|
| 35 |
A |
308 |
295 |
109.46 |
|
|
|
| 36 |
A |
254 |
243 |
0.45 |
|
|
|
| 37 |
A |
235 |
225 |
1.87 |
|
|
|
| 38 |
A |
218 |
208 |
0.07 |
|
|
|
| 39 |
A |
117 |
112 |
0.10 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30173.9 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 28867.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.623 |
|
|
-0.642 |
| 2 |
H |
0.393 |
|
|
0.403 |
| 3 |
C |
-0.524 |
|
|
-0.670 |
| 4 |
H |
0.153 |
|
|
0.152 |
| 5 |
H |
0.167 |
|
|
0.155 |
| 6 |
C |
0.101 |
|
|
0.436 |
| 7 |
H |
0.163 |
|
|
0.051 |
| 8 |
C |
-0.318 |
|
|
-0.111 |
| 9 |
H |
0.158 |
|
|
0.043 |
| 10 |
H |
0.143 |
|
|
0.047 |
| 11 |
C |
-0.507 |
|
|
-0.263 |
| 12 |
H |
0.165 |
|
|
0.068 |
| 13 |
H |
0.182 |
|
|
0.073 |
| 14 |
H |
0.163 |
|
|
0.078 |
| 15 |
H |
0.183 |
|
|
0.179 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.183 |
-1.233 |
1.002 |
1.600 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.177 |
-1.220 |
0.991 |
1.582 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.063 |
0.258 |
-0.718 |
| y |
0.258 |
-35.207 |
2.415 |
| z |
-0.718 |
2.415 |
-30.381 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.269 |
0.258 |
-0.718 |
| y |
0.258 |
-3.485 |
2.415 |
| z |
-0.718 |
2.415 |
3.755 |
|
| Polar |
|---|
| 3z2-r2 | 7.509 |
|---|
| x2-y2 | 2.144 |
|---|
| xy | 0.258 |
|---|
| xz | -0.718 |
|---|
| yz | 2.415 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
146.087 |
| (<r2>)1/2 |
12.087 |