return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H10O (2-Butanol, (.+/-.)-)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-233.380289
Energy at 298.15K-233.391557
HF Energy-233.148098
Nuclear repulsion energy193.205781
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3738 3548 8.66      
2 A 3175 3014 28.07      
3 A 3169 3008 21.02      
4 A 3151 2991 46.26      
5 A 3143 2983 38.18      
6 A 3101 2944 32.23      
7 A 3089 2932 19.32      
8 A 3084 2927 24.15      
9 A 3069 2913 18.38      
10 A 3045 2890 22.02      
11 A 1560 1480 5.58      
12 A 1547 1469 9.95      
13 A 1546 1467 0.08      
14 A 1543 1465 5.26      
15 A 1531 1453 0.44      
16 A 1460 1386 15.73      
17 A 1453 1379 17.72      
18 A 1444 1371 24.50      
19 A 1431 1358 3.22      
20 A 1372 1303 3.16      
21 A 1358 1289 6.22      
22 A 1292 1227 5.46      
23 A 1211 1150 13.54      
24 A 1160 1101 20.75      
25 A 1115 1058 69.49      
26 A 1068 1014 4.51      
27 A 1024 972 14.90      
28 A 1001 950 11.20      
29 A 944 896 12.48      
30 A 835 792 6.13      
31 A 790 750 0.65      
32 A 502 477 13.16      
33 A 469 445 7.90      
34 A 380 361 14.76      
35 A 312 296 111.03      
36 A 259 246 0.46      
37 A 242 230 1.79      
38 A 225 213 0.08      
39 A 120 114 0.11      

Unscaled Zero Point Vibrational Energy (zpe) 30478.3 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 28930.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.26650 0.11464 0.08903

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
O1 -0.516 1.412 0.054
H2 -0.632 1.428 1.018
C3 -1.816 -0.657 -0.001
H4 -1.966 -0.678 1.083
H5 -1.826 -1.689 -0.365
C6 -0.486 0.035 -0.331
H7 -0.367 0.067 -1.418
C8 0.730 -0.700 0.266
H9 0.723 -1.740 -0.078
H10 0.610 -0.722 1.356
C11 2.075 -0.045 -0.099
H12 2.215 -0.039 -1.182
H13 2.098 0.988 0.250
H14 2.915 -0.582 0.350
H15 -2.654 -0.120 -0.452

Atom - Atom Distances (Å)
  O1 H2 C3 H4 H5 C6 H7 C8 H9 H10 C11 H12 H13 H14 H15
O10.97142.44462.74383.39201.43021.99942.46133.38982.74162.97633.33052.65583.97962.6785
H20.97142.60592.49353.61291.94522.80312.63653.61652.50603.27803.88612.87074.13182.9406
C32.44462.60591.09471.09371.53612.15242.56032.76212.78123.94004.24584.25384.74521.0930
H42.74382.49351.09471.77192.16823.06062.81713.11642.59144.25734.79794.47094.93711.7726
H53.39203.61291.09371.77192.18382.51382.81182.56613.13594.24104.44024.78984.92091.7761
C61.43021.94521.53612.16822.18381.09341.54062.16272.14962.57192.83212.81453.52282.1775
H71.99942.80312.15243.06062.51381.09342.15072.50013.04502.77752.59463.11543.78422.4896
C82.46132.63652.56032.81712.81181.54062.15071.09541.09711.53972.17702.17332.19043.5074
H93.38983.61652.76213.11642.56612.16272.50011.09541.76272.16772.51733.07252.51573.7648
H102.74162.50602.78122.59143.13592.14963.04501.09711.76272.17203.07952.52212.51873.7802
C112.97633.27803.94004.25734.24102.57192.77751.53972.16772.17201.09281.09051.09364.7425
H123.33053.88614.24584.79794.44022.83212.59462.17702.51733.07951.09281.76611.77024.9239
H132.65582.87074.25384.47094.78982.81453.11542.17333.07252.52211.09051.76611.77264.9301
H143.97964.13184.74524.93714.92093.52283.78422.19042.51572.51871.09361.77021.77265.6458
H152.67852.94061.09301.77261.77612.17752.48963.50743.76483.78024.74254.92394.93015.6458

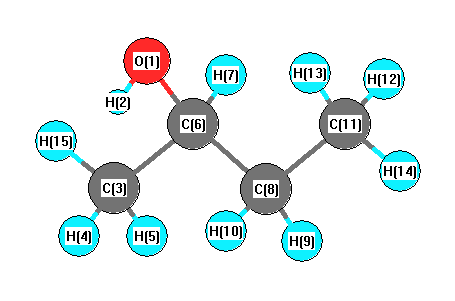
picture of 2-Butanol, (.+/-.)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
O1 C6 C3 110.949 O1 C6 H7 103.994
O1 C6 C8 111.832 H2 O1 C6 106.620
C3 C6 H7 108.709 C3 C6 C8 112.637
H4 C3 H5 108.131 H4 C3 C6 109.868
H4 C3 H15 108.249 H5 C3 C6 111.166
H5 C3 H15 108.631 C6 C3 H15 110.708
C6 C8 H9 109.092 C6 C8 H10 107.984
C6 C8 C11 113.223 H7 C6 C8 108.276
C8 C11 H12 110.424 C8 C11 H13 110.272
C8 C11 H14 111.448 H9 C8 H10 107.021
H9 C8 C11 109.543 H10 C8 C11 109.781
H12 C11 H13 107.977 H12 C11 H14 108.118
H13 C11 H14 108.497
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 O -0.674      
2 H 0.399      
3 C -0.475      
4 H 0.138      
5 H 0.152      
6 C 0.151      
7 H 0.149      
8 C -0.289      
9 H 0.143      
10 H 0.128      
11 C -0.460      
12 H 0.149      
13 H 0.170      
14 H 0.148      
15 H 0.169      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 7.433 0.047 -0.049
y 0.047 6.635 0.153
z -0.049 0.153 6.582


<r2> (average value of r2) Å2
<r2> 146.280
(<r2>)1/2 12.095