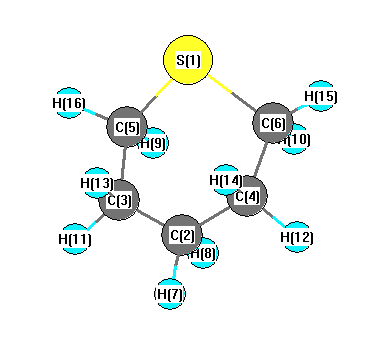
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3126 |
2990 |
21.75 |
|
|
|
| 2 |
A |
3100 |
2966 |
59.47 |
|
|
|
| 3 |
A |
3087 |
2954 |
40.59 |
|
|
|
| 4 |
A |
3055 |
2923 |
21.50 |
|
|
|
| 5 |
A |
3044 |
2913 |
24.39 |
|
|
|
| 6 |
A |
3032 |
2901 |
20.57 |
|
|
|
| 7 |
A |
1518 |
1452 |
2.23 |
|
|
|
| 8 |
A |
1503 |
1438 |
11.49 |
|
|
|
| 9 |
A |
1491 |
1427 |
9.13 |
|
|
|
| 10 |
A |
1399 |
1338 |
1.08 |
|
|
|
| 11 |
A |
1356 |
1297 |
6.73 |
|
|
|
| 12 |
A |
1283 |
1227 |
0.97 |
|
|
|
| 13 |
A |
1256 |
1202 |
7.52 |
|
|
|
| 14 |
A |
1095 |
1047 |
2.63 |
|
|
|
| 15 |
A |
1041 |
996 |
0.62 |
|
|
|
| 16 |
A |
988 |
945 |
9.29 |
|
|
|
| 17 |
A |
843 |
807 |
5.18 |
|
|
|
| 18 |
A |
833 |
797 |
0.95 |
|
|
|
| 19 |
A |
667 |
639 |
3.97 |
|
|
|
| 20 |
A |
499 |
477 |
0.73 |
|
|
|
| 21 |
A |
356 |
341 |
0.73 |
|
|
|
| 22 |
A |
344 |
329 |
0.03 |
|
|
|
| 23 |
A |
176 |
168 |
2.07 |
|
|
|
| 24 |
A |
3127 |
2991 |
11.35 |
|
|
|
| 25 |
A |
3093 |
2959 |
19.44 |
|
|
|
| 26 |
A |
3057 |
2924 |
21.83 |
|
|
|
| 27 |
A |
3044 |
2913 |
45.93 |
|
|
|
| 28 |
A |
1497 |
1432 |
3.94 |
|
|
|
| 29 |
A |
1486 |
1422 |
0.84 |
|
|
|
| 30 |
A |
1404 |
1343 |
0.20 |
|
|
|
| 31 |
A |
1383 |
1323 |
0.22 |
|
|
|
| 32 |
A |
1316 |
1259 |
18.44 |
|
|
|
| 33 |
A |
1302 |
1245 |
3.26 |
|
|
|
| 34 |
A |
1180 |
1129 |
0.02 |
|
|
|
| 35 |
A |
1116 |
1067 |
0.65 |
|
|
|
| 36 |
A |
1103 |
1055 |
0.32 |
|
|
|
| 37 |
A |
954 |
913 |
0.32 |
|
|
|
| 38 |
A |
921 |
881 |
4.49 |
|
|
|
| 39 |
A |
805 |
770 |
0.13 |
|
|
|
| 40 |
A |
699 |
669 |
0.35 |
|
|
|
| 41 |
A |
400 |
382 |
0.25 |
|
|
|
| 42 |
A |
233 |
223 |
0.57 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31605.4 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 30236.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.095 |
|
|
-0.230 |
| 2 |
C |
-0.475 |
|
|
-0.255 |
| 3 |
C |
-0.475 |
|
|
-0.255 |
| 4 |
C |
-0.309 |
|
|
0.016 |
| 5 |
C |
-0.309 |
|
|
0.016 |
| 6 |
C |
-0.310 |
|
|
-0.056 |
| 7 |
H |
0.200 |
|
|
0.161 |
| 8 |
H |
0.188 |
|
|
0.125 |
| 9 |
H |
0.200 |
|
|
0.161 |
| 10 |
H |
0.188 |
|
|
0.125 |
| 11 |
H |
0.176 |
|
|
0.047 |
| 12 |
H |
0.166 |
|
|
0.014 |
| 13 |
H |
0.176 |
|
|
0.047 |
| 14 |
H |
0.166 |
|
|
0.014 |
| 15 |
H |
0.166 |
|
|
0.018 |
| 16 |
H |
0.156 |
|
|
0.053 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.314 |
1.454 |
0.000 |
1.960 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-1.371 |
1.401 |
0.000 |
1.960 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.537 |
1.352 |
0.000 |
| y |
1.352 |
-48.065 |
0.000 |
| z |
0.000 |
0.000 |
-41.415 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.798 |
1.352 |
0.000 |
| y |
1.352 |
-4.089 |
0.000 |
| z |
0.000 |
0.000 |
5.887 |
|
| Polar |
|---|
| 3z2-r2 | 11.773 |
|---|
| x2-y2 | 1.528 |
|---|
| xy | 1.352 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
197.039 |
| (<r2>)1/2 |
14.037 |