return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10S (2H-Thiopyran, tetrahydro-)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-594.334266
Energy at 298.15K-594.346621
HF Energy-594.077289
Nuclear repulsion energy311.149961
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3145 2985 23.63      
2 A' 3121 2962 58.44      
3 A' 3107 2949 39.88      
4 A' 3079 2922 20.97      
5 A' 3067 2911 22.29      
6 A' 3054 2899 19.34      
7 A' 1545 1466 1.47      
8 A' 1532 1454 9.51      
9 A' 1518 1440 7.36      
10 A' 1420 1348 1.68      
11 A' 1378 1308 8.12      
12 A' 1299 1233 1.23      
13 A' 1274 1209 6.96      
14 A' 1113 1056 2.67      
15 A' 1048 994 0.65      
16 A' 1002 951 8.34      
17 A' 856 813 3.85      
18 A' 836 793 1.21      
19 A' 670 636 3.75      
20 A' 510 484 0.51      
21 A' 364 345 0.53      
22 A' 349 331 0.09      
23 A' 180 171 1.97      
24 A" 3146 2986 11.67      
25 A" 3114 2956 19.50      
26 A" 3080 2924 21.80      
27 A" 3067 2911 43.41      
28 A" 1525 1448 2.31      
29 A" 1513 1436 0.58      
30 A" 1424 1351 0.36      
31 A" 1405 1333 0.58      
32 A" 1340 1272 20.93      
33 A" 1318 1251 2.38      
34 A" 1194 1134 0.03      
35 A" 1129 1072 0.69      
36 A" 1112 1056 0.09      
37 A" 962 913 0.05      
38 A" 934 887 3.91      
39 A" 819 777 0.13      
40 A" 703 667 0.58      
41 A" 403 383 0.24      
42 A" 239 227 0.49      

Unscaled Zero Point Vibrational Energy (zpe) 31945.2 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 30322.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.13244 0.09942 0.06326

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 -0.758 -1.312 0.000
C2 0.813 1.524 0.000
C3 0.142 1.010 1.280
C4 0.142 1.010 -1.280
C5 0.142 -0.517 1.375
C6 0.142 -0.517 -1.375
H7 0.806 2.620 0.000
H8 1.868 1.219 0.000
H9 1.170 -0.897 1.405
H10 1.170 -0.897 -1.405
H11 0.665 1.417 2.155
H12 0.665 1.417 -2.155
H13 -0.892 1.371 1.325
H14 -0.892 1.371 -1.325
H15 -0.358 -0.850 -2.288
H16 -0.358 -0.850 2.288

Atom - Atom Distances (Å)
  S1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
S13.24172.80012.80011.82551.82554.23063.64682.42082.42083.75673.75672.99452.99452.36792.3679
C23.24171.53401.53402.55082.55081.09551.09832.82172.82172.16242.16242.16422.16423.49933.4993
C32.80011.53402.56071.53013.06332.16092.15912.17013.45001.09703.49841.09572.82584.05532.1744
C42.80011.53402.56073.06331.53012.16092.15913.45002.17013.49841.09702.82581.09572.17444.0553
C51.82552.55081.53013.06332.75023.48832.80761.09592.98772.14974.05862.15273.45273.71211.0932
C61.82552.55083.06331.53012.75023.48832.80762.98771.09594.05862.14973.45272.15271.09323.7121
H74.23061.09552.16092.16093.48833.48831.75803.80423.80422.47162.47162.48902.48904.31634.3163
H83.64681.09832.15912.15912.80762.80761.75802.63412.63412.47552.47553.06493.06493.80453.8045
H92.42082.82172.17013.45001.09592.98773.80422.63412.80912.48424.27513.06554.10373.99651.7656
H102.42082.82173.45002.17012.98771.09593.80422.63412.80914.27512.48424.10373.06551.76563.9965
H113.75672.16241.09703.49842.14974.05862.47162.47552.48424.27514.30931.76493.81225.09172.4913
H123.75672.16243.49841.09704.05862.14972.47162.47554.27512.48424.30933.81221.76492.49135.0917
H132.99452.16421.09572.82582.15273.45272.48903.06493.06554.10371.76493.81222.64964.27432.4789
H142.99452.16422.82581.09573.45272.15272.48903.06494.10373.06553.81221.76492.64962.47894.2743
H152.36793.49934.05532.17443.71211.09324.31633.80453.99651.76565.09172.49134.27432.47894.5759
H162.36793.49932.17444.05531.09323.71214.31633.80451.76563.99652.49135.09172.47894.27434.5759

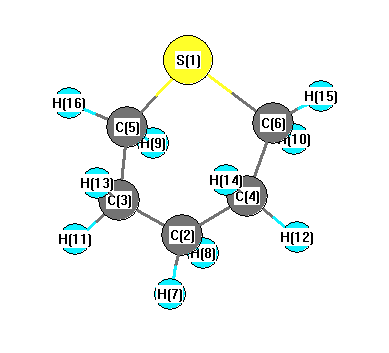
picture of 2H-Thiopyran, tetrahydro- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 C5 C3 112.819 S1 C5 H9 109.362
S1 C5 H16 105.692 S1 C6 C4 112.819
S1 C6 H10 109.362 S1 C6 H15 105.692
C2 C3 C5 112.703 C2 C3 H11 109.419
C2 C3 H13 109.646 C2 C4 C6 112.703
C2 C4 H12 109.419 C2 C4 H14 109.646
C3 C2 C4 113.156 C3 C2 H7 109.391
C3 C2 H8 109.087 C3 C5 H9 110.360
C3 C5 H16 110.864 C4 C2 H7 109.391
C4 C2 H8 109.087 C4 C6 H10 110.360
C4 C6 H15 110.864 C5 S1 C6 97.747
C5 C3 H11 108.700 C5 C3 H13 109.014
C6 C4 H12 108.700 C6 C4 H14 109.014
H7 C2 H8 106.512 H9 C5 H16 107.516
H10 C6 H15 107.516 H11 C3 H13 107.204
H12 C4 H14 107.204
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 S 0.090      
2 C -0.283      
3 C -0.277      
4 C -0.277      
5 C -0.440      
6 C -0.440      
7 H 0.151      
8 H 0.140      
9 H 0.173      
10 H 0.173      
11 H 0.151      
12 H 0.151      
13 H 0.161      
14 H 0.161      
15 H 0.184      
16 H 0.184      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 8.711 0.944 0.000
y 0.944 9.785 0.000
z 0.000 0.000 10.642


<r2> (average value of r2) Å2
<r2> 197.590
(<r2>)1/2 14.057