return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10 (Cyclopropane, 1,1-dimethyl-)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-195.846168
Energy at 298.15K-195.857038
HF Energy-195.131557
Nuclear repulsion energy184.202173
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3163 3012 10.98      
2 A 3133 2983 38.84      
3 A 3053 2907 32.23      
4 A 1574 1499 0.17      
5 A 1554 1480 5.28      
6 A 1476 1405 0.00      
7 A 1407 1340 0.83      
8 A 1123 1070 1.58      
9 A 1087 1036 6.79      
10 A 981 934 12.45      
11 A 708 674 1.61      
12 A 360 343 0.02      
13 A 3232 3078 0.00      
14 A 3117 2968 0.00      
15 A 1533 1460 0.00      
16 A 1220 1161 0.00      
17 A 1095 1043 0.00      
18 A 927 883 0.00      
19 A 339 323 0.00      
20 A 225 214 0.00      
21 A 3156 3005 20.57      
22 A 3120 2972 61.06      
23 A 1550 1477 5.43      
24 A 1514 1442 3.05      
25 A 1206 1148 4.94      
26 A 1109 1056 0.86      
27 A 892 849 6.18      
28 A 402 383 0.99      
29 A 251 239 0.01      
30 A 3245 3090 26.46      
31 A 3130 2981 16.96      
32 A 3048 2903 30.23      
33 A 1546 1472 4.27      
34 A 1466 1396 7.58      
35 A 1366 1301 0.46      
36 A 997 949 0.53      
37 A 980 933 0.42      
38 A 811 772 0.08      
39 A 368 351 0.74      

Unscaled Zero Point Vibrational Energy (zpe) 30730.1 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 29264.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
0.20499 0.17286 0.12681

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.150
C2 0.757 0.000 -1.154
C3 -0.757 0.000 -1.154
C4 0.000 1.274 0.971
C5 0.000 -1.274 0.971
H6 1.269 0.913 -1.453
H7 1.269 -0.913 -1.453
H8 -1.269 -0.913 -1.453
H9 -1.269 0.913 -1.453
H10 -0.889 1.325 1.616
H11 -0.889 -1.325 1.616
H12 0.889 -1.325 1.616
H13 0.889 1.325 1.616
H14 0.000 2.162 0.325
H15 0.000 -2.162 0.325

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15
C11.50771.50771.51601.51602.23902.23902.23902.23902.16642.16642.16642.16642.16882.1688
C21.50771.51322.59112.59111.08881.08882.24162.24163.48353.48353.07323.07322.72672.7267
C31.50771.51322.59112.59112.24162.24161.08881.08883.07323.07323.48353.48352.72672.7267
C41.51602.59112.59112.54852.76003.50323.50322.76001.09902.82132.82131.09901.09753.4961
C51.51602.59112.59112.54853.50322.76002.76003.50322.82131.09901.09902.82133.49611.0975
H62.23901.08882.24162.76003.50321.82653.12622.53713.77384.36813.81723.11962.51623.7720
H72.23901.08882.24163.50322.76001.82652.53713.12624.36813.77383.11963.81723.77202.5162
H82.23902.24161.08883.50322.76003.12622.53711.82653.81723.11963.77384.36813.77202.5162
H92.23902.24161.08882.76003.50322.53713.12621.82653.11963.81724.36813.77382.51623.7720
H102.16643.48353.07321.09902.82133.77384.36813.81723.11962.64933.19031.77751.77633.8222
H112.16643.48353.07322.82131.09904.36813.77383.11963.81722.64931.77753.19033.82221.7763
H122.16643.07323.48352.82131.09903.81723.11963.77384.36813.19031.77752.64933.82221.7763
H132.16643.07323.48351.09902.82133.11963.81724.36813.77381.77753.19032.64931.77633.8222
H142.16882.72672.72671.09753.49612.51623.77203.77202.51621.77633.82223.82221.77634.3234
H152.16882.72672.72673.49611.09753.77202.51622.51623.77203.82221.77631.77633.82224.3234

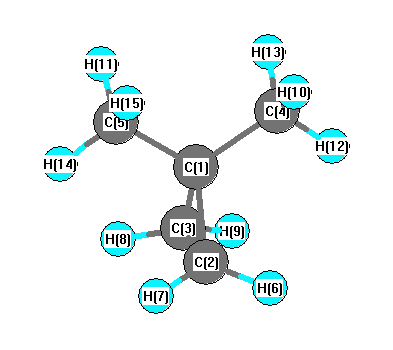
picture of Cyclopropane, 1,1-dimethyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 59.879 C1 C2 H6 118.259
C1 C2 H7 118.259 C1 C3 C2 59.879
C1 C3 H8 118.259 C1 C3 H9 118.259
C1 C4 H10 110.870 C1 C4 H13 110.870
C1 C4 H14 111.154 C1 C5 H11 110.870
C1 C5 H12 110.870 C1 C5 H15 111.154
C2 C1 C3 60.243 C2 C1 C4 117.944
C2 C1 C5 117.944 C2 C3 H8 118.046
C2 C3 H9 118.046 C3 C1 C4 117.944
C3 C1 C5 117.944 C3 C2 H6 118.046
C3 C2 H7 118.046 C4 C1 C5 114.392
H6 C2 H7 114.021 H8 C3 H9 114.021
H10 C4 H13 107.936 H10 C4 H14 107.938
H11 C5 H12 107.936 H11 C5 H15 107.938
H12 C5 H15 107.938 H13 C4 H14 107.938
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability