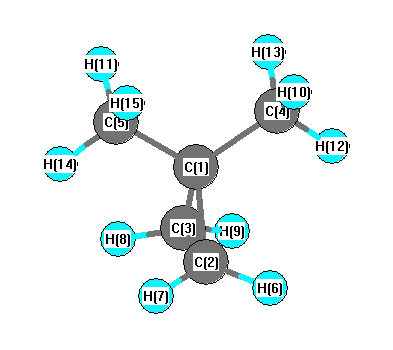
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3166 |
3011 |
11.65 |
|
|
|
| 2 |
A |
3140 |
2986 |
38.26 |
|
|
|
| 3 |
A |
3056 |
2906 |
38.41 |
|
|
|
| 4 |
A |
1552 |
1476 |
0.08 |
|
|
|
| 5 |
A |
1530 |
1455 |
6.21 |
|
|
|
| 6 |
A |
1449 |
1378 |
0.00 |
|
|
|
| 7 |
A |
1387 |
1319 |
1.34 |
|
|
|
| 8 |
A |
1116 |
1061 |
0.79 |
|
|
|
| 9 |
A |
1055 |
1003 |
7.76 |
|
|
|
| 10 |
A |
978 |
930 |
14.67 |
|
|
|
| 11 |
A |
710 |
675 |
1.56 |
|
|
|
| 12 |
A |
349 |
332 |
0.00 |
|
|
|
| 13 |
A |
3240 |
3081 |
0.00 |
|
|
|
| 14 |
A |
3122 |
2969 |
0.00 |
|
|
|
| 15 |
A |
1507 |
1433 |
0.00 |
|
|
|
| 16 |
A |
1201 |
1143 |
0.00 |
|
|
|
| 17 |
A |
1077 |
1025 |
0.00 |
|
|
|
| 18 |
A |
905 |
860 |
0.00 |
|
|
|
| 19 |
A |
331 |
315 |
0.00 |
|
|
|
| 20 |
A |
218 |
207 |
0.00 |
|
|
|
| 21 |
A |
3161 |
3006 |
23.16 |
|
|
|
| 22 |
A |
3125 |
2972 |
59.12 |
|
|
|
| 23 |
A |
1527 |
1452 |
6.30 |
|
|
|
| 24 |
A |
1490 |
1417 |
4.04 |
|
|
|
| 25 |
A |
1175 |
1118 |
6.93 |
|
|
|
| 26 |
A |
1081 |
1028 |
1.32 |
|
|
|
| 27 |
A |
886 |
842 |
5.13 |
|
|
|
| 28 |
A |
392 |
373 |
1.26 |
|
|
|
| 29 |
A |
245 |
233 |
0.02 |
|
|
|
| 30 |
A |
3253 |
3094 |
23.79 |
|
|
|
| 31 |
A |
3138 |
2984 |
16.75 |
|
|
|
| 32 |
A |
3051 |
2902 |
34.64 |
|
|
|
| 33 |
A |
1521 |
1447 |
4.72 |
|
|
|
| 34 |
A |
1439 |
1368 |
8.56 |
|
|
|
| 35 |
A |
1345 |
1279 |
0.84 |
|
|
|
| 36 |
A |
981 |
933 |
0.76 |
|
|
|
| 37 |
A |
969 |
921 |
0.49 |
|
|
|
| 38 |
A |
798 |
759 |
0.02 |
|
|
|
| 39 |
A |
357 |
340 |
0.75 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30509.4 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 29014.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.092 |
|
|
|
| 2 |
C |
-0.370 |
|
|
|
| 3 |
C |
-0.370 |
|
|
|
| 4 |
C |
-0.521 |
|
|
|
| 5 |
C |
-0.521 |
|
|
|
| 6 |
H |
0.173 |
|
|
|
| 7 |
H |
0.173 |
|
|
|
| 8 |
H |
0.173 |
|
|
|
| 9 |
H |
0.173 |
|
|
|
| 10 |
H |
0.168 |
|
|
|
| 11 |
H |
0.168 |
|
|
|
| 12 |
H |
0.168 |
|
|
|
| 13 |
H |
0.168 |
|
|
|
| 14 |
H |
0.163 |
|
|
|
| 15 |
H |
0.163 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.103 |
0.103 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.071 |
0.000 |
0.000 |
| y |
0.000 |
-31.770 |
0.000 |
| z |
0.000 |
0.000 |
-33.621 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.376 |
0.000 |
0.000 |
| y |
0.000 |
1.576 |
0.000 |
| z |
0.000 |
0.000 |
-1.200 |
|
| Polar |
|---|
| 3z2-r2 | -2.400 |
|---|
| x2-y2 | -1.301 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.186 |
0.000 |
0.000 |
| y |
0.000 |
7.538 |
0.000 |
| z |
0.000 |
0.000 |
7.903 |
<r2> (average value of r
2) Å
2
| <r2> |
122.213 |
| (<r2>)1/2 |
11.055 |