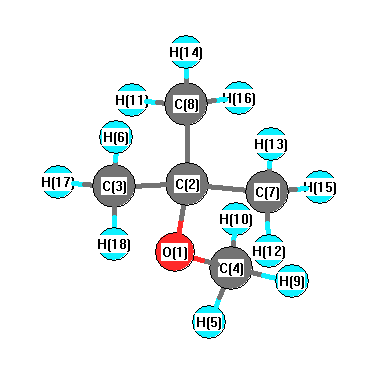
Vibrational Frequencies calculated at PBE1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3168 |
3011 |
46.27 |
|
|
|
| 2 |
A |
3160 |
3003 |
10.72 |
|
|
|
| 3 |
A |
3157 |
3000 |
27.89 |
|
|
|
| 4 |
A |
3153 |
2996 |
49.22 |
|
|
|
| 5 |
A |
3082 |
2928 |
4.39 |
|
|
|
| 6 |
A |
3075 |
2922 |
23.20 |
|
|
|
| 7 |
A |
3028 |
2877 |
51.04 |
|
|
|
| 8 |
A |
1551 |
1474 |
11.96 |
|
|
|
| 9 |
A |
1542 |
1465 |
7.25 |
|
|
|
| 10 |
A |
1528 |
1452 |
2.73 |
|
|
|
| 11 |
A |
1513 |
1438 |
0.67 |
|
|
|
| 12 |
A |
1496 |
1421 |
0.33 |
|
|
|
| 13 |
A |
1444 |
1372 |
13.90 |
|
|
|
| 14 |
A |
1422 |
1351 |
19.66 |
|
|
|
| 15 |
A |
1309 |
1244 |
13.63 |
|
|
|
| 16 |
A |
1264 |
1201 |
127.37 |
|
|
|
| 17 |
A |
1221 |
1160 |
2.82 |
|
|
|
| 18 |
A |
1169 |
1111 |
80.86 |
|
|
|
| 19 |
A |
1055 |
1003 |
1.28 |
|
|
|
| 20 |
A |
940 |
894 |
0.26 |
|
|
|
| 21 |
A |
893 |
849 |
15.60 |
|
|
|
| 22 |
A |
749 |
711 |
4.31 |
|
|
|
| 23 |
A |
512 |
486 |
1.70 |
|
|
|
| 24 |
A |
413 |
392 |
0.65 |
|
|
|
| 25 |
A |
366 |
348 |
0.90 |
|
|
|
| 26 |
A |
292 |
278 |
0.56 |
|
|
|
| 27 |
A |
257 |
244 |
0.35 |
|
|
|
| 28 |
A |
3166 |
3009 |
29.04 |
|
|
|
| 29 |
A |
3159 |
3002 |
4.42 |
|
|
|
| 30 |
A |
3150 |
2994 |
0.12 |
|
|
|
| 31 |
A |
3085 |
2932 |
75.04 |
|
|
|
| 32 |
A |
3071 |
2918 |
3.67 |
|
|
|
| 33 |
A |
1535 |
1459 |
0.56 |
|
|
|
| 34 |
A |
1518 |
1443 |
5.11 |
|
|
|
| 35 |
A |
1509 |
1434 |
0.00 |
|
|
|
| 36 |
A |
1495 |
1421 |
0.00 |
|
|
|
| 37 |
A |
1417 |
1347 |
21.84 |
|
|
|
| 38 |
A |
1280 |
1216 |
20.83 |
|
|
|
| 39 |
A |
1190 |
1131 |
1.16 |
|
|
|
| 40 |
A |
1049 |
997 |
1.86 |
|
|
|
| 41 |
A |
962 |
914 |
0.03 |
|
|
|
| 42 |
A |
929 |
882 |
0.03 |
|
|
|
| 43 |
A |
461 |
438 |
5.73 |
|
|
|
| 44 |
A |
343 |
326 |
1.22 |
|
|
|
| 45 |
A |
290 |
276 |
1.55 |
|
|
|
| 46 |
A |
238 |
226 |
1.25 |
|
|
|
| 47 |
A |
181 |
172 |
1.17 |
|
|
|
| 48 |
A |
41 |
39 |
3.08 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36412.4 cm
-1
Scaled (by 0.9503) Zero Point Vibrational Energy (zpe) 34602.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.479 |
|
|
|
| 2 |
C |
0.272 |
|
|
|
| 3 |
C |
-0.512 |
|
|
|
| 4 |
C |
-0.267 |
|
|
|
| 5 |
H |
0.182 |
|
|
|
| 6 |
H |
0.162 |
|
|
|
| 7 |
C |
-0.531 |
|
|
|
| 8 |
C |
-0.531 |
|
|
|
| 9 |
H |
0.152 |
|
|
|
| 10 |
H |
0.152 |
|
|
|
| 11 |
H |
0.181 |
|
|
|
| 12 |
H |
0.181 |
|
|
|
| 13 |
H |
0.171 |
|
|
|
| 14 |
H |
0.171 |
|
|
|
| 15 |
H |
0.169 |
|
|
|
| 16 |
H |
0.169 |
|
|
|
| 17 |
H |
0.180 |
|
|
|
| 18 |
H |
0.180 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.089 |
0.461 |
0.000 |
1.182 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.737 |
-1.890 |
0.000 |
| y |
-1.890 |
-37.709 |
0.000 |
| z |
0.000 |
0.000 |
-39.427 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.169 |
-1.890 |
0.000 |
| y |
-1.890 |
1.873 |
0.000 |
| z |
0.000 |
0.000 |
-0.704 |
|
| Polar |
|---|
| 3z2-r2 | -1.408 |
|---|
| x2-y2 | -2.028 |
|---|
| xy | -1.890 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.204 |
-0.436 |
0.000 |
| y |
-0.436 |
9.093 |
0.000 |
| z |
0.000 |
0.000 |
8.151 |
<r2> (average value of r
2) Å
2
| <r2> |
178.649 |
| (<r2>)1/2 |
13.366 |