return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C2H4N4 (2H-Tetrazole, 2-methyl-)

using model chemistry: MP3/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP3/6-31G*
 hartrees
Energy at 0K-296.713967
Energy at 298.15K-296.721580
HF Energy-295.791488
Nuclear repulsion energy231.999226
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3358 3152 0.21      
2 A 3256 3056 0.45      
3 A 3229 3031 4.72      
4 A 3143 2950 14.12      
5 A 1575 1479 3.50      
6 A 1548 1453 10.77      
7 A 1535 1440 10.92      
8 A 1503 1410 0.89      
9 A 1489 1398 19.70      
10 A 1413 1326 10.10      
11 A 1368 1284 20.77      
12 A 1293 1214 13.95      
13 A 1207 1132 10.93      
14 A 1186 1113 0.02      
15 A 1103 1035 18.46      
16 A 1085 1018 16.84      
17 A 1052 987 4.84      
18 A 899 844 12.50      
19 A 741 695 16.45      
20 A 729 685 5.73      
21 A 698 655 6.23      
22 A 381 358 5.32      
23 A 240 226 5.81      
24 A 69 65 0.01      

Unscaled Zero Point Vibrational Energy (zpe) 17049.3 cm-1
Scaled (by 0.9386) Zero Point Vibrational Energy (zpe) 16002.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP3/6-31G*
ABC
0.32747 0.13121 0.09538

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP3/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 2.306 -1.199 0.000
C2 1.410 -0.598 0.000
N3 0.185 -1.117 0.000
N4 1.410 0.752 -0.000
N5 0.145 1.108 0.000
N6 -0.553 -0.015 -0.000
H7 -2.358 -0.555 -0.891
H8 -2.338 1.002 -0.004
H9 -2.358 -0.548 0.895
C10 -2.004 -0.035 0.000

Atom - Atom Distances (Å)
  H1 C2 N3 N4 N5 N6 H7 H8 H9 C10
H11.07862.12292.14643.16163.09454.79255.13914.79424.4653
C21.07861.33031.35012.12492.04793.87284.07543.87373.4610
N32.12291.33032.23422.22551.32572.75293.29442.75562.4423
N42.14641.35012.23421.31482.10764.08683.75634.08553.5040
N53.16162.12492.22551.31481.32273.13452.48473.13192.4342
N63.09452.04791.32572.10761.32272.08412.05442.08441.4517
H74.79253.87282.75294.08683.13452.08411.79161.78591.0906
H85.13914.07543.29443.75632.48472.05441.79161.79161.0887
H94.79423.87372.75564.08553.13192.08441.78591.79161.0906
C104.46533.46102.44233.50402.43421.45171.09061.08871.0906

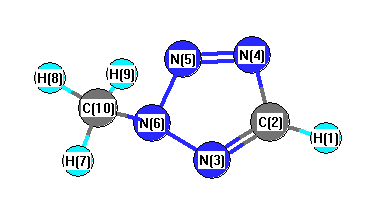
picture of 2H-Tetrazole, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C2 N3 123.257 H1 C2 N4 123.820
C2 N3 N6 100.896 C2 N4 N5 105.750
N3 C2 N4 112.923 N3 N6 N5 114.348
N3 N6 C10 123.065 N4 N5 N6 106.083
N5 N6 C10 122.587 N6 C10 H7 109.303
N6 C10 H8 107.067 N6 C10 H9 109.320
H7 C10 H8 110.591 H7 C10 H9 109.918
H8 C10 H9 110.583
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability