return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10S (Cyclopentanethiol)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-594.323287
Energy at 298.15K-594.334876
HF Energy-594.067272
Nuclear repulsion energy300.900015
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3158 2998 70.26      
2 A 3146 2987 17.08      
3 A 3141 2981 5.25      
4 A 3129 2970 19.65      
5 A 3105 2947 37.56      
6 A 3096 2939 31.38      
7 A 3091 2934 27.85      
8 A 3088 2931 7.83      
9 A 3083 2926 9.60      
10 A 2725 2587 23.30      
11 A 1570 1490 0.35      
12 A 1547 1469 4.55      
13 A 1544 1466 2.10      
14 A 1534 1456 0.41      
15 A 1395 1324 5.04      
16 A 1384 1313 0.64      
17 A 1368 1299 0.31      
18 A 1360 1291 2.16      
19 A 1326 1259 0.79      
20 A 1314 1247 18.40      
21 A 1288 1223 1.54      
22 A 1261 1197 6.45      
23 A 1229 1167 0.71      
24 A 1202 1141 4.91      
25 A 1138 1080 0.75      
26 A 1076 1021 1.24      
27 A 1011 960 0.35      
28 A 1000 949 0.78      
29 A 979 929 1.35      
30 A 953 904 1.42      
31 A 908 862 1.55      
32 A 890 845 4.71      
33 A 838 796 4.51      
34 A 831 789 2.51      
35 A 760 722 1.90      
36 A 632 600 0.53      
37 A 469 446 0.37      
38 A 373 354 1.38      
39 A 252 240 4.31      
40 A 202 191 19.52      
41 A 167 159 0.78      
42 A 43 40 0.04      

Unscaled Zero Point Vibrational Energy (zpe) 31302.9 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 29712.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.21049 0.06081 0.05042

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 -2.393 0.997 -0.891
H2 -2.281 1.252 0.876
C3 -1.829 0.768 0.006
H4 -2.586 -1.294 -0.310
H5 -1.783 -0.995 1.262
C6 -1.774 -0.768 0.190
H7 -0.333 -1.008 -1.405
H8 -0.086 -2.141 -0.035
C9 -0.386 -1.133 -0.325
H10 -0.059 2.139 0.228
H11 -0.161 1.192 -1.278
C12 -0.344 1.175 -0.201
H13 0.292 -0.089 1.425
C14 0.432 -0.025 0.348
H15 2.350 0.804 0.492
S16 1.903 -0.060 -0.029

Atom - Atom Distances (Å)
  H1 H2 C3 H4 H5 C6 H7 H8 C9 H10 H11 C12 H13 C14 H15 S16
H11.78941.08402.37142.99562.15962.92033.98682.98012.82862.27332.16893.70833.24904.94434.5069
H21.78941.09382.82582.33372.19253.75644.14213.27422.47783.02312.21722.95273.04424.66854.4772
C31.08401.09382.21942.16511.54752.71753.39072.40902.24922.14721.55322.69192.41974.20743.8227
H42.37142.82582.21941.79011.08912.52202.65352.20634.29633.60533.33693.57073.33905.42354.6643
H52.99562.33372.16511.79011.09573.03622.42372.11903.72303.72382.98662.27102.58484.57354.0165
C62.15962.19251.54751.08911.09572.16392.18761.52543.37482.93232.44382.50162.33294.42433.7515
H72.92033.75642.71752.52203.03622.16391.79451.08863.55592.21012.49333.04082.15013.75232.7910
H83.98684.14213.39072.65352.42372.18761.79451.09094.28743.55743.32972.54692.21113.85812.8788
C92.98013.27422.40902.20632.11901.52541.08861.09093.33362.52212.31122.14771.53193.45022.5451
H102.82862.47782.24924.29633.72303.37483.55594.28743.33361.78241.09262.55272.22222.76732.9582
H112.27333.02312.14723.60533.72382.93232.21013.55742.52211.78241.09313.02532.11603.09702.7178
C122.16892.21721.55323.33692.98662.44382.49333.32972.31121.09261.09312.15521.53102.80682.5699
H133.70832.95272.69193.57072.27102.50163.04082.54692.14772.55273.02532.15521.08782.42952.1704
C143.24903.04422.41973.33902.58482.33292.15012.21111.53192.22222.11601.53101.08782.09531.5196
H154.94434.66854.20745.42354.57354.42433.75233.85813.45022.76733.09702.80682.42952.09531.1035
S164.50694.47723.82274.66434.01653.75152.79102.87882.54512.95822.71782.56992.17041.51961.1035

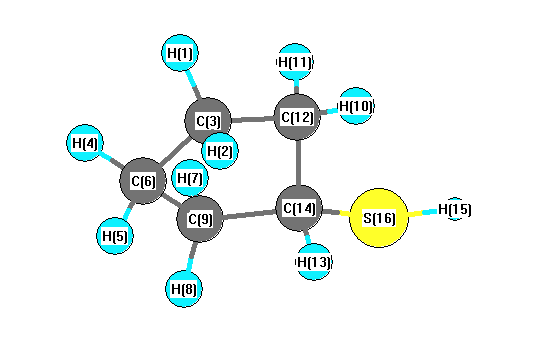
picture of Cyclopentanethiol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C3 H2 110.505 H1 C3 C6 109.035
H1 C3 C12 109.372 H2 C3 C6 111.046
H2 C3 C12 112.628 C3 C6 H4 113.513
C3 C6 H5 108.791 C3 C6 C9 103.245
C3 C12 H10 115.343 C3 C12 H11 107.180
C3 C12 C14 103.361 H4 C6 H5 110.041
H4 C6 C9 114.061 H5 C6 C9 106.745
C6 C3 C12 104.026 C6 C9 H7 110.635
C6 C9 H8 112.413 C6 C9 C14 99.468
H7 C9 H8 110.841 H7 C9 C14 109.097
H8 C9 C14 113.866 C9 C14 C12 97.979
C9 C14 H13 108.953 C9 C14 S16 113.033
H10 C12 H11 109.266 H10 C12 C14 114.751
H11 C12 C14 106.284 C12 C14 H13 109.609
C12 C14 S16 114.798 H13 C14 S16 111.618
C14 S16 H15 104.923
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 H 0.157      
2 H 0.152      
3 C -0.301      
4 H 0.157      
5 H 0.155      
6 C -0.299      
7 H 0.165      
8 H 0.166      
9 C -0.300      
10 H 0.151      
11 H 0.166      
12 C -0.282      
13 H 0.178      
14 C -0.279      
15 H 0.087      
16 S -0.073      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 11.645 0.210 -0.069
y 0.210 9.212 0.215
z -0.069 0.215 8.079


<r2> (average value of r2) Å2
<r2> 231.082
(<r2>)1/2 15.201