Jump to
S1C2
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -246.414177 |
| Energy at 298.15K | -246.419857 |
| HF Energy | -245.791927 |
| Nuclear repulsion energy | 157.781460 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3287 |
3037 |
19.12 |
|
|
|
| 2 |
A' |
3138 |
2899 |
42.99 |
|
|
|
| 3 |
A' |
3130 |
2892 |
19.75 |
|
|
|
| 4 |
A' |
2512 |
2320 |
0.31 |
|
|
|
| 5 |
A' |
1615 |
1491 |
2.10 |
|
|
|
| 6 |
A' |
1597 |
1475 |
7.74 |
|
|
|
| 7 |
A' |
1573 |
1453 |
0.20 |
|
|
|
| 8 |
A' |
1504 |
1389 |
52.71 |
|
|
|
| 9 |
A' |
1314 |
1214 |
89.63 |
|
|
|
| 10 |
A' |
1245 |
1150 |
102.07 |
|
|
|
| 11 |
A' |
1070 |
988 |
36.47 |
|
|
|
| 12 |
A' |
1004 |
927 |
11.16 |
|
|
|
| 13 |
A' |
572 |
528 |
1.84 |
|
|
|
| 14 |
A' |
397 |
367 |
2.93 |
|
|
|
| 15 |
A' |
189 |
174 |
2.71 |
|
|
|
| 16 |
A" |
3198 |
2954 |
48.28 |
|
|
|
| 17 |
A" |
3173 |
2931 |
28.58 |
|
|
|
| 18 |
A" |
1589 |
1467 |
7.68 |
|
|
|
| 19 |
A" |
1337 |
1235 |
3.65 |
|
|
|
| 20 |
A" |
1248 |
1153 |
4.41 |
|
|
|
| 21 |
A" |
1100 |
1016 |
2.49 |
|
|
|
| 22 |
A" |
390 |
360 |
2.90 |
|
|
|
| 23 |
A" |
232 |
215 |
5.51 |
|
|
|
| 24 |
A" |
90 |
83 |
0.13 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18251.5 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 16858.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.123 |
-0.355 |
0.000 |
| O2 |
-0.734 |
-0.592 |
0.000 |
| C3 |
0.000 |
0.595 |
0.000 |
| C4 |
1.432 |
0.261 |
0.000 |
| N5 |
2.559 |
0.036 |
0.000 |
| H6 |
-2.603 |
-1.327 |
0.000 |
| H7 |
-2.430 |
0.200 |
0.889 |
| H8 |
-2.430 |
0.200 |
-0.889 |
| H9 |
-0.214 |
1.203 |
-0.884 |
| H10 |
-0.214 |
1.203 |
0.884 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4088 | 2.3261 | 3.6078 | 4.6986 | 1.0842 | 1.0923 | 1.0923 | 2.6180 | 2.6180 |
O2 | 1.4088 | | 1.3960 | 2.3279 | 3.3530 | 2.0085 | 2.0723 | 2.0723 | 2.0673 | 2.0673 | C3 | 2.3261 | 1.3960 | | 1.4703 | 2.6195 | 3.2363 | 2.6181 | 2.6181 | 1.0936 | 1.0936 | C4 | 3.6078 | 2.3279 | 1.4703 | | 1.1496 | 4.3364 | 3.9637 | 3.9637 | 2.0920 | 2.0920 | N5 | 4.6986 | 3.3530 | 2.6195 | 1.1496 | | 5.3397 | 5.0709 | 5.0709 | 3.1355 | 3.1355 | H6 | 1.0842 | 2.0085 | 3.2363 | 4.3364 | 5.3397 | | 1.7754 | 1.7754 | 3.5906 | 3.5906 | H7 | 1.0923 | 2.0723 | 2.6181 | 3.9637 | 5.0709 | 1.7754 | | 1.7786 | 3.0105 | 2.4331 | H8 | 1.0923 | 2.0723 | 2.6181 | 3.9637 | 5.0709 | 1.7754 | 1.7786 | | 2.4331 | 3.0105 | H9 | 2.6180 | 2.0673 | 1.0936 | 2.0920 | 3.1355 | 3.5906 | 3.0105 | 2.4331 | | 1.7672 | H10 | 2.6180 | 2.0673 | 1.0936 | 2.0920 | 3.1355 | 3.5906 | 2.4331 | 3.0105 | 1.7672 | |
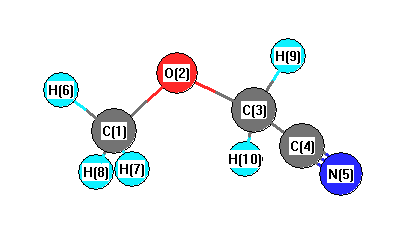 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
112.059 |
|
O2 |
C1 |
H6 |
106.622 |
| O2 |
C1 |
H7 |
111.270 |
|
O2 |
C1 |
H8 |
111.270 |
| O2 |
C3 |
C4 |
108.590 |
|
O2 |
C3 |
H9 |
111.703 |
| O2 |
C3 |
H10 |
111.703 |
|
C3 |
C4 |
N5 |
178.107 |
| C4 |
C3 |
H9 |
108.474 |
|
C4 |
C3 |
H10 |
108.474 |
| H6 |
C1 |
H7 |
109.311 |
|
H6 |
C1 |
H8 |
109.311 |
| H7 |
C1 |
H8 |
109.004 |
|
H9 |
C3 |
H10 |
107.800 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -246.416582 |
| Energy at 298.15K | -246.422382 |
| HF Energy | -245.793890 |
| Nuclear repulsion energy | 160.575941 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3288 |
3037 |
20.06 |
|
|
|
| 2 |
A |
3253 |
3004 |
6.08 |
|
|
|
| 3 |
A |
3215 |
2970 |
37.13 |
|
|
|
| 4 |
A |
3155 |
2914 |
26.78 |
|
|
|
| 5 |
A |
3143 |
2903 |
31.47 |
|
|
|
| 6 |
A |
2497 |
2306 |
0.23 |
|
|
|
| 7 |
A |
1605 |
1482 |
9.65 |
|
|
|
| 8 |
A |
1591 |
1470 |
6.99 |
|
|
|
| 9 |
A |
1586 |
1465 |
2.01 |
|
|
|
| 10 |
A |
1571 |
1451 |
0.32 |
|
|
|
| 11 |
A |
1476 |
1364 |
22.47 |
|
|
|
| 12 |
A |
1388 |
1282 |
10.48 |
|
|
|
| 13 |
A |
1306 |
1206 |
95.86 |
|
|
|
| 14 |
A |
1254 |
1158 |
25.97 |
|
|
|
| 15 |
A |
1241 |
1146 |
49.53 |
|
|
|
| 16 |
A |
1100 |
1016 |
8.07 |
|
|
|
| 17 |
A |
1015 |
937 |
34.65 |
|
|
|
| 18 |
A |
950 |
878 |
26.77 |
|
|
|
| 19 |
A |
628 |
580 |
2.03 |
|
|
|
| 20 |
A |
414 |
382 |
3.01 |
|
|
|
| 21 |
A |
381 |
352 |
1.96 |
|
|
|
| 22 |
A |
270 |
250 |
13.89 |
|
|
|
| 23 |
A |
184 |
170 |
4.09 |
|
|
|
| 24 |
A |
121 |
112 |
9.73 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18315.1 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 16917.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.570 |
-0.762 |
0.136 |
| O2 |
1.068 |
0.428 |
-0.437 |
| C3 |
-0.061 |
0.926 |
0.208 |
| C4 |
-1.244 |
0.050 |
0.035 |
| N5 |
-2.158 |
-0.637 |
-0.093 |
| H6 |
2.468 |
-1.015 |
-0.416 |
| H7 |
0.851 |
-1.578 |
0.051 |
| H8 |
1.821 |
-0.612 |
1.188 |
| H9 |
-0.281 |
1.895 |
-0.229 |
| H10 |
0.108 |
1.060 |
1.280 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4128 | 2.3479 | 2.9300 | 3.7364 | 1.0843 | 1.0904 | 1.0921 | 3.2589 | 2.6006 |
O2 | 1.4128 | | 1.3929 | 2.3898 | 3.4144 | 2.0101 | 2.0757 | 2.0715 | 2.0046 | 2.0663 | C3 | 2.3479 | 1.3929 | | 1.4814 | 2.6316 | 3.2482 | 2.6691 | 2.6209 | 1.0862 | 1.0929 | C4 | 2.9300 | 2.3898 | 1.4814 | | 1.1503 | 3.8875 | 2.6532 | 3.3412 | 2.0977 | 2.0965 | N5 | 3.7364 | 3.4144 | 2.6316 | 1.1503 | | 4.6521 | 3.1560 | 4.1802 | 3.1542 | 3.1453 | H6 | 1.0843 | 2.0101 | 3.2482 | 3.8875 | 4.6521 | | 1.7744 | 1.7760 | 4.0077 | 3.5706 | H7 | 1.0904 | 2.0757 | 2.6691 | 2.6532 | 3.1560 | 1.7744 | | 1.7796 | 3.6639 | 3.0032 | H8 | 1.0921 | 2.0715 | 2.6209 | 3.3412 | 4.1802 | 1.7760 | 1.7796 | | 3.5661 | 2.3958 | H9 | 3.2589 | 2.0046 | 1.0862 | 2.0977 | 3.1542 | 4.0077 | 3.6639 | 3.5661 | | 1.7677 | H10 | 2.6006 | 2.0663 | 1.0929 | 2.0965 | 3.1453 | 3.5706 | 3.0032 | 2.3958 | 1.7677 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
113.616 |
|
O2 |
C1 |
H6 |
106.475 |
| O2 |
C1 |
H7 |
111.385 |
|
O2 |
C1 |
H8 |
110.932 |
| O2 |
C3 |
C4 |
112.454 |
|
O2 |
C3 |
H9 |
107.272 |
| O2 |
C3 |
H10 |
111.884 |
|
C3 |
C4 |
N5 |
179.466 |
| C4 |
C3 |
H9 |
108.586 |
|
C4 |
C3 |
H10 |
108.102 |
| H6 |
C1 |
H7 |
109.355 |
|
H6 |
C1 |
H8 |
109.381 |
| H7 |
C1 |
H8 |
109.255 |
|
H9 |
C3 |
H10 |
108.426 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
