Jump to
S1C2
Energy calculated at B3PW91/6-31G*
| | hartrees |
|---|
| Energy at 0K | -247.158663 |
| Energy at 298.15K | -247.164235 |
| HF Energy | -247.158663 |
| Nuclear repulsion energy | 157.140465 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3171 |
3034 |
17.72 |
|
|
|
| 2 |
A' |
3008 |
2878 |
54.60 |
|
|
|
| 3 |
A' |
2990 |
2861 |
25.33 |
|
|
|
| 4 |
A' |
2391 |
2287 |
0.46 |
|
|
|
| 5 |
A' |
1540 |
1473 |
4.27 |
|
|
|
| 6 |
A' |
1517 |
1452 |
8.94 |
|
|
|
| 7 |
A' |
1497 |
1432 |
0.76 |
|
|
|
| 8 |
A' |
1426 |
1364 |
41.64 |
|
|
|
| 9 |
A' |
1237 |
1184 |
61.92 |
|
|
|
| 10 |
A' |
1184 |
1133 |
118.57 |
|
|
|
| 11 |
A' |
1017 |
973 |
33.69 |
|
|
|
| 12 |
A' |
962 |
920 |
6.72 |
|
|
|
| 13 |
A' |
551 |
527 |
1.53 |
|
|
|
| 14 |
A' |
379 |
363 |
2.15 |
|
|
|
| 15 |
A' |
181 |
174 |
2.30 |
|
|
|
| 16 |
A" |
3063 |
2930 |
52.40 |
|
|
|
| 17 |
A" |
3019 |
2889 |
35.71 |
|
|
|
| 18 |
A" |
1512 |
1447 |
8.71 |
|
|
|
| 19 |
A" |
1271 |
1216 |
3.51 |
|
|
|
| 20 |
A" |
1191 |
1139 |
2.69 |
|
|
|
| 21 |
A" |
1040 |
995 |
1.33 |
|
|
|
| 22 |
A" |
382 |
365 |
1.79 |
|
|
|
| 23 |
A" |
234 |
224 |
4.55 |
|
|
|
| 24 |
A" |
89 |
85 |
0.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17426.0 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 16671.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.127 |
-0.358 |
0.000 |
| O2 |
-0.738 |
-0.607 |
0.000 |
| C3 |
0.000 |
0.585 |
0.000 |
| C4 |
1.432 |
0.270 |
0.000 |
| N5 |
2.572 |
0.056 |
0.000 |
| H6 |
-2.619 |
-1.332 |
0.000 |
| H7 |
-2.439 |
0.203 |
0.895 |
| H8 |
-2.439 |
0.203 |
-0.895 |
| H9 |
-0.220 |
1.203 |
-0.888 |
| H10 |
-0.220 |
1.203 |
0.888 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4112 | 2.3263 | 3.6141 | 4.7168 | 1.0917 | 1.1013 | 1.1013 | 2.6187 | 2.6187 |
O2 | 1.4112 | | 1.4018 | 2.3408 | 3.3753 | 2.0162 | 2.0861 | 2.0861 | 2.0811 | 2.0811 | C3 | 2.3263 | 1.4018 | | 1.4666 | 2.6259 | 3.2457 | 2.6258 | 2.6258 | 1.1038 | 1.1038 | C4 | 3.6141 | 2.3408 | 1.4666 | | 1.1596 | 4.3568 | 3.9741 | 3.9741 | 2.0952 | 2.0952 | N5 | 4.7168 | 3.3753 | 2.6259 | 1.1596 | | 5.3732 | 5.0924 | 5.0924 | 3.1468 | 3.1468 | H6 | 1.0917 | 2.0162 | 3.2457 | 4.3568 | 5.3732 | | 1.7863 | 1.7863 | 3.6010 | 3.6010 | H7 | 1.1013 | 2.0861 | 2.6258 | 3.9741 | 5.0924 | 1.7863 | | 1.7892 | 3.0163 | 2.4333 | H8 | 1.1013 | 2.0861 | 2.6258 | 3.9741 | 5.0924 | 1.7863 | 1.7892 | | 2.4333 | 3.0163 | H9 | 2.6187 | 2.0811 | 1.1038 | 2.0952 | 3.1468 | 3.6010 | 3.0163 | 2.4333 | | 1.7756 | H10 | 2.6187 | 2.0811 | 1.1038 | 2.0952 | 3.1468 | 3.6010 | 2.4333 | 3.0163 | 1.7756 | |
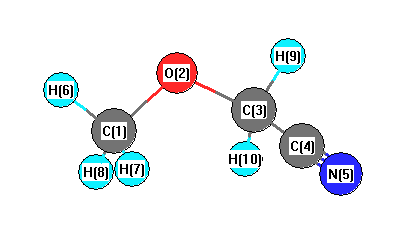 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
111.581 |
|
O2 |
C1 |
H6 |
106.629 |
| O2 |
C1 |
H7 |
111.665 |
|
O2 |
C1 |
H8 |
111.665 |
| O2 |
C3 |
C4 |
109.362 |
|
O2 |
C3 |
H9 |
111.767 |
| O2 |
C3 |
H10 |
111.767 |
|
C3 |
C4 |
N5 |
178.292 |
| C4 |
C3 |
H9 |
108.369 |
|
C4 |
C3 |
H10 |
108.369 |
| H6 |
C1 |
H7 |
109.084 |
|
H6 |
C1 |
H8 |
109.084 |
| H7 |
C1 |
H8 |
108.646 |
|
H9 |
C3 |
H10 |
107.082 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.264 |
|
|
|
| 2 |
O |
-0.445 |
|
|
|
| 3 |
C |
-0.108 |
|
|
|
| 4 |
C |
0.373 |
|
|
|
| 5 |
N |
-0.476 |
|
|
|
| 6 |
H |
0.201 |
|
|
|
| 7 |
H |
0.161 |
|
|
|
| 8 |
H |
0.161 |
|
|
|
| 9 |
H |
0.198 |
|
|
|
| 10 |
H |
0.198 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-4.318 |
1.759 |
0.000 |
4.662 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.497 |
0.694 |
0.000 |
| y |
0.694 |
-28.772 |
0.000 |
| z |
0.000 |
0.000 |
-28.884 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.669 |
0.694 |
0.000 |
| y |
0.694 |
3.419 |
0.000 |
| z |
0.000 |
0.000 |
3.250 |
|
| Polar |
|---|
| 3z2-r2 | 6.500 |
|---|
| x2-y2 | -6.726 |
|---|
| xy | 0.694 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.953 |
-0.007 |
0.000 |
| y |
-0.007 |
4.604 |
0.000 |
| z |
0.000 |
0.000 |
4.341 |
<r2> (average value of r
2) Å
2
| <r2> |
142.512 |
| (<r2>)1/2 |
11.938 |
Jump to
S1C1
Energy calculated at B3PW91/6-31G*
| | hartrees |
|---|
| Energy at 0K | -247.161212 |
| Energy at 298.15K | -247.166863 |
| HF Energy | -247.161212 |
| Nuclear repulsion energy | 159.882476 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3173 |
3036 |
18.48 |
|
|
|
| 2 |
A |
3125 |
2990 |
5.41 |
|
|
|
| 3 |
A |
3086 |
2953 |
37.94 |
|
|
|
| 4 |
A |
3020 |
2889 |
44.97 |
|
|
|
| 5 |
A |
3012 |
2881 |
28.48 |
|
|
|
| 6 |
A |
2373 |
2270 |
0.12 |
|
|
|
| 7 |
A |
1532 |
1465 |
10.27 |
|
|
|
| 8 |
A |
1514 |
1448 |
7.58 |
|
|
|
| 9 |
A |
1503 |
1438 |
2.68 |
|
|
|
| 10 |
A |
1489 |
1425 |
2.56 |
|
|
|
| 11 |
A |
1403 |
1342 |
16.20 |
|
|
|
| 12 |
A |
1323 |
1265 |
8.48 |
|
|
|
| 13 |
A |
1230 |
1177 |
54.99 |
|
|
|
| 14 |
A |
1192 |
1140 |
15.70 |
|
|
|
| 15 |
A |
1178 |
1127 |
84.48 |
|
|
|
| 16 |
A |
1039 |
994 |
7.29 |
|
|
|
| 17 |
A |
965 |
923 |
30.97 |
|
|
|
| 18 |
A |
909 |
870 |
19.28 |
|
|
|
| 19 |
A |
602 |
576 |
2.04 |
|
|
|
| 20 |
A |
392 |
375 |
2.41 |
|
|
|
| 21 |
A |
366 |
350 |
1.53 |
|
|
|
| 22 |
A |
255 |
244 |
10.55 |
|
|
|
| 23 |
A |
176 |
168 |
3.62 |
|
|
|
| 24 |
A |
118 |
113 |
8.81 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17487.3 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 16730.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.574 |
-0.763 |
0.136 |
| O2 |
1.076 |
0.431 |
-0.439 |
| C3 |
-0.063 |
0.923 |
0.208 |
| C4 |
-1.248 |
0.056 |
0.038 |
| N5 |
-2.167 |
-0.640 |
-0.096 |
| H6 |
2.476 |
-1.022 |
-0.423 |
| H7 |
0.850 |
-1.586 |
0.055 |
| H8 |
1.834 |
-0.618 |
1.196 |
| H9 |
-0.281 |
1.903 |
-0.229 |
| H10 |
0.105 |
1.063 |
1.289 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4160 | 2.3514 | 2.9411 | 3.7508 | 1.0917 | 1.0988 | 1.1007 | 3.2682 | 2.6119 |
O2 | 1.4160 | | 1.3991 | 2.4023 | 3.4325 | 2.0177 | 2.0886 | 2.0851 | 2.0124 | 2.0802 | C3 | 2.3514 | 1.3991 | | 1.4783 | 2.6387 | 3.2597 | 2.6742 | 2.6361 | 1.0947 | 1.1029 | C4 | 2.9411 | 2.4023 | 1.4783 | | 1.1605 | 3.9046 | 2.6651 | 3.3611 | 2.1017 | 2.0999 | N5 | 3.7508 | 3.4325 | 2.6387 | 1.1605 | | 4.6701 | 3.1661 | 4.2048 | 3.1688 | 3.1593 | H6 | 1.0917 | 2.0177 | 3.2597 | 3.9046 | 4.6701 | | 1.7855 | 1.7873 | 4.0235 | 3.5911 | H7 | 1.0988 | 2.0886 | 2.6742 | 2.6651 | 3.1661 | 1.7855 | | 1.7903 | 3.6782 | 3.0155 | H8 | 1.1007 | 2.0851 | 2.6361 | 3.3611 | 4.2048 | 1.7873 | 1.7903 | | 3.5857 | 2.4132 | H9 | 3.2682 | 2.0124 | 1.0947 | 2.1017 | 3.1688 | 4.0235 | 3.6782 | 3.5857 | | 1.7774 | H10 | 2.6119 | 2.0802 | 1.1029 | 2.0999 | 3.1593 | 3.5911 | 3.0155 | 2.4132 | 1.7774 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
113.286 |
|
O2 |
C1 |
H6 |
106.423 |
| O2 |
C1 |
H7 |
111.684 |
|
O2 |
C1 |
H8 |
111.276 |
| O2 |
C3 |
C4 |
113.176 |
|
O2 |
C3 |
H9 |
106.968 |
| O2 |
C3 |
H10 |
111.949 |
|
C3 |
C4 |
N5 |
179.017 |
| C4 |
C3 |
H9 |
108.620 |
|
C4 |
C3 |
H10 |
108.004 |
| H6 |
C1 |
H7 |
109.197 |
|
H6 |
C1 |
H8 |
109.221 |
| H7 |
C1 |
H8 |
108.973 |
|
H9 |
C3 |
H10 |
107.960 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.281 |
|
|
|
| 2 |
O |
-0.436 |
|
|
|
| 3 |
C |
-0.136 |
|
|
|
| 4 |
C |
0.357 |
|
|
|
| 5 |
N |
-0.474 |
|
|
|
| 6 |
H |
0.197 |
|
|
|
| 7 |
H |
0.184 |
|
|
|
| 8 |
H |
0.160 |
|
|
|
| 9 |
H |
0.230 |
|
|
|
| 10 |
H |
0.199 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.526 |
1.221 |
1.411 |
3.140 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
4.330 |
1.795 |
0.000 |
4.688 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.701 |
-5.514 |
0.574 |
| y |
-5.514 |
-27.576 |
0.565 |
| z |
0.574 |
0.565 |
-29.335 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.246 |
-5.514 |
0.574 |
| y |
-5.514 |
4.942 |
0.565 |
| z |
0.574 |
0.565 |
2.304 |
|
| Polar |
|---|
| 3z2-r2 | 4.607 |
|---|
| x2-y2 | -8.125 |
|---|
| xy | -5.514 |
|---|
| xz | 0.574 |
|---|
| yz | 0.565 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.760 |
0.492 |
0.187 |
| y |
0.492 |
5.402 |
0.214 |
| z |
0.187 |
0.214 |
4.329 |
<r2> (average value of r
2) Å
2
| <r2> |
122.953 |
| (<r2>)1/2 |
11.088 |
