Jump to
S1C2
Energy calculated at QCISD/6-31G*
| | hartrees |
|---|
| Energy at 0K | -246.540774 |
| Energy at 298.15K | -246.546325 |
| HF Energy | -245.788260 |
| Nuclear repulsion energy | 156.452875 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3184 |
3032 |
19.17 |
|
|
|
| 2 |
A' |
3032 |
2888 |
47.70 |
|
|
|
| 3 |
A' |
3022 |
2878 |
18.54 |
|
|
|
| 4 |
A' |
2357 |
2244 |
1.76 |
|
|
|
| 5 |
A' |
1567 |
1493 |
1.56 |
|
|
|
| 6 |
A' |
1549 |
1475 |
8.32 |
|
|
|
| 7 |
A' |
1518 |
1446 |
0.45 |
|
|
|
| 8 |
A' |
1448 |
1379 |
44.49 |
|
|
|
| 9 |
A' |
1256 |
1196 |
61.01 |
|
|
|
| 10 |
A' |
1189 |
1133 |
106.05 |
|
|
|
| 11 |
A' |
1019 |
970 |
31.64 |
|
|
|
| 12 |
A' |
970 |
924 |
9.59 |
|
|
|
| 13 |
A' |
542 |
516 |
1.15 |
|
|
|
| 14 |
A' |
385 |
366 |
2.38 |
|
|
|
| 15 |
A' |
178 |
170 |
2.39 |
|
|
|
| 16 |
A" |
3090 |
2943 |
48.30 |
|
|
|
| 17 |
A" |
3058 |
2912 |
32.33 |
|
|
|
| 18 |
A" |
1537 |
1464 |
7.16 |
|
|
|
| 19 |
A" |
1286 |
1224 |
3.21 |
|
|
|
| 20 |
A" |
1203 |
1146 |
3.71 |
|
|
|
| 21 |
A" |
1062 |
1011 |
2.30 |
|
|
|
| 22 |
A" |
360 |
343 |
2.21 |
|
|
|
| 23 |
A" |
230 |
219 |
4.92 |
|
|
|
| 24 |
A" |
88 |
84 |
0.12 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17565.6 cm
-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 16727.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.138 |
-0.344 |
0.000 |
| O2 |
-0.739 |
-0.603 |
0.000 |
| C3 |
0.000 |
0.598 |
0.000 |
| C4 |
1.437 |
0.258 |
0.000 |
| N5 |
2.581 |
0.029 |
0.000 |
| H6 |
-2.631 |
-1.319 |
0.000 |
| H7 |
-2.440 |
0.220 |
0.896 |
| H8 |
-2.440 |
0.220 |
-0.896 |
| H9 |
-0.218 |
1.211 |
-0.891 |
| H10 |
-0.218 |
1.211 |
0.891 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4225 | 2.3366 | 3.6258 | 4.7344 | 1.0924 | 1.1010 | 1.1010 | 2.6263 | 2.6263 |
O2 | 1.4225 | | 1.4104 | 2.3408 | 3.3804 | 2.0228 | 2.0910 | 2.0910 | 2.0867 | 2.0867 | C3 | 2.3366 | 1.4104 | | 1.4768 | 2.6437 | 3.2556 | 2.6272 | 2.6272 | 1.1028 | 1.1028 | C4 | 3.6258 | 2.3408 | 1.4768 | | 1.1672 | 4.3635 | 3.9799 | 3.9799 | 2.1075 | 2.1075 | N5 | 4.7344 | 3.3804 | 2.6437 | 1.1672 | | 5.3840 | 5.1047 | 5.1047 | 3.1671 | 3.1671 | H6 | 1.0924 | 2.0228 | 3.2556 | 4.3635 | 5.3840 | | 1.7905 | 1.7905 | 3.6077 | 3.6077 | H7 | 1.1010 | 2.0910 | 2.6272 | 3.9799 | 5.1047 | 1.7905 | | 1.7929 | 3.0189 | 2.4332 | H8 | 1.1010 | 2.0910 | 2.6272 | 3.9799 | 5.1047 | 1.7905 | 1.7929 | | 2.4332 | 3.0189 | H9 | 2.6263 | 2.0867 | 1.1028 | 2.1075 | 3.1671 | 3.6077 | 3.0189 | 2.4332 | | 1.7812 | H10 | 2.6263 | 2.0867 | 1.1028 | 2.1075 | 3.1671 | 3.6077 | 2.4332 | 3.0189 | 1.7812 | |
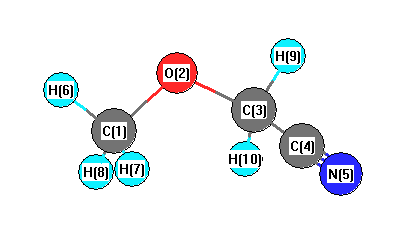 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
111.141 |
|
O2 |
C1 |
H6 |
106.354 |
| O2 |
C1 |
H7 |
111.275 |
|
O2 |
C1 |
H8 |
111.275 |
| O2 |
C3 |
C4 |
108.313 |
|
O2 |
C3 |
H9 |
111.674 |
| O2 |
C3 |
H10 |
111.674 |
|
C3 |
C4 |
N5 |
178.055 |
| C4 |
C3 |
H9 |
108.694 |
|
C4 |
C3 |
H10 |
108.694 |
| H6 |
C1 |
H7 |
109.435 |
|
H6 |
C1 |
H8 |
109.435 |
| H7 |
C1 |
H8 |
109.016 |
|
H9 |
C3 |
H10 |
107.716 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/6-31G*
| | hartrees |
|---|
| Energy at 0K | -246.543334 |
| Energy at 298.15K | -246.549001 |
| HF Energy | -245.790255 |
| Nuclear repulsion energy | 159.343516 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3186 |
3034 |
19.94 |
|
|
|
| 2 |
A |
3148 |
2998 |
6.85 |
|
|
|
| 3 |
A |
3111 |
2962 |
36.79 |
|
|
|
| 4 |
A |
3046 |
2901 |
32.66 |
|
|
|
| 5 |
A |
3041 |
2896 |
28.78 |
|
|
|
| 6 |
A |
2343 |
2231 |
2.32 |
|
|
|
| 7 |
A |
1556 |
1482 |
9.12 |
|
|
|
| 8 |
A |
1541 |
1467 |
7.17 |
|
|
|
| 9 |
A |
1535 |
1462 |
1.56 |
|
|
|
| 10 |
A |
1518 |
1446 |
0.18 |
|
|
|
| 11 |
A |
1422 |
1354 |
18.97 |
|
|
|
| 12 |
A |
1337 |
1273 |
6.88 |
|
|
|
| 13 |
A |
1247 |
1188 |
58.65 |
|
|
|
| 14 |
A |
1208 |
1150 |
15.40 |
|
|
|
| 15 |
A |
1187 |
1130 |
75.80 |
|
|
|
| 16 |
A |
1059 |
1008 |
6.75 |
|
|
|
| 17 |
A |
969 |
922 |
32.10 |
|
|
|
| 18 |
A |
916 |
872 |
22.15 |
|
|
|
| 19 |
A |
597 |
569 |
2.22 |
|
|
|
| 20 |
A |
403 |
384 |
3.18 |
|
|
|
| 21 |
A |
352 |
336 |
1.29 |
|
|
|
| 22 |
A |
259 |
247 |
11.26 |
|
|
|
| 23 |
A |
177 |
168 |
4.42 |
|
|
|
| 24 |
A |
119 |
113 |
8.99 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17637.4 cm
-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 16796.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.560 |
-0.779 |
0.140 |
| O2 |
1.083 |
0.432 |
-0.445 |
| C3 |
-0.055 |
0.937 |
0.213 |
| C4 |
-1.244 |
0.061 |
0.037 |
| N5 |
-2.168 |
-0.641 |
-0.097 |
| H6 |
2.469 |
-1.043 |
-0.407 |
| H7 |
0.825 |
-1.589 |
0.040 |
| H8 |
1.801 |
-0.634 |
1.204 |
| H9 |
-0.268 |
1.917 |
-0.227 |
| H10 |
0.121 |
1.069 |
1.292 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4267 | 2.3579 | 2.9296 | 3.7385 | 1.0923 | 1.0988 | 1.1006 | 3.2781 | 2.6109 |
O2 | 1.4267 | | 1.4074 | 2.4051 | 3.4407 | 2.0238 | 2.0943 | 2.0899 | 2.0192 | 2.0851 | C3 | 2.3579 | 1.4074 | | 1.4883 | 2.6559 | 3.2666 | 2.6808 | 2.6254 | 1.0946 | 1.1019 | C4 | 2.9296 | 2.4051 | 1.4883 | | 1.1677 | 3.8988 | 2.6464 | 3.3340 | 2.1136 | 2.1112 | N5 | 3.7385 | 3.4407 | 2.6559 | 1.1677 | | 4.6644 | 3.1425 | 4.1765 | 3.1886 | 3.1773 | H6 | 1.0923 | 2.0238 | 3.2666 | 3.8988 | 4.6644 | | 1.7894 | 1.7912 | 4.0347 | 3.5859 | H7 | 1.0988 | 2.0943 | 2.6808 | 2.6464 | 3.1425 | 1.7894 | | 1.7940 | 3.6821 | 3.0216 | H8 | 1.1006 | 2.0899 | 2.6254 | 3.3340 | 4.1765 | 1.7912 | 1.7940 | | 3.5823 | 2.3935 | H9 | 3.2781 | 2.0192 | 1.0946 | 2.1136 | 3.1886 | 4.0347 | 3.6821 | 3.5823 | | 1.7830 | H10 | 2.6109 | 2.0851 | 1.1019 | 2.1112 | 3.1773 | 3.5859 | 3.0216 | 2.3935 | 1.7830 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
112.605 |
|
O2 |
C1 |
H6 |
106.148 |
| O2 |
C1 |
H7 |
111.384 |
|
O2 |
C1 |
H8 |
110.908 |
| O2 |
C3 |
C4 |
112.292 |
|
O2 |
C3 |
H9 |
106.951 |
| O2 |
C3 |
H10 |
111.821 |
|
C3 |
C4 |
N5 |
179.154 |
| C4 |
C3 |
H9 |
108.878 |
|
C4 |
C3 |
H10 |
108.266 |
| H6 |
C1 |
H7 |
109.500 |
|
H6 |
C1 |
H8 |
109.531 |
| H7 |
C1 |
H8 |
109.310 |
|
H9 |
C3 |
H10 |
108.534 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
