Jump to
S1C2
Energy calculated at B2PLYP/6-31G*
| | hartrees |
|---|
| Energy at 0K | -247.001994 |
| Energy at 298.15K | -247.007558 |
| HF Energy | -246.761940 |
| Nuclear repulsion energy | 156.721017 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3199 |
3037 |
18.19 |
|
|
|
| 2 |
A' |
3040 |
2885 |
51.38 |
|
|
|
| 3 |
A' |
3024 |
2870 |
22.57 |
|
|
|
| 4 |
A' |
2316 |
2198 |
1.74 |
|
|
|
| 5 |
A' |
1564 |
1485 |
2.59 |
|
|
|
| 6 |
A' |
1546 |
1467 |
7.56 |
|
|
|
| 7 |
A' |
1518 |
1441 |
1.05 |
|
|
|
| 8 |
A' |
1444 |
1371 |
46.36 |
|
|
|
| 9 |
A' |
1247 |
1184 |
43.64 |
|
|
|
| 10 |
A' |
1175 |
1115 |
134.48 |
|
|
|
| 11 |
A' |
1005 |
954 |
32.69 |
|
|
|
| 12 |
A' |
963 |
914 |
7.66 |
|
|
|
| 13 |
A' |
544 |
517 |
1.43 |
|
|
|
| 14 |
A' |
381 |
362 |
2.28 |
|
|
|
| 15 |
A' |
180 |
171 |
2.30 |
|
|
|
| 16 |
A" |
3096 |
2939 |
51.35 |
|
|
|
| 17 |
A" |
3056 |
2901 |
34.50 |
|
|
|
| 18 |
A" |
1537 |
1459 |
7.13 |
|
|
|
| 19 |
A" |
1283 |
1218 |
3.31 |
|
|
|
| 20 |
A" |
1203 |
1142 |
3.42 |
|
|
|
| 21 |
A" |
1058 |
1004 |
1.84 |
|
|
|
| 22 |
A" |
371 |
352 |
1.93 |
|
|
|
| 23 |
A" |
234 |
222 |
4.58 |
|
|
|
| 24 |
A" |
89 |
85 |
0.10 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17536.9 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 16646.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.134 |
-0.355 |
0.000 |
| O2 |
-0.738 |
-0.606 |
0.000 |
| C3 |
0.000 |
0.594 |
0.000 |
| C4 |
1.433 |
0.269 |
0.000 |
| N5 |
2.579 |
0.044 |
0.000 |
| H6 |
-2.622 |
-1.329 |
0.000 |
| H7 |
-2.441 |
0.205 |
0.893 |
| H8 |
-2.441 |
0.205 |
-0.893 |
| H9 |
-0.222 |
1.207 |
-0.887 |
| H10 |
-0.222 |
1.207 |
0.887 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4185 | 2.3354 | 3.6214 | 4.7293 | 1.0892 | 1.0983 | 1.0983 | 2.6231 | 2.6231 |
O2 | 1.4185 | | 1.4082 | 2.3405 | 3.3793 | 2.0186 | 2.0874 | 2.0874 | 2.0825 | 2.0825 | C3 | 2.3354 | 1.4082 | | 1.4699 | 2.6366 | 3.2516 | 2.6282 | 2.6282 | 1.1006 | 1.1006 | C4 | 3.6214 | 2.3405 | 1.4699 | | 1.1669 | 4.3589 | 3.9766 | 3.9766 | 2.0995 | 2.0995 | N5 | 4.7293 | 3.3793 | 2.6366 | 1.1669 | | 5.3790 | 5.1010 | 5.1010 | 3.1595 | 3.1595 | H6 | 1.0892 | 2.0186 | 3.2516 | 4.3589 | 5.3790 | | 1.7844 | 1.7844 | 3.6021 | 3.6021 | H7 | 1.0983 | 2.0874 | 2.6282 | 3.9766 | 5.1010 | 1.7844 | | 1.7862 | 3.0157 | 2.4343 | H8 | 1.0983 | 2.0874 | 2.6282 | 3.9766 | 5.1010 | 1.7844 | 1.7862 | | 2.4343 | 3.0157 | H9 | 2.6231 | 2.0825 | 1.1006 | 2.0995 | 3.1595 | 3.6021 | 3.0157 | 2.4343 | | 1.7739 | H10 | 2.6231 | 2.0825 | 1.1006 | 2.0995 | 3.1595 | 3.6021 | 2.4343 | 3.0157 | 1.7739 | |
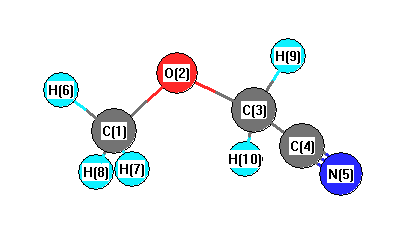 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
111.418 |
|
O2 |
C1 |
H6 |
106.474 |
| O2 |
C1 |
H7 |
111.437 |
|
O2 |
C1 |
H8 |
111.437 |
| O2 |
C3 |
C4 |
108.795 |
|
O2 |
C3 |
H9 |
111.621 |
| O2 |
C3 |
H10 |
111.621 |
|
C3 |
C4 |
N5 |
178.294 |
| C4 |
C3 |
H9 |
108.668 |
|
C4 |
C3 |
H10 |
108.668 |
| H6 |
C1 |
H7 |
109.318 |
|
H6 |
C1 |
H8 |
109.318 |
| H7 |
C1 |
H8 |
108.805 |
|
H9 |
C3 |
H10 |
107.383 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.206 |
|
|
|
| 2 |
O |
-0.470 |
|
|
|
| 3 |
C |
-0.065 |
|
|
|
| 4 |
C |
0.320 |
|
|
|
| 5 |
N |
-0.420 |
|
|
|
| 6 |
H |
0.186 |
|
|
|
| 7 |
H |
0.147 |
|
|
|
| 8 |
H |
0.147 |
|
|
|
| 9 |
H |
0.181 |
|
|
|
| 10 |
H |
0.181 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-4.286 |
1.805 |
0.000 |
4.651 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.725 |
0.750 |
0.000 |
| y |
0.750 |
-29.067 |
0.000 |
| z |
0.000 |
0.000 |
-29.149 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.618 |
0.750 |
0.000 |
| y |
0.750 |
3.371 |
0.000 |
| z |
0.000 |
0.000 |
3.247 |
|
| Polar |
|---|
| 3z2-r2 | 6.494 |
|---|
| x2-y2 | -6.659 |
|---|
| xy | 0.750 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.780 |
-0.007 |
0.000 |
| y |
-0.007 |
4.557 |
0.000 |
| z |
0.000 |
0.000 |
4.280 |
<r2> (average value of r
2) Å
2
| <r2> |
143.181 |
| (<r2>)1/2 |
11.966 |
Jump to
S1C1
Energy calculated at B2PLYP/6-31G*
| | hartrees |
|---|
| Energy at 0K | -247.004518 |
| Energy at 298.15K | -247.010182 |
| HF Energy | -246.764091 |
| Nuclear repulsion energy | 159.546483 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3201 |
3038 |
18.93 |
|
|
|
| 2 |
A |
3155 |
2995 |
6.06 |
|
|
|
| 3 |
A |
3118 |
2959 |
37.69 |
|
|
|
| 4 |
A |
3051 |
2896 |
42.27 |
|
|
|
| 5 |
A |
3045 |
2890 |
26.24 |
|
|
|
| 6 |
A |
2298 |
2181 |
1.88 |
|
|
|
| 7 |
A |
1557 |
1478 |
9.16 |
|
|
|
| 8 |
A |
1539 |
1461 |
6.36 |
|
|
|
| 9 |
A |
1527 |
1450 |
3.00 |
|
|
|
| 10 |
A |
1516 |
1439 |
0.21 |
|
|
|
| 11 |
A |
1419 |
1347 |
19.04 |
|
|
|
| 12 |
A |
1334 |
1266 |
6.67 |
|
|
|
| 13 |
A |
1240 |
1177 |
35.36 |
|
|
|
| 14 |
A |
1205 |
1144 |
7.80 |
|
|
|
| 15 |
A |
1173 |
1113 |
111.54 |
|
|
|
| 16 |
A |
1053 |
1000 |
7.18 |
|
|
|
| 17 |
A |
955 |
907 |
32.45 |
|
|
|
| 18 |
A |
909 |
862 |
20.73 |
|
|
|
| 19 |
A |
598 |
568 |
2.08 |
|
|
|
| 20 |
A |
397 |
377 |
2.69 |
|
|
|
| 21 |
A |
359 |
341 |
1.28 |
|
|
|
| 22 |
A |
258 |
245 |
10.41 |
|
|
|
| 23 |
A |
179 |
170 |
4.08 |
|
|
|
| 24 |
A |
120 |
114 |
8.82 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17602.0 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 16707.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.585 |
-0.760 |
0.138 |
| O2 |
1.070 |
0.425 |
-0.445 |
| C3 |
-0.059 |
0.925 |
0.209 |
| C4 |
-1.256 |
0.059 |
0.042 |
| N5 |
-2.175 |
-0.643 |
-0.097 |
| H6 |
2.480 |
-1.018 |
-0.428 |
| H7 |
0.869 |
-1.588 |
0.076 |
| H8 |
1.856 |
-0.602 |
1.191 |
| H9 |
-0.277 |
1.904 |
-0.224 |
| H10 |
0.110 |
1.057 |
1.288 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4174 | 2.3548 | 2.9577 | 3.7691 | 1.0896 | 1.0971 | 1.0987 | 3.2698 | 2.6075 |
O2 | 1.4174 | | 1.3971 | 2.4045 | 3.4342 | 2.0173 | 2.0896 | 2.0857 | 2.0124 | 2.0799 | C3 | 2.3548 | 1.3971 | | 1.4872 | 2.6516 | 3.2594 | 2.6824 | 2.6390 | 1.0921 | 1.1003 | C4 | 2.9577 | 2.4045 | 1.4872 | | 1.1646 | 3.9157 | 2.6881 | 3.3824 | 2.1056 | 2.1010 | N5 | 3.7691 | 3.4342 | 2.6516 | 1.1646 | | 4.6816 | 3.1918 | 4.2322 | 3.1786 | 3.1666 | H6 | 1.0896 | 2.0173 | 3.2594 | 3.9157 | 4.6816 | | 1.7817 | 1.7836 | 4.0220 | 3.5868 | H7 | 1.0971 | 2.0896 | 2.6824 | 2.6881 | 3.1918 | 1.7817 | | 1.7865 | 3.6873 | 3.0072 | H8 | 1.0987 | 2.0857 | 2.6390 | 3.3824 | 4.2322 | 1.7836 | 1.7865 | | 3.5823 | 2.4109 | H9 | 3.2698 | 2.0124 | 1.0921 | 2.1056 | 3.1786 | 4.0220 | 3.6873 | 3.5823 | | 1.7758 | H10 | 2.6075 | 2.0799 | 1.1003 | 2.1010 | 3.1666 | 3.5868 | 3.0072 | 2.4109 | 1.7758 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
113.579 |
|
O2 |
C1 |
H6 |
106.429 |
| O2 |
C1 |
H7 |
111.776 |
|
O2 |
C1 |
H8 |
111.355 |
| O2 |
C3 |
C4 |
112.912 |
|
O2 |
C3 |
H9 |
107.251 |
| O2 |
C3 |
H10 |
112.231 |
|
C3 |
C4 |
N5 |
178.523 |
| C4 |
C3 |
H9 |
108.466 |
|
C4 |
C3 |
H10 |
107.641 |
| H6 |
C1 |
H7 |
109.136 |
|
H6 |
C1 |
H8 |
109.187 |
| H7 |
C1 |
H8 |
108.890 |
|
H9 |
C3 |
H10 |
108.191 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.211 |
|
|
|
| 2 |
O |
-0.504 |
|
|
|
| 3 |
C |
-0.058 |
|
|
|
| 4 |
C |
0.324 |
|
|
|
| 5 |
N |
-0.463 |
|
|
|
| 6 |
H |
0.184 |
|
|
|
| 7 |
H |
0.172 |
|
|
|
| 8 |
H |
0.147 |
|
|
|
| 9 |
H |
0.219 |
|
|
|
| 10 |
H |
0.189 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.527 |
1.243 |
1.524 |
3.202 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.285 |
-5.729 |
0.678 |
| y |
-5.729 |
-27.784 |
0.607 |
| z |
0.678 |
0.607 |
-29.619 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.584 |
-5.729 |
0.678 |
| y |
-5.729 |
5.168 |
0.607 |
| z |
0.678 |
0.607 |
2.415 |
|
| Polar |
|---|
| 3z2-r2 | 4.830 |
|---|
| x2-y2 | -8.501 |
|---|
| xy | -5.729 |
|---|
| xz | 0.678 |
|---|
| yz | 0.607 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.601 |
0.461 |
0.174 |
| y |
0.461 |
5.349 |
0.202 |
| z |
0.174 |
0.202 |
4.278 |
<r2> (average value of r
2) Å
2
| <r2> |
123.225 |
| (<r2>)1/2 |
11.101 |
