Jump to
S1C2
Energy calculated at PBEPBEultrafine/6-31G*
| | hartrees |
|---|
| Energy at 0K | -246.956037 |
| Energy at 298.15K | -246.961521 |
| HF Energy | -246.956037 |
| Nuclear repulsion energy | 156.049779 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3091 |
3040 |
16.28 |
|
|
|
| 2 |
A' |
2921 |
2872 |
56.54 |
|
|
|
| 3 |
A' |
2891 |
2844 |
29.88 |
|
|
|
| 4 |
A' |
2295 |
2257 |
0.33 |
|
|
|
| 5 |
A' |
1495 |
1470 |
4.32 |
|
|
|
| 6 |
A' |
1470 |
1445 |
9.46 |
|
|
|
| 7 |
A' |
1447 |
1423 |
1.53 |
|
|
|
| 8 |
A' |
1372 |
1350 |
36.24 |
|
|
|
| 9 |
A' |
1186 |
1167 |
30.99 |
|
|
|
| 10 |
A' |
1132 |
1114 |
136.70 |
|
|
|
| 11 |
A' |
975 |
959 |
29.56 |
|
|
|
| 12 |
A' |
934 |
919 |
5.85 |
|
|
|
| 13 |
A' |
533 |
524 |
1.25 |
|
|
|
| 14 |
A' |
370 |
364 |
1.70 |
|
|
|
| 15 |
A' |
174 |
171 |
2.03 |
|
|
|
| 16 |
A" |
2969 |
2920 |
54.43 |
|
|
|
| 17 |
A" |
2912 |
2864 |
38.58 |
|
|
|
| 18 |
A" |
1463 |
1439 |
8.32 |
|
|
|
| 19 |
A" |
1220 |
1200 |
3.19 |
|
|
|
| 20 |
A" |
1151 |
1132 |
2.04 |
|
|
|
| 21 |
A" |
1005 |
988 |
0.99 |
|
|
|
| 22 |
A" |
370 |
363 |
1.24 |
|
|
|
| 23 |
A" |
238 |
234 |
3.99 |
|
|
|
| 24 |
A" |
89 |
87 |
0.04 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16851.2 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 16573.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.138 |
-0.354 |
0.000 |
| O2 |
-0.741 |
-0.620 |
0.000 |
| C3 |
0.000 |
0.585 |
0.000 |
| C4 |
1.437 |
0.275 |
0.000 |
| N5 |
2.589 |
0.059 |
0.000 |
| H6 |
-2.641 |
-1.333 |
0.000 |
| H7 |
-2.449 |
0.215 |
0.901 |
| H8 |
-2.449 |
0.215 |
-0.901 |
| H9 |
-0.227 |
1.210 |
-0.894 |
| H10 |
-0.227 |
1.210 |
0.894 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4219 | 2.3350 | 3.6295 | 4.7447 | 1.1001 | 1.1106 | 1.1106 | 2.6262 | 2.6262 |
O2 | 1.4219 | | 1.4144 | 2.3542 | 3.3982 | 2.0295 | 2.1042 | 2.1042 | 2.1003 | 2.1003 | C3 | 2.3350 | 1.4144 | | 1.4700 | 2.6420 | 3.2637 | 2.6356 | 2.6356 | 1.1142 | 1.1142 | C4 | 3.6295 | 2.3542 | 1.4700 | | 1.1723 | 4.3831 | 3.9894 | 3.9894 | 2.1078 | 2.1078 | N5 | 4.7447 | 3.3982 | 2.6420 | 1.1723 | | 5.4119 | 5.1204 | 5.1204 | 3.1710 | 3.1710 | H6 | 1.1001 | 2.0295 | 3.2637 | 4.3831 | 5.4119 | | 1.8013 | 1.8013 | 3.6181 | 3.6181 | H7 | 1.1106 | 2.1042 | 2.6356 | 3.9894 | 5.1204 | 1.8013 | | 1.8024 | 3.0248 | 2.4344 | H8 | 1.1106 | 2.1042 | 2.6356 | 3.9894 | 5.1204 | 1.8013 | 1.8024 | | 2.4344 | 3.0248 | H9 | 2.6262 | 2.1003 | 1.1142 | 2.1078 | 3.1710 | 3.6181 | 3.0248 | 2.4344 | | 1.7882 | H10 | 2.6262 | 2.1003 | 1.1142 | 2.1078 | 3.1710 | 3.6181 | 2.4344 | 3.0248 | 1.7882 | |
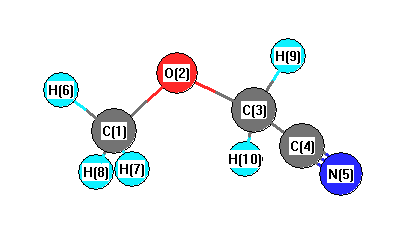 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
110.821 |
|
O2 |
C1 |
H6 |
106.464 |
| O2 |
C1 |
H7 |
111.784 |
|
O2 |
C1 |
H8 |
111.784 |
| O2 |
C3 |
C4 |
109.390 |
|
O2 |
C3 |
H9 |
111.780 |
| O2 |
C3 |
H10 |
111.780 |
|
C3 |
C4 |
N5 |
178.422 |
| C4 |
C3 |
H9 |
108.517 |
|
C4 |
C3 |
H10 |
108.517 |
| H6 |
C1 |
H7 |
109.136 |
|
H6 |
C1 |
H8 |
109.136 |
| H7 |
C1 |
H8 |
108.473 |
|
H9 |
C3 |
H10 |
106.735 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.258 |
|
|
|
| 2 |
O |
-0.399 |
|
|
|
| 3 |
C |
-0.124 |
|
|
|
| 4 |
C |
0.369 |
|
|
|
| 5 |
N |
-0.454 |
|
|
|
| 6 |
H |
0.190 |
|
|
|
| 7 |
H |
0.153 |
|
|
|
| 8 |
H |
0.153 |
|
|
|
| 9 |
H |
0.185 |
|
|
|
| 10 |
H |
0.185 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-4.161 |
1.644 |
0.000 |
4.474 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.441 |
0.685 |
0.000 |
| y |
0.685 |
-28.913 |
0.000 |
| z |
0.000 |
0.000 |
-29.072 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.449 |
0.685 |
0.000 |
| y |
0.685 |
3.344 |
0.000 |
| z |
0.000 |
0.000 |
3.105 |
|
| Polar |
|---|
| 3z2-r2 | 6.209 |
|---|
| x2-y2 | -6.529 |
|---|
| xy | 0.685 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.400 |
0.017 |
0.000 |
| y |
0.017 |
4.751 |
0.000 |
| z |
0.000 |
0.000 |
4.449 |
<r2> (average value of r
2) Å
2
| <r2> |
144.018 |
| (<r2>)1/2 |
12.001 |
Jump to
S1C1
Energy calculated at PBEPBEultrafine/6-31G*
| | hartrees |
|---|
| Energy at 0K | -246.958856 |
| Energy at 298.15K | -246.964405 |
| HF Energy | -246.958856 |
| Nuclear repulsion energy | 158.875535 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3095 |
3044 |
16.69 |
|
|
|
| 2 |
A |
3039 |
2989 |
5.17 |
|
|
|
| 3 |
A |
2999 |
2950 |
36.74 |
|
|
|
| 4 |
A |
2936 |
2888 |
46.88 |
|
|
|
| 5 |
A |
2916 |
2868 |
32.49 |
|
|
|
| 6 |
A |
2275 |
2238 |
0.05 |
|
|
|
| 7 |
A |
1487 |
1463 |
10.07 |
|
|
|
| 8 |
A |
1465 |
1441 |
7.10 |
|
|
|
| 9 |
A |
1452 |
1428 |
2.41 |
|
|
|
| 10 |
A |
1439 |
1415 |
3.34 |
|
|
|
| 11 |
A |
1352 |
1330 |
13.91 |
|
|
|
| 12 |
A |
1276 |
1255 |
6.03 |
|
|
|
| 13 |
A |
1180 |
1161 |
20.26 |
|
|
|
| 14 |
A |
1151 |
1132 |
6.44 |
|
|
|
| 15 |
A |
1121 |
1103 |
112.82 |
|
|
|
| 16 |
A |
1002 |
985 |
6.21 |
|
|
|
| 17 |
A |
925 |
909 |
28.92 |
|
|
|
| 18 |
A |
883 |
869 |
15.49 |
|
|
|
| 19 |
A |
584 |
574 |
2.14 |
|
|
|
| 20 |
A |
382 |
376 |
2.37 |
|
|
|
| 21 |
A |
351 |
346 |
1.09 |
|
|
|
| 22 |
A |
249 |
245 |
8.41 |
|
|
|
| 23 |
A |
174 |
171 |
3.89 |
|
|
|
| 24 |
A |
116 |
114 |
7.95 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16924.4 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 16645.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.565 |
-0.777 |
0.140 |
| O2 |
1.086 |
0.434 |
-0.445 |
| C3 |
-0.061 |
0.935 |
0.210 |
| C4 |
-1.251 |
0.068 |
0.043 |
| N5 |
-2.167 |
-0.650 |
-0.101 |
| H6 |
2.471 |
-1.051 |
-0.422 |
| H7 |
0.823 |
-1.595 |
0.058 |
| H8 |
1.826 |
-0.635 |
1.209 |
| H9 |
-0.272 |
1.923 |
-0.234 |
| H10 |
0.113 |
1.079 |
1.300 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4278 | 2.3627 | 2.9422 | 3.7428 | 1.0998 | 1.1075 | 1.1097 | 3.2875 | 2.6274 |
O2 | 1.4278 | | 1.4131 | 2.4155 | 3.4465 | 2.0304 | 2.1066 | 2.1035 | 2.0267 | 2.0997 | C3 | 2.3627 | 1.4131 | | 1.4818 | 2.6545 | 3.2793 | 2.6842 | 2.6499 | 1.1034 | 1.1126 | C4 | 2.9422 | 2.4155 | 1.4818 | | 1.1732 | 3.9140 | 2.6583 | 3.3646 | 2.1152 | 2.1122 | N5 | 3.7428 | 3.4465 | 2.6545 | 1.1732 | | 4.6664 | 3.1400 | 4.2030 | 3.1980 | 3.1864 | H6 | 1.0998 | 2.0304 | 3.2793 | 3.9140 | 4.6664 | | 1.8002 | 1.8021 | 4.0502 | 3.6141 | H7 | 1.1075 | 2.1066 | 2.6842 | 2.6583 | 3.1400 | 1.8002 | | 1.8039 | 3.6957 | 3.0329 | H8 | 1.1097 | 2.1035 | 2.6499 | 3.3646 | 4.2030 | 1.8021 | 1.8039 | | 3.6090 | 2.4251 | H9 | 3.2875 | 2.0267 | 1.1034 | 2.1152 | 3.1980 | 4.0502 | 3.6957 | 3.6090 | | 1.7921 | H10 | 2.6274 | 2.0997 | 1.1126 | 2.1122 | 3.1864 | 3.6141 | 3.0329 | 2.4251 | 1.7921 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
112.542 |
|
O2 |
C1 |
H6 |
106.168 |
| O2 |
C1 |
H7 |
111.761 |
|
O2 |
C1 |
H8 |
111.368 |
| O2 |
C3 |
C4 |
113.085 |
|
O2 |
C3 |
H9 |
106.642 |
| O2 |
C3 |
H10 |
111.927 |
|
C3 |
C4 |
N5 |
177.916 |
| C4 |
C3 |
H9 |
108.934 |
|
C4 |
C3 |
H10 |
108.162 |
| H6 |
C1 |
H7 |
109.287 |
|
H6 |
C1 |
H8 |
109.294 |
| H7 |
C1 |
H8 |
108.897 |
|
H9 |
C3 |
H10 |
107.934 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.277 |
|
|
|
| 2 |
O |
-0.388 |
|
|
|
| 3 |
C |
-0.150 |
|
|
|
| 4 |
C |
0.354 |
|
|
|
| 5 |
N |
-0.454 |
|
|
|
| 6 |
H |
0.186 |
|
|
|
| 7 |
H |
0.176 |
|
|
|
| 8 |
H |
0.151 |
|
|
|
| 9 |
H |
0.215 |
|
|
|
| 10 |
H |
0.186 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.452 |
1.210 |
1.326 |
3.038 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.577 |
-5.326 |
0.492 |
| y |
-5.326 |
-27.714 |
0.512 |
| z |
0.492 |
0.512 |
-29.468 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.986 |
-5.326 |
0.492 |
| y |
-5.326 |
4.809 |
0.512 |
| z |
0.492 |
0.512 |
2.177 |
|
| Polar |
|---|
| 3z2-r2 | 4.355 |
|---|
| x2-y2 | -7.863 |
|---|
| xy | -5.326 |
|---|
| xz | 0.492 |
|---|
| yz | 0.512 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.069 |
0.478 |
0.198 |
| y |
0.478 |
5.634 |
0.228 |
| z |
0.198 |
0.228 |
4.450 |
<r2> (average value of r
2) Å
2
| <r2> |
123.664 |
| (<r2>)1/2 |
11.120 |
