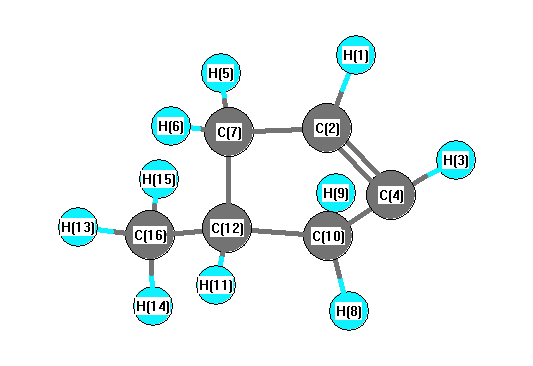
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3225 |
3085 |
33.98 |
|
|
|
| 2 |
A |
3200 |
3061 |
9.58 |
|
|
|
| 3 |
A |
3123 |
2988 |
27.77 |
|
|
|
| 4 |
A |
3121 |
2986 |
48.42 |
|
|
|
| 5 |
A |
3081 |
2948 |
38.52 |
|
|
|
| 6 |
A |
3076 |
2943 |
35.58 |
|
|
|
| 7 |
A |
3047 |
2915 |
39.20 |
|
|
|
| 8 |
A |
3044 |
2912 |
8.24 |
|
|
|
| 9 |
A |
3013 |
2883 |
27.33 |
|
|
|
| 10 |
A |
3013 |
2883 |
43.70 |
|
|
|
| 11 |
A |
1709 |
1635 |
2.73 |
|
|
|
| 12 |
A |
1527 |
1461 |
7.32 |
|
|
|
| 13 |
A |
1523 |
1457 |
2.49 |
|
|
|
| 14 |
A |
1517 |
1451 |
0.33 |
|
|
|
| 15 |
A |
1509 |
1444 |
1.15 |
|
|
|
| 16 |
A |
1433 |
1371 |
3.04 |
|
|
|
| 17 |
A |
1390 |
1330 |
0.26 |
|
|
|
| 18 |
A |
1390 |
1330 |
3.85 |
|
|
|
| 19 |
A |
1350 |
1291 |
2.29 |
|
|
|
| 20 |
A |
1320 |
1263 |
0.11 |
|
|
|
| 21 |
A |
1307 |
1250 |
3.47 |
|
|
|
| 22 |
A |
1249 |
1195 |
0.42 |
|
|
|
| 23 |
A |
1182 |
1131 |
0.00 |
|
|
|
| 24 |
A |
1161 |
1111 |
2.52 |
|
|
|
| 25 |
A |
1142 |
1092 |
0.21 |
|
|
|
| 26 |
A |
1121 |
1072 |
0.59 |
|
|
|
| 27 |
A |
1079 |
1032 |
1.67 |
|
|
|
| 28 |
A |
1003 |
959 |
9.02 |
|
|
|
| 29 |
A |
995 |
951 |
0.38 |
|
|
|
| 30 |
A |
971 |
929 |
0.00 |
|
|
|
| 31 |
A |
940 |
900 |
4.92 |
|
|
|
| 32 |
A |
936 |
895 |
4.04 |
|
|
|
| 33 |
A |
914 |
875 |
0.02 |
|
|
|
| 34 |
A |
832 |
796 |
1.28 |
|
|
|
| 35 |
A |
778 |
744 |
0.45 |
|
|
|
| 36 |
A |
708 |
677 |
33.87 |
|
|
|
| 37 |
A |
573 |
548 |
3.84 |
|
|
|
| 38 |
A |
422 |
404 |
1.18 |
|
|
|
| 39 |
A |
395 |
378 |
0.09 |
|
|
|
| 40 |
A |
313 |
299 |
0.10 |
|
|
|
| 41 |
A |
254 |
243 |
0.01 |
|
|
|
| 42 |
A |
104 |
100 |
0.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31994.1 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 30608.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.152 |
|
|
0.139 |
| 2 |
C |
-0.137 |
|
|
-0.222 |
| 3 |
H |
0.152 |
|
|
0.139 |
| 4 |
C |
-0.137 |
|
|
-0.222 |
| 5 |
H |
0.166 |
|
|
0.059 |
| 6 |
H |
0.165 |
|
|
0.035 |
| 7 |
C |
-0.353 |
|
|
-0.063 |
| 8 |
H |
0.165 |
|
|
0.035 |
| 9 |
H |
0.166 |
|
|
0.059 |
| 10 |
C |
-0.353 |
|
|
-0.063 |
| 11 |
H |
0.164 |
|
|
0.017 |
| 12 |
C |
-0.128 |
|
|
0.240 |
| 13 |
H |
0.166 |
|
|
0.123 |
| 14 |
H |
0.166 |
|
|
0.123 |
| 15 |
H |
0.164 |
|
|
0.135 |
| 16 |
C |
-0.517 |
|
|
-0.535 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.163 |
0.000 |
0.017 |
0.164 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.161 |
0.000 |
-0.005 |
0.161 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.574 |
0.000 |
-0.373 |
| y |
0.000 |
-36.491 |
0.000 |
| z |
-0.373 |
0.000 |
-38.586 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.035 |
0.000 |
-0.373 |
| y |
0.000 |
1.589 |
0.000 |
| z |
-0.373 |
0.000 |
-1.553 |
|
| Polar |
|---|
| 3z2-r2 | -3.107 |
|---|
| x2-y2 | -1.083 |
|---|
| xy | 0.000 |
|---|
| xz | -0.373 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
163.818 |
| (<r2>)1/2 |
12.799 |