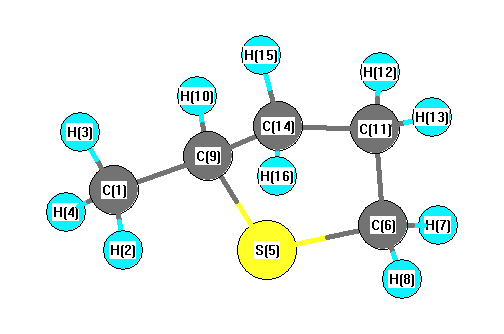
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3145 |
3009 |
27.76 |
|
|
|
| 2 |
A |
3142 |
3006 |
16.61 |
|
|
|
| 3 |
A |
3128 |
2992 |
25.65 |
|
|
|
| 4 |
A |
3115 |
2981 |
34.88 |
|
|
|
| 5 |
A |
3104 |
2970 |
26.61 |
|
|
|
| 6 |
A |
3083 |
2949 |
28.47 |
|
|
|
| 7 |
A |
3064 |
2931 |
13.56 |
|
|
|
| 8 |
A |
3064 |
2931 |
25.57 |
|
|
|
| 9 |
A |
3055 |
2922 |
44.92 |
|
|
|
| 10 |
A |
3051 |
2918 |
7.97 |
|
|
|
| 11 |
A |
1529 |
1463 |
1.57 |
|
|
|
| 12 |
A |
1523 |
1457 |
6.28 |
|
|
|
| 13 |
A |
1514 |
1448 |
6.00 |
|
|
|
| 14 |
A |
1509 |
1444 |
4.22 |
|
|
|
| 15 |
A |
1505 |
1440 |
3.28 |
|
|
|
| 16 |
A |
1434 |
1371 |
4.12 |
|
|
|
| 17 |
A |
1395 |
1334 |
1.57 |
|
|
|
| 18 |
A |
1372 |
1312 |
0.50 |
|
|
|
| 19 |
A |
1333 |
1276 |
0.19 |
|
|
|
| 20 |
A |
1317 |
1260 |
5.50 |
|
|
|
| 21 |
A |
1299 |
1243 |
26.23 |
|
|
|
| 22 |
A |
1251 |
1197 |
4.69 |
|
|
|
| 23 |
A |
1214 |
1161 |
6.55 |
|
|
|
| 24 |
A |
1171 |
1120 |
0.48 |
|
|
|
| 25 |
A |
1121 |
1073 |
0.10 |
|
|
|
| 26 |
A |
1088 |
1041 |
2.45 |
|
|
|
| 27 |
A |
1050 |
1004 |
3.94 |
|
|
|
| 28 |
A |
1049 |
1003 |
1.39 |
|
|
|
| 29 |
A |
983 |
941 |
3.16 |
|
|
|
| 30 |
A |
964 |
922 |
2.09 |
|
|
|
| 31 |
A |
913 |
874 |
0.86 |
|
|
|
| 32 |
A |
875 |
837 |
2.51 |
|
|
|
| 33 |
A |
859 |
821 |
1.88 |
|
|
|
| 34 |
A |
700 |
670 |
1.94 |
|
|
|
| 35 |
A |
642 |
614 |
2.97 |
|
|
|
| 36 |
A |
529 |
506 |
0.78 |
|
|
|
| 37 |
A |
461 |
441 |
0.09 |
|
|
|
| 38 |
A |
413 |
395 |
0.76 |
|
|
|
| 39 |
A |
284 |
272 |
1.09 |
|
|
|
| 40 |
A |
257 |
246 |
0.16 |
|
|
|
| 41 |
A |
242 |
231 |
0.21 |
|
|
|
| 42 |
A |
94 |
90 |
1.50 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31418.3 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 30057.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.506 |
|
|
-0.515 |
| 2 |
H |
0.182 |
|
|
0.126 |
| 3 |
H |
0.169 |
|
|
0.116 |
| 4 |
H |
0.182 |
|
|
0.175 |
| 5 |
S |
0.081 |
|
|
-0.327 |
| 6 |
C |
-0.472 |
|
|
-0.052 |
| 7 |
H |
0.199 |
|
|
0.107 |
| 8 |
H |
0.196 |
|
|
0.101 |
| 9 |
C |
-0.282 |
|
|
0.265 |
| 10 |
H |
0.191 |
|
|
0.037 |
| 11 |
C |
-0.323 |
|
|
-0.309 |
| 12 |
H |
0.177 |
|
|
0.116 |
| 13 |
H |
0.174 |
|
|
0.114 |
| 14 |
C |
-0.313 |
|
|
-0.098 |
| 15 |
H |
0.172 |
|
|
0.060 |
| 16 |
H |
0.173 |
|
|
0.086 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.378 |
2.137 |
0.007 |
2.170 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.380 |
2.136 |
-0.011 |
2.169 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.801 |
-0.332 |
0.983 |
| y |
-0.332 |
-47.792 |
-0.350 |
| z |
0.983 |
-0.350 |
-45.488 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.839 |
-0.332 |
0.983 |
| y |
-0.332 |
-4.148 |
-0.350 |
| z |
0.983 |
-0.350 |
-0.691 |
|
| Polar |
|---|
| 3z2-r2 | -1.382 |
|---|
| x2-y2 | 5.991 |
|---|
| xy | -0.332 |
|---|
| xz | 0.983 |
|---|
| yz | -0.350 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
203.887 |
| (<r2>)1/2 |
14.279 |