return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10S (Thiophene, tetrahydro-2-methyl-)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-594.333909
Energy at 298.15K-594.345768
HF Energy-594.076202
Nuclear repulsion energy310.305113
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3167 3006 30.59      
2 A 3166 3005 13.86      
3 A 3149 2989 25.11      
4 A 3136 2976 36.39      
5 A 3124 2966 25.89      
6 A 3107 2949 26.99      
7 A 3089 2932 12.68      
8 A 3085 2928 22.94      
9 A 3077 2921 40.12      
10 A 3072 2916 9.98      
11 A 1554 1475 1.47      
12 A 1548 1470 5.31      
13 A 1539 1461 4.89      
14 A 1535 1457 3.82      
15 A 1531 1453 2.13      
16 A 1457 1383 2.74      
17 A 1414 1342 1.62      
18 A 1391 1320 0.61      
19 A 1354 1285 0.70      
20 A 1339 1271 6.64      
21 A 1320 1253 27.83      
22 A 1268 1203 4.04      
23 A 1229 1166 5.75      
24 A 1183 1123 0.35      
25 A 1134 1076 0.13      
26 A 1102 1046 2.26      
27 A 1063 1009 3.77      
28 A 1054 1000 0.48      
29 A 998 948 2.37      
30 A 973 924 1.96      
31 A 918 872 0.67      
32 A 886 841 2.25      
33 A 866 822 1.41      
34 A 701 665 2.12      
35 A 644 611 3.07      
36 A 536 509 0.62      
37 A 465 441 0.06      
38 A 418 397 0.65      
39 A 289 275 0.94      
40 A 262 248 0.14      
41 A 248 235 0.21      
42 A 91 87 1.44      

Unscaled Zero Point Vibrational Energy (zpe) 31738.8 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 30126.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.13955 0.09256 0.06107

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 2.306 0.141 -0.021
H2 2.410 0.030 -1.102
H3 2.804 1.061 0.289
H4 2.801 -0.701 0.468
S5 -0.106 -1.283 -0.084
C6 -1.707 -0.388 0.001
H7 -2.286 -0.729 0.861
H8 -2.278 -0.602 -0.903
C9 0.828 0.211 0.367
H10 0.748 0.338 1.449
C11 -1.396 1.122 0.135
H12 -1.443 1.389 1.193
H13 -2.122 1.735 -0.405
C14 0.051 1.332 -0.323
H15 0.431 2.322 -0.056
H16 0.134 1.190 -1.403

Atom - Atom Distances (Å)
  C1 H2 H3 H4 S5 C6 H7 H8 C9 H10 C11 H12 H13 C14 H15 H16
C11.09151.09151.09192.80154.04804.75634.72691.52982.15173.83344.13434.72232.56822.87672.7806
H21.09151.77561.77503.01454.28225.14564.73412.16593.06044.14824.68624.89192.80443.20372.5725
H31.09151.77561.77163.75534.74715.42625.47862.15282.46994.20374.35535.02042.83322.70933.1637
H41.09191.77501.77163.01534.54295.10235.26152.17602.50214.58814.78675.56183.51013.87683.7672
S52.80153.01453.75533.01531.83612.43972.41911.81842.38882.73773.24963.64352.63023.64442.8133
C64.04804.28224.74714.54291.83611.09151.09042.63052.94131.54742.15632.20082.48063.45212.8019
H74.75635.14565.42625.10232.43971.09151.76873.29013.26932.17812.30392.77493.33334.18683.8295
H84.72694.73415.47865.26152.41911.09041.76873.45263.94622.19723.00932.39463.08234.07503.0461
C91.52982.16592.15282.17601.81842.63053.29013.45261.09312.41462.68903.40911.52832.18902.1385
H102.15173.06042.46992.50212.38882.94133.26933.94621.09312.63402.44393.69142.14822.51013.0396
C113.83344.14824.20374.58812.73771.54742.17812.19722.41462.63401.09191.09301.53252.19422.1705
H124.13434.68624.35534.78673.24962.15632.30393.00932.68902.44391.09191.76992.12952.43743.0438
H134.72234.89195.02045.56183.64352.20082.77492.39463.40913.69141.09301.76992.21192.64302.5262
C142.56822.80442.83323.51012.63022.48063.33333.08231.52832.14821.53252.12952.21191.09361.0922
H152.87673.20372.70933.87683.64443.45214.18684.07502.18902.51012.19422.43742.64301.09361.7842
H162.78062.57253.16373.76722.81332.80193.82953.04612.13853.03962.17053.04382.52621.09221.7842

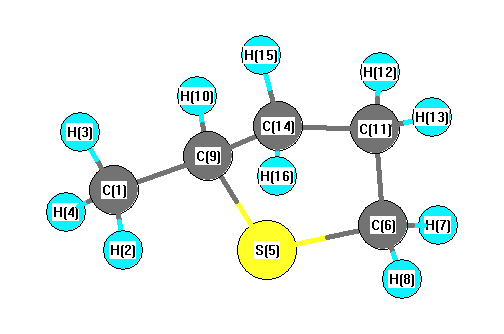
picture of Thiophene, tetrahydro-2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C9 S5 113.311 C1 C9 H10 109.110
C1 C9 C14 114.239 H2 C1 H3 108.856
H2 C1 H4 108.768 H2 C1 C9 110.318
H3 C1 H4 108.467 H3 C1 C9 109.288
H4 C1 C9 111.094 S5 C6 H7 110.278
S5 C6 H8 108.823 S5 C6 C11 107.718
S5 C9 H10 107.647 S5 C9 C14 103.268
C6 S5 C9 92.071 C6 C11 H12 108.341
C6 C11 H13 111.769 C6 C11 C14 107.300
H7 C6 H8 108.320 H7 C6 C11 110.054
H8 C6 C11 111.645 C9 C14 C11 104.161
C9 C14 H15 112.140 C9 C14 H16 108.236
H10 C9 C14 108.933 C11 C14 H15 112.264
C11 C14 H16 110.453 H12 C11 H13 108.205
H12 C11 C14 107.274 H13 C11 C14 113.743
H15 C14 H16 109.431
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.457      
2 H 0.167      
3 H 0.153      
4 H 0.166      
5 S 0.072      
6 C -0.436      
7 H 0.184      
8 H 0.180      
9 C -0.262      
10 H 0.175      
11 C -0.293      
12 H 0.161      
13 H 0.160      
14 C -0.284      
15 H 0.158      
16 H 0.157      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 11.095 -0.126 0.282
y -0.126 10.117 -0.027
z 0.282 -0.027 8.070


<r2> (average value of r2) Å2
<r2> 204.494
(<r2>)1/2 14.300