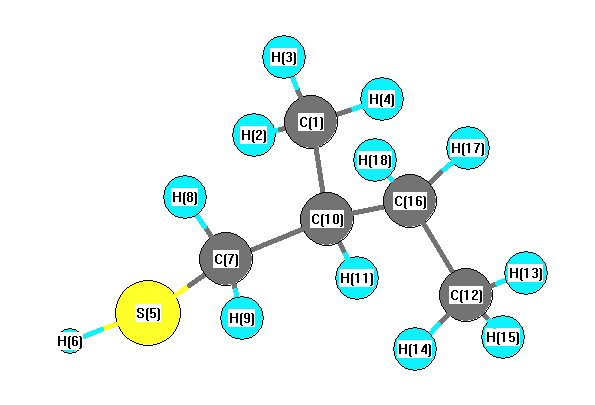
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3141 |
3005 |
26.35 |
|
|
|
| 2 |
A |
3139 |
3004 |
20.86 |
|
|
|
| 3 |
A |
3132 |
2996 |
44.84 |
|
|
|
| 4 |
A |
3129 |
2993 |
47.77 |
|
|
|
| 5 |
A |
3124 |
2989 |
16.00 |
|
|
|
| 6 |
A |
3078 |
2944 |
24.18 |
|
|
|
| 7 |
A |
3072 |
2939 |
20.66 |
|
|
|
| 8 |
A |
3056 |
2923 |
26.41 |
|
|
|
| 9 |
A |
3053 |
2921 |
30.30 |
|
|
|
| 10 |
A |
3040 |
2908 |
20.45 |
|
|
|
| 11 |
A |
3032 |
2901 |
5.06 |
|
|
|
| 12 |
A |
2715 |
2597 |
25.20 |
|
|
|
| 13 |
A |
1534 |
1467 |
9.12 |
|
|
|
| 14 |
A |
1526 |
1460 |
8.31 |
|
|
|
| 15 |
A |
1526 |
1460 |
5.71 |
|
|
|
| 16 |
A |
1523 |
1457 |
2.17 |
|
|
|
| 17 |
A |
1510 |
1444 |
2.33 |
|
|
|
| 18 |
A |
1502 |
1437 |
6.61 |
|
|
|
| 19 |
A |
1438 |
1376 |
11.34 |
|
|
|
| 20 |
A |
1438 |
1375 |
2.16 |
|
|
|
| 21 |
A |
1404 |
1344 |
2.83 |
|
|
|
| 22 |
A |
1392 |
1332 |
2.23 |
|
|
|
| 23 |
A |
1341 |
1283 |
1.02 |
|
|
|
| 24 |
A |
1324 |
1267 |
14.92 |
|
|
|
| 25 |
A |
1290 |
1234 |
7.25 |
|
|
|
| 26 |
A |
1243 |
1189 |
7.78 |
|
|
|
| 27 |
A |
1193 |
1141 |
0.55 |
|
|
|
| 28 |
A |
1159 |
1109 |
3.59 |
|
|
|
| 29 |
A |
1102 |
1055 |
1.40 |
|
|
|
| 30 |
A |
1058 |
1012 |
0.43 |
|
|
|
| 31 |
A |
1040 |
995 |
2.54 |
|
|
|
| 32 |
A |
989 |
946 |
7.27 |
|
|
|
| 33 |
A |
954 |
913 |
0.88 |
|
|
|
| 34 |
A |
906 |
867 |
4.06 |
|
|
|
| 35 |
A |
880 |
842 |
1.25 |
|
|
|
| 36 |
A |
807 |
772 |
0.43 |
|
|
|
| 37 |
A |
791 |
757 |
8.88 |
|
|
|
| 38 |
A |
728 |
697 |
1.56 |
|
|
|
| 39 |
A |
456 |
436 |
0.16 |
|
|
|
| 40 |
A |
400 |
383 |
0.05 |
|
|
|
| 41 |
A |
380 |
364 |
1.14 |
|
|
|
| 42 |
A |
254 |
243 |
0.17 |
|
|
|
| 43 |
A |
244 |
234 |
0.39 |
|
|
|
| 44 |
A |
208 |
199 |
0.33 |
|
|
|
| 45 |
A |
200 |
191 |
2.08 |
|
|
|
| 46 |
A |
170 |
163 |
19.28 |
|
|
|
| 47 |
A |
97 |
92 |
0.64 |
|
|
|
| 48 |
A |
67 |
64 |
6.34 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35391.4 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 33859.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.515 |
|
|
-0.641 |
| 2 |
H |
0.193 |
|
|
0.216 |
| 3 |
H |
0.165 |
|
|
0.145 |
| 4 |
H |
0.168 |
|
|
0.134 |
| 5 |
S |
-0.095 |
|
|
-0.351 |
| 6 |
H |
0.102 |
|
|
0.231 |
| 7 |
C |
-0.462 |
|
|
-0.559 |
| 8 |
H |
0.192 |
|
|
0.204 |
| 9 |
H |
0.200 |
|
|
0.203 |
| 10 |
C |
-0.129 |
|
|
0.463 |
| 11 |
H |
0.171 |
|
|
0.017 |
| 12 |
C |
-0.517 |
|
|
-0.271 |
| 13 |
H |
0.172 |
|
|
0.088 |
| 14 |
H |
0.166 |
|
|
0.062 |
| 15 |
H |
0.170 |
|
|
0.060 |
| 16 |
C |
-0.301 |
|
|
-0.079 |
| 17 |
H |
0.165 |
|
|
0.036 |
| 18 |
H |
0.157 |
|
|
0.041 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.488 |
-0.697 |
0.850 |
1.850 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
1.489 |
-0.722 |
0.875 |
1.872 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-49.279 |
2.452 |
-1.748 |
| y |
2.452 |
-45.550 |
-1.732 |
| z |
-1.748 |
-1.732 |
-47.523 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.743 |
2.452 |
-1.748 |
| y |
2.452 |
2.851 |
-1.732 |
| z |
-1.748 |
-1.732 |
-0.108 |
|
| Polar |
|---|
| 3z2-r2 | -0.216 |
|---|
| x2-y2 | -3.730 |
|---|
| xy | 2.452 |
|---|
| xz | -1.748 |
|---|
| yz | -1.732 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
287.757 |
| (<r2>)1/2 |
16.963 |