Jump to
S1C2
Vibrational Frequencies calculated at CID/6-31G*
Geometric Data calculated at CID/6-31G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -234.862447 |
| Energy at 298.15K | -234.876215 |
| Nuclear repulsion energy | 256.124893 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3252 |
3004 |
70.69 |
|
|
|
| 2 |
A' |
3234 |
2987 |
48.27 |
|
|
|
| 3 |
A' |
3231 |
2984 |
24.41 |
|
|
|
| 4 |
A' |
3223 |
2977 |
12.42 |
|
|
|
| 5 |
A' |
3197 |
2953 |
25.05 |
|
|
|
| 6 |
A' |
3179 |
2937 |
11.91 |
|
|
|
| 7 |
A' |
3155 |
2914 |
27.40 |
|
|
|
| 8 |
A' |
3149 |
2909 |
25.23 |
|
|
|
| 9 |
A' |
1619 |
1496 |
1.54 |
|
|
|
| 10 |
A' |
1603 |
1481 |
2.37 |
|
|
|
| 11 |
A' |
1594 |
1472 |
5.94 |
|
|
|
| 12 |
A' |
1582 |
1461 |
0.02 |
|
|
|
| 13 |
A' |
1520 |
1404 |
2.07 |
|
|
|
| 14 |
A' |
1508 |
1393 |
10.15 |
|
|
|
| 15 |
A' |
1419 |
1311 |
6.14 |
|
|
|
| 16 |
A' |
1396 |
1289 |
0.92 |
|
|
|
| 17 |
A' |
1318 |
1218 |
1.20 |
|
|
|
| 18 |
A' |
1285 |
1187 |
0.21 |
|
|
|
| 19 |
A' |
1074 |
992 |
0.59 |
|
|
|
| 20 |
A' |
1051 |
971 |
0.48 |
|
|
|
| 21 |
A' |
1012 |
934 |
0.40 |
|
|
|
| 22 |
A' |
984 |
909 |
0.41 |
|
|
|
| 23 |
A' |
930 |
859 |
0.44 |
|
|
|
| 24 |
A' |
753 |
695 |
0.86 |
|
|
|
| 25 |
A' |
639 |
590 |
1.18 |
|
|
|
| 26 |
A' |
417 |
385 |
0.34 |
|
|
|
| 27 |
A' |
369 |
340 |
0.01 |
|
|
|
| 28 |
A' |
170 |
157 |
0.00 |
|
|
|
| 29 |
A" |
3230 |
2984 |
57.15 |
|
|
|
| 30 |
A" |
3227 |
2981 |
7.44 |
|
|
|
| 31 |
A" |
3221 |
2976 |
7.53 |
|
|
|
| 32 |
A" |
3175 |
2932 |
66.07 |
|
|
|
| 33 |
A" |
1607 |
1485 |
5.93 |
|
|
|
| 34 |
A" |
1585 |
1464 |
0.49 |
|
|
|
| 35 |
A" |
1581 |
1460 |
3.10 |
|
|
|
| 36 |
A" |
1372 |
1267 |
0.10 |
|
|
|
| 37 |
A" |
1341 |
1239 |
0.51 |
|
|
|
| 38 |
A" |
1307 |
1207 |
0.34 |
|
|
|
| 39 |
A" |
1270 |
1173 |
2.11 |
|
|
|
| 40 |
A" |
1153 |
1065 |
0.05 |
|
|
|
| 41 |
A" |
1048 |
968 |
0.00 |
|
|
|
| 42 |
A" |
1004 |
927 |
1.92 |
|
|
|
| 43 |
A" |
945 |
873 |
0.00 |
|
|
|
| 44 |
A" |
839 |
775 |
0.18 |
|
|
|
| 45 |
A" |
406 |
375 |
0.07 |
|
|
|
| 46 |
A" |
316 |
292 |
0.02 |
|
|
|
| 47 |
A" |
286 |
264 |
0.00 |
|
|
|
| 48 |
A" |
242 |
223 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 38506.3 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 35568.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.098 |
0.400 |
0.000 |
| C2 |
0.362 |
-1.720 |
0.000 |
| C3 |
-0.098 |
-0.716 |
1.075 |
| C4 |
-0.098 |
-0.716 |
-1.075 |
| C5 |
1.209 |
1.183 |
0.000 |
| C6 |
-1.288 |
1.343 |
0.000 |
| H7 |
1.441 |
-1.861 |
0.000 |
| H8 |
-0.109 |
-2.700 |
0.000 |
| H9 |
0.543 |
-0.570 |
1.943 |
| H10 |
0.543 |
-0.570 |
-1.943 |
| H11 |
-1.106 |
-0.935 |
1.424 |
| H12 |
-1.106 |
-0.935 |
-1.424 |
| H13 |
2.074 |
0.520 |
0.000 |
| H14 |
-2.226 |
0.789 |
0.000 |
| H15 |
1.277 |
1.820 |
-0.882 |
| H16 |
1.277 |
1.820 |
0.882 |
| H17 |
-1.277 |
1.985 |
0.881 |
| H18 |
-1.277 |
1.985 |
-0.881 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 2.1690 | 1.5491 | 1.5491 | 1.5232 | 1.5181 | 2.7350 | 3.0999 | 2.2644 | 2.2644 | 2.1967 | 2.1967 | 2.1747 | 2.1632 | 2.1645 | 2.1645 | 2.1633 | 2.1633 |
C2 | 2.1690 | | 1.5409 | 1.5409 | 3.0234 | 3.4785 | 1.0877 | 1.0875 | 2.2652 | 2.2652 | 2.1908 | 2.1908 | 2.8189 | 3.6042 | 3.7608 | 3.7608 | 4.1460 | 4.1460 | C3 | 1.5491 | 1.5409 | | 2.1493 | 2.5430 | 2.6092 | 2.1986 | 2.2565 | 1.0893 | 3.0887 | 1.0894 | 2.7032 | 2.7198 | 2.8190 | 3.4855 | 2.8909 | 2.9534 | 3.5371 | C4 | 1.5491 | 1.5409 | 2.1493 | | 2.5430 | 2.6092 | 2.1986 | 2.2565 | 3.0887 | 1.0893 | 2.7032 | 1.0894 | 2.7198 | 2.8190 | 2.8909 | 3.4855 | 3.5371 | 2.9534 | C5 | 1.5232 | 3.0234 | 2.5430 | 2.5430 | | 2.5017 | 3.0529 | 4.1002 | 2.7003 | 2.7003 | 3.4454 | 3.4454 | 1.0894 | 3.4572 | 1.0902 | 1.0902 | 2.7570 | 2.7570 | C6 | 1.5181 | 3.4785 | 2.6092 | 2.6092 | 2.5017 | | 4.2083 | 4.2110 | 3.2842 | 3.2842 | 2.6920 | 2.6920 | 3.4605 | 1.0894 | 2.7541 | 2.7541 | 1.0906 | 1.0906 | H7 | 2.7350 | 1.0877 | 2.1986 | 2.1986 | 3.0529 | 4.2083 | | 1.7618 | 2.4999 | 2.4999 | 3.0613 | 3.0613 | 2.4642 | 4.5239 | 3.7889 | 3.7889 | 4.7914 | 4.7914 | H8 | 3.0999 | 1.0875 | 2.2565 | 2.2565 | 4.1002 | 4.2110 | 1.7618 | | 2.9560 | 2.9560 | 2.4775 | 2.4775 | 3.8900 | 4.0807 | 4.8091 | 4.8091 | 4.9082 | 4.9082 | H9 | 2.2644 | 2.2652 | 1.0893 | 3.0887 | 2.7003 | 3.2842 | 2.4999 | 2.9560 | | 3.8866 | 1.7672 | 3.7670 | 2.7032 | 3.6454 | 3.7726 | 2.7159 | 3.3119 | 4.2213 | H10 | 2.2644 | 2.2652 | 3.0887 | 1.0893 | 2.7003 | 3.2842 | 2.4999 | 2.9560 | 3.8866 | | 3.7670 | 1.7672 | 2.7032 | 3.6454 | 2.7159 | 3.7726 | 4.2213 | 3.3119 | H11 | 2.1967 | 2.1908 | 1.0894 | 2.7032 | 3.4454 | 2.6920 | 3.0613 | 2.4775 | 1.7672 | 3.7670 | | 2.8476 | 3.7756 | 2.5003 | 4.3111 | 3.6827 | 2.9746 | 3.7240 | H12 | 2.1967 | 2.1908 | 2.7032 | 1.0894 | 3.4454 | 2.6920 | 3.0613 | 2.4775 | 3.7670 | 1.7672 | 2.8476 | | 3.7756 | 2.5003 | 3.6827 | 4.3111 | 3.7240 | 2.9746 | H13 | 2.1747 | 2.8189 | 2.7198 | 2.7198 | 1.0894 | 3.4605 | 2.4642 | 3.8900 | 2.7032 | 2.7032 | 3.7756 | 3.7756 | | 4.3078 | 1.7610 | 1.7610 | 3.7617 | 3.7617 | H14 | 2.1632 | 3.6042 | 2.8190 | 2.8190 | 3.4572 | 1.0894 | 4.5239 | 4.0807 | 3.6454 | 3.6454 | 2.5003 | 2.5003 | 4.3078 | | 3.7567 | 3.7567 | 1.7628 | 1.7628 | H15 | 2.1645 | 3.7608 | 3.4855 | 2.8909 | 1.0902 | 2.7541 | 3.7889 | 4.8091 | 3.7726 | 2.7159 | 4.3111 | 3.6827 | 1.7610 | 3.7567 | | 1.7640 | 3.1084 | 2.5599 | H16 | 2.1645 | 3.7608 | 2.8909 | 3.4855 | 1.0902 | 2.7541 | 3.7889 | 4.8091 | 2.7159 | 3.7726 | 3.6827 | 4.3111 | 1.7610 | 3.7567 | 1.7640 | | 2.5599 | 3.1084 | H17 | 2.1633 | 4.1460 | 2.9534 | 3.5371 | 2.7570 | 1.0906 | 4.7914 | 4.9082 | 3.3119 | 4.2213 | 2.9746 | 3.7240 | 3.7617 | 1.7628 | 3.1084 | 2.5599 | | 1.7627 | H18 | 2.1633 | 4.1460 | 3.5371 | 2.9534 | 2.7570 | 1.0906 | 4.7914 | 4.9082 | 4.2213 | 3.3119 | 3.7240 | 2.9746 | 3.7617 | 1.7628 | 2.5599 | 3.1084 | 1.7627 | |
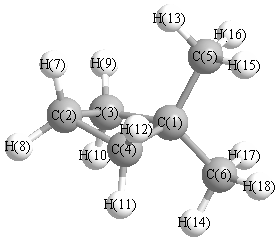 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
C2 |
89.165 |
|
C1 |
C3 |
H9 |
117.177 |
| C1 |
C3 |
H11 |
111.540 |
|
C1 |
C4 |
C2 |
89.165 |
| C1 |
C4 |
H10 |
117.177 |
|
C1 |
C4 |
H12 |
111.540 |
| C1 |
C5 |
H13 |
111.615 |
|
C1 |
C5 |
H15 |
110.741 |
| C1 |
C5 |
H16 |
110.741 |
|
C1 |
C6 |
H14 |
111.053 |
| C1 |
C6 |
H17 |
110.988 |
|
C1 |
C6 |
H18 |
110.988 |
| C2 |
C3 |
H9 |
117.889 |
|
C2 |
C3 |
H11 |
111.646 |
| C2 |
C4 |
H10 |
117.889 |
|
C2 |
C4 |
H12 |
111.646 |
| C3 |
C1 |
C4 |
87.851 |
|
C3 |
C1 |
C5 |
111.724 |
| C3 |
C1 |
C6 |
116.567 |
|
C3 |
C2 |
C4 |
88.442 |
| C3 |
C2 |
H7 |
112.384 |
|
C3 |
C2 |
H8 |
117.256 |
| C4 |
C1 |
C5 |
111.724 |
|
C4 |
C1 |
C6 |
116.567 |
| C4 |
C2 |
H7 |
112.384 |
|
C4 |
C2 |
H8 |
117.256 |
| C5 |
C1 |
C6 |
110.690 |
|
H7 |
C2 |
H8 |
108.181 |
| H9 |
C3 |
H11 |
108.419 |
|
H10 |
C4 |
H12 |
108.419 |
| H13 |
C5 |
H15 |
107.797 |
|
H13 |
C5 |
H16 |
107.797 |
| H14 |
C6 |
H17 |
107.923 |
|
H14 |
C6 |
H18 |
107.923 |
| H15 |
C5 |
H16 |
108.003 |
|
H17 |
C6 |
H18 |
107.825 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
