Jump to
S1C2
Vibrational Frequencies calculated at QCISD/6-31G*
Geometric Data calculated at QCISD/6-31G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/6-31G*
| | hartrees |
|---|
| Energy at 0K | -235.028240 |
| Energy at 298.15K | -235.041829 |
| Nuclear repulsion energy | 254.803179 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3145 |
2995 |
66.95 |
|
|
|
| 2 |
A' |
3130 |
2981 |
47.05 |
|
|
|
| 3 |
A' |
3126 |
2977 |
22.88 |
|
|
|
| 4 |
A' |
3117 |
2969 |
13.72 |
|
|
|
| 5 |
A' |
3092 |
2944 |
25.17 |
|
|
|
| 6 |
A' |
3074 |
2927 |
10.48 |
|
|
|
| 7 |
A' |
3052 |
2906 |
26.76 |
|
|
|
| 8 |
A' |
3046 |
2901 |
24.48 |
|
|
|
| 9 |
A' |
1567 |
1492 |
1.61 |
|
|
|
| 10 |
A' |
1553 |
1479 |
2.19 |
|
|
|
| 11 |
A' |
1545 |
1471 |
6.02 |
|
|
|
| 12 |
A' |
1533 |
1460 |
0.02 |
|
|
|
| 13 |
A' |
1468 |
1398 |
1.93 |
|
|
|
| 14 |
A' |
1454 |
1385 |
9.76 |
|
|
|
| 15 |
A' |
1365 |
1300 |
5.99 |
|
|
|
| 16 |
A' |
1348 |
1284 |
1.29 |
|
|
|
| 17 |
A' |
1273 |
1213 |
0.82 |
|
|
|
| 18 |
A' |
1241 |
1182 |
0.14 |
|
|
|
| 19 |
A' |
1040 |
991 |
0.64 |
|
|
|
| 20 |
A' |
1018 |
969 |
0.41 |
|
|
|
| 21 |
A' |
979 |
933 |
0.35 |
|
|
|
| 22 |
A' |
952 |
906 |
0.42 |
|
|
|
| 23 |
A' |
903 |
860 |
0.43 |
|
|
|
| 24 |
A' |
729 |
695 |
0.84 |
|
|
|
| 25 |
A' |
620 |
590 |
1.16 |
|
|
|
| 26 |
A' |
406 |
386 |
0.37 |
|
|
|
| 27 |
A' |
360 |
343 |
0.01 |
|
|
|
| 28 |
A' |
174 |
166 |
0.00 |
|
|
|
| 29 |
A" |
3125 |
2976 |
62.24 |
|
|
|
| 30 |
A" |
3122 |
2973 |
0.80 |
|
|
|
| 31 |
A" |
3117 |
2969 |
8.12 |
|
|
|
| 32 |
A" |
3069 |
2923 |
61.87 |
|
|
|
| 33 |
A" |
1556 |
1482 |
5.88 |
|
|
|
| 34 |
A" |
1535 |
1462 |
0.38 |
|
|
|
| 35 |
A" |
1530 |
1457 |
2.79 |
|
|
|
| 36 |
A" |
1317 |
1254 |
0.11 |
|
|
|
| 37 |
A" |
1288 |
1227 |
0.46 |
|
|
|
| 38 |
A" |
1261 |
1201 |
0.27 |
|
|
|
| 39 |
A" |
1222 |
1164 |
2.10 |
|
|
|
| 40 |
A" |
1113 |
1060 |
0.04 |
|
|
|
| 41 |
A" |
1013 |
965 |
0.00 |
|
|
|
| 42 |
A" |
969 |
922 |
1.77 |
|
|
|
| 43 |
A" |
910 |
867 |
0.00 |
|
|
|
| 44 |
A" |
813 |
774 |
0.18 |
|
|
|
| 45 |
A" |
393 |
374 |
0.06 |
|
|
|
| 46 |
A" |
306 |
291 |
0.02 |
|
|
|
| 47 |
A" |
275 |
262 |
0.00 |
|
|
|
| 48 |
A" |
236 |
225 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 37238.7 cm
-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 35462.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.101 |
0.402 |
0.000 |
| C2 |
0.376 |
-1.723 |
0.000 |
| C3 |
-0.101 |
-0.720 |
1.080 |
| C4 |
-0.101 |
-0.720 |
-1.080 |
| C5 |
1.213 |
1.185 |
0.000 |
| C6 |
-1.294 |
1.350 |
0.000 |
| H7 |
1.465 |
-1.846 |
0.000 |
| H8 |
-0.081 |
-2.720 |
0.000 |
| H9 |
0.535 |
-0.569 |
1.961 |
| H10 |
0.535 |
-0.569 |
-1.961 |
| H11 |
-1.120 |
-0.947 |
1.421 |
| H12 |
-1.120 |
-0.947 |
-1.421 |
| H13 |
2.082 |
0.514 |
0.000 |
| H14 |
-2.239 |
0.791 |
0.000 |
| H15 |
1.282 |
1.827 |
-0.889 |
| H16 |
1.282 |
1.827 |
0.889 |
| H17 |
-1.282 |
1.997 |
0.889 |
| H18 |
-1.282 |
1.997 |
-0.889 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 2.1788 | 1.5573 | 1.5573 | 1.5293 | 1.5235 | 2.7405 | 3.1221 | 2.2794 | 2.2794 | 2.2088 | 2.2088 | 2.1858 | 2.1731 | 2.1756 | 2.1756 | 2.1741 | 2.1741 |
C2 | 2.1788 | | 1.5493 | 1.5493 | 3.0264 | 3.4979 | 1.0964 | 1.0962 | 2.2815 | 2.2815 | 2.2043 | 2.2043 | 2.8134 | 3.6280 | 3.7703 | 3.7703 | 4.1690 | 4.1690 | C3 | 1.5573 | 1.5493 | | 2.1592 | 2.5535 | 2.6217 | 2.2110 | 2.2727 | 1.0980 | 3.1105 | 1.0984 | 2.7095 | 2.7300 | 2.8319 | 3.5036 | 2.9043 | 2.9685 | 3.5566 | C4 | 1.5573 | 1.5493 | 2.1592 | | 2.5535 | 2.6217 | 2.2110 | 2.2727 | 3.1105 | 1.0980 | 2.7095 | 1.0984 | 2.7300 | 2.8319 | 2.9043 | 3.5036 | 3.5566 | 2.9685 | C5 | 1.5293 | 3.0264 | 2.5535 | 2.5535 | | 2.5123 | 3.0415 | 4.1134 | 2.7168 | 2.7168 | 3.4652 | 3.4652 | 1.0979 | 3.4744 | 1.0988 | 1.0988 | 2.7700 | 2.7700 | C6 | 1.5235 | 3.4979 | 2.6217 | 2.6217 | 2.5123 | | 4.2226 | 4.2466 | 3.2980 | 3.2980 | 2.7068 | 2.7068 | 3.4779 | 1.0981 | 2.7669 | 2.7669 | 1.0992 | 1.0992 | H7 | 2.7405 | 1.0964 | 2.2110 | 2.2110 | 3.0415 | 4.2226 | | 1.7765 | 2.5185 | 2.5185 | 3.0841 | 3.0841 | 2.4390 | 4.5475 | 3.7832 | 3.7832 | 4.8070 | 4.8070 | H8 | 3.1221 | 1.0962 | 2.2727 | 2.2727 | 4.1134 | 4.2466 | 1.7765 | | 2.9754 | 2.9754 | 2.4976 | 2.4976 | 3.8903 | 4.1209 | 4.8289 | 4.8289 | 4.9475 | 4.9475 | H9 | 2.2794 | 2.2815 | 1.0980 | 3.1105 | 2.7168 | 3.2980 | 2.5185 | 2.9754 | | 3.9226 | 1.7825 | 3.7845 | 2.7221 | 3.6600 | 3.7976 | 2.7285 | 3.3223 | 4.2437 | H10 | 2.2794 | 2.2815 | 3.1105 | 1.0980 | 2.7168 | 3.2980 | 2.5185 | 2.9754 | 3.9226 | | 3.7845 | 1.7825 | 2.7221 | 3.6600 | 2.7285 | 3.7976 | 4.2437 | 3.3223 | H11 | 2.2088 | 2.2043 | 1.0984 | 2.7095 | 3.4652 | 2.7068 | 3.0841 | 2.4976 | 1.7825 | 3.7845 | | 2.8412 | 3.7955 | 2.5086 | 4.3362 | 3.7080 | 2.9965 | 3.7454 | H12 | 2.2088 | 2.2043 | 2.7095 | 1.0984 | 3.4652 | 2.7068 | 3.0841 | 2.4976 | 3.7845 | 1.7825 | 2.8412 | | 3.7955 | 2.5086 | 3.7080 | 4.3362 | 3.7454 | 2.9965 | H13 | 2.1858 | 2.8134 | 2.7300 | 2.7300 | 1.0979 | 3.4779 | 2.4390 | 3.8903 | 2.7221 | 2.7221 | 3.7955 | 3.7955 | | 4.3299 | 1.7758 | 1.7758 | 3.7821 | 3.7821 | H14 | 2.1731 | 3.6280 | 2.8319 | 2.8319 | 3.4744 | 1.0981 | 4.5475 | 4.1209 | 3.6600 | 3.6600 | 2.5086 | 2.5086 | 4.3299 | | 3.7768 | 3.7768 | 1.7777 | 1.7777 | H15 | 2.1756 | 3.7703 | 3.5036 | 2.9043 | 1.0988 | 2.7669 | 3.7832 | 4.8289 | 3.7976 | 2.7285 | 4.3362 | 3.7080 | 1.7758 | 3.7768 | | 1.7786 | 3.1250 | 2.5699 | H16 | 2.1756 | 3.7703 | 2.9043 | 3.5036 | 1.0988 | 2.7669 | 3.7832 | 4.8289 | 2.7285 | 3.7976 | 3.7080 | 4.3362 | 1.7758 | 3.7768 | 1.7786 | | 2.5699 | 3.1250 | H17 | 2.1741 | 4.1690 | 2.9685 | 3.5566 | 2.7700 | 1.0992 | 4.8070 | 4.9475 | 3.3223 | 4.2437 | 2.9965 | 3.7454 | 3.7821 | 1.7777 | 3.1250 | 2.5699 | | 1.7772 | H18 | 2.1741 | 4.1690 | 3.5566 | 2.9685 | 2.7700 | 1.0992 | 4.8070 | 4.9475 | 4.2437 | 3.3223 | 3.7454 | 2.9965 | 3.7821 | 1.7777 | 2.5699 | 3.1250 | 1.7772 | |
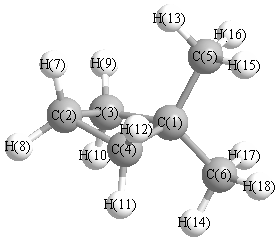 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
C2 |
89.067 |
|
C1 |
C3 |
H9 |
117.229 |
| C1 |
C3 |
H11 |
111.380 |
|
C1 |
C4 |
C2 |
89.067 |
| C1 |
C4 |
H10 |
117.229 |
|
C1 |
C4 |
H12 |
111.380 |
| C1 |
C5 |
H13 |
111.547 |
|
C1 |
C5 |
H15 |
110.679 |
| C1 |
C5 |
H16 |
110.679 |
|
C1 |
C6 |
H14 |
110.937 |
| C1 |
C6 |
H17 |
110.945 |
|
C1 |
C6 |
H18 |
110.945 |
| C2 |
C3 |
H9 |
118.036 |
|
C2 |
C3 |
H11 |
111.583 |
| C2 |
C4 |
H10 |
118.036 |
|
C2 |
C4 |
H12 |
111.583 |
| C3 |
C1 |
C4 |
87.779 |
|
C3 |
C1 |
C5 |
111.640 |
| C3 |
C1 |
C6 |
116.632 |
|
C3 |
C2 |
C4 |
88.346 |
| C3 |
C2 |
H7 |
112.240 |
|
C3 |
C2 |
H8 |
117.400 |
| C4 |
C1 |
C5 |
111.640 |
|
C4 |
C1 |
C6 |
116.632 |
| C4 |
C2 |
H7 |
112.240 |
|
C4 |
C2 |
H8 |
117.400 |
| C5 |
C1 |
C6 |
110.761 |
|
H7 |
C2 |
H8 |
108.236 |
| H9 |
C3 |
H11 |
108.495 |
|
H10 |
C4 |
H12 |
108.495 |
| H13 |
C5 |
H15 |
107.870 |
|
H13 |
C5 |
H16 |
107.870 |
| H14 |
C6 |
H17 |
108.000 |
|
H14 |
C6 |
H18 |
108.000 |
| H15 |
C5 |
H16 |
108.058 |
|
H17 |
C6 |
H18 |
107.881 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
