return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H9N (Butanenitrile, 2-methyl-)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-250.395909
Energy at 298.15K-250.405384
HF Energy-250.126295
Nuclear repulsion energy219.525938
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3183 3022 18.01      
2 A 3174 3013 18.45      
3 A 3166 3005 35.55      
4 A 3156 2996 32.23      
5 A 3127 2968 7.75      
6 A 3096 2939 12.05      
7 A 3087 2930 27.55      
8 A 3083 2927 11.21      
9 A 3070 2914 3.45      
10 A 2296 2179 2.42      
11 A 1561 1482 2.85      
12 A 1554 1475 8.41      
13 A 1549 1470 6.00      
14 A 1544 1466 7.94      
15 A 1535 1457 0.14      
16 A 1463 1389 1.92      
17 A 1457 1383 4.35      
18 A 1417 1345 0.52      
19 A 1381 1311 1.38      
20 A 1350 1282 3.06      
21 A 1316 1249 1.46      
22 A 1216 1154 2.14      
23 A 1163 1104 0.11      
24 A 1142 1084 6.33      
25 A 1067 1013 1.06      
26 A 1017 965 0.48      
27 A 995 945 4.80      
28 A 914 867 0.13      
29 A 828 785 0.76      
30 A 780 741 2.88      
31 A 564 535 0.12      
32 A 538 511 1.01      
33 A 404 383 0.28      
34 A 327 310 0.72      
35 A 289 274 0.13      
36 A 226 214 0.10      
37 A 207 196 2.42      
38 A 163 155 5.55      
39 A 88 83 1.47      

Unscaled Zero Point Vibrational Energy (zpe) 29245.4 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 27759.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.21572 0.07322 0.05856

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.493 -0.324 0.120
N2 -2.557 -0.737 -0.114
C3 -0.116 1.657 -0.150
H4 -0.272 1.666 -1.230
H5 -0.894 2.265 0.316
H6 0.845 2.115 0.071
C7 2.351 -0.383 0.046
H8 2.536 -0.345 1.122
H9 3.018 -1.129 -0.391
H10 2.613 0.582 -0.384
C11 0.882 -0.759 -0.234
H12 0.717 -0.813 -1.312
H13 0.711 -1.755 0.184
C14 -0.182 0.200 0.379
H15 -0.030 0.223 1.463

Atom - Atom Distances (Å)
  C1 N2 C3 H4 H5 H6 C7 H8 H9 H10 C11 H12 H13 C14 H15
C11.16492.42722.69732.66443.37913.84594.15244.61064.23482.44092.67852.62851.43592.0608
N21.16493.41873.49893.45834.44314.92395.25595.59545.34223.44163.48703.43562.60063.1303
C32.42723.41871.09171.09171.08773.20763.55864.20022.94252.61552.85423.52621.55062.1597
H42.69733.49891.09171.77091.77283.56524.17954.39803.19602.86512.67103.83032.17863.0654
H52.66443.45831.09171.77091.76274.19744.38555.22713.95233.55013.83694.33032.18492.4964
H63.37914.44311.08771.77281.76272.91723.16533.93182.38382.89063.24123.87392.19422.5063
C73.84594.92393.20763.56524.19742.91721.09291.09141.08801.54202.16802.14332.62082.8366
H84.15245.25593.55864.17954.38553.16531.09291.77051.77032.17843.07472.48992.87042.6502
H94.61065.59544.20024.39805.22713.93181.09141.77051.75852.17282.49842.45853.54923.8148
H104.23485.34222.94253.19603.95232.38381.08801.77031.75852.19442.53043.06612.92183.2438
C112.44093.44162.61552.86513.55012.89061.54202.17842.17282.19441.09211.09341.55832.1626
H122.67853.48702.85422.67103.83693.24122.16803.07472.49842.53041.09211.76752.16653.0551
H132.62853.43563.52623.83034.33033.87392.14332.48992.45853.06611.09341.76752.15812.4693
C141.43592.60061.55062.17862.18492.19422.62082.87043.54922.92181.55832.16652.15811.0955
H152.06083.13032.15973.06542.49642.50632.83662.65023.81483.24382.16263.05512.46931.0955

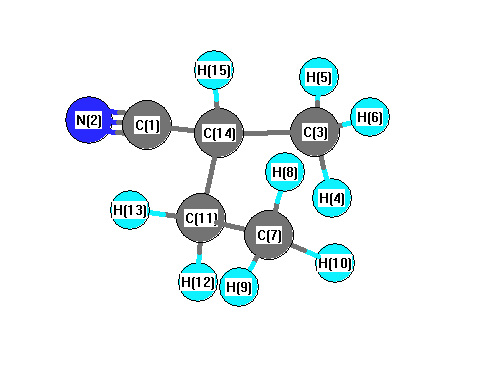
picture of Butanenitrile, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C14 C3 108.664 C1 C14 C11 109.148
C1 C14 H15 108.245 N2 C1 C14 178.670
C3 C14 C11 114.553 C3 C14 H15 108.174
H4 C3 H5 108.400 H4 C3 H6 108.864
H4 C3 C14 109.853 H5 C3 H6 107.964
H5 C3 C14 110.350 H6 C3 C14 111.334
C7 C11 H12 109.599 C7 C11 H13 107.610
C7 C11 C14 115.411 H8 C7 H9 108.301
H8 C7 H10 108.532 H8 C7 C11 110.367
H9 C7 H10 107.575 H9 C7 C11 110.012
H10 C7 C11 111.940 C11 C14 H15 107.886
H12 C11 H13 107.951 H12 C11 C14 108.368
H13 C11 C14 107.656
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C 0.335      
2 N -0.481      
3 C -0.465      
4 H 0.173      
5 H 0.179      
6 H 0.171      
7 C -0.473      
8 H 0.156      
9 H 0.168      
10 H 0.161      
11 C -0.278      
12 H 0.166      
13 H 0.172      
14 C -0.179      
15 H 0.195      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 9.887 0.484 0.255
y 0.484 7.517 0.110
z 0.255 0.110 6.632


<r2> (average value of r2) Å2
<r2> 199.762
(<r2>)1/2 14.134